return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SO2NH2 (methanesulfonamide)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-643.212864
Energy at 298.15K-643.220704
HF Energy-642.384966
Nuclear repulsion energy276.632708
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3523 3355 35.39      
2 A' 3210 3057 0.43      
3 A' 3107 2959 0.30      
4 A' 1659 1580 48.22      
5 A' 1505 1433 6.70      
6 A' 1424 1356 15.01      
7 A' 1160 1105 186.81      
8 A' 1040 990 27.14      
9 A' 938 894 112.38      
10 A' 765 729 45.92      
11 A' 731 697 220.19      
12 A' 505 481 45.84      
13 A' 481 458 25.66      
14 A' 298 284 5.35      
15 A" 3631 3458 44.03      
16 A" 3219 3065 0.70      
17 A" 1505 1433 2.44      
18 A" 1385 1319 232.23      
19 A" 1145 1091 4.92      
20 A" 1026 977 0.78      
21 A" 404 385 0.03      
22 A" 332 316 3.97      
23 A" 236 224 0.67      
24 A" 167 159 49.50      

Unscaled Zero Point Vibrational Energy (zpe) 16697.6 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 15901.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.15548 0.14485 0.14204

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.672 -0.057 0.000
S2 0.106 -0.134 0.000
N3 0.532 1.495 0.000
O4 0.532 -0.702 1.279
O5 0.532 -0.702 -1.279
H6 -2.040 -1.086 0.000
H7 -2.003 0.467 0.899
H8 -2.003 0.467 -0.899
H9 1.073 1.704 0.839
H10 1.073 1.704 -0.839

Atom - Atom Distances (Å)
  C1 S2 N3 O4 O5 H6 H7 H8 H9 H10
C11.77892.69502.62842.62841.09361.09231.09233.36653.3665
S21.77891.68381.46271.46272.34742.36982.36982.23982.2398
N32.69501.68382.54152.54153.64392.87932.87931.01951.0195
O42.62841.46272.54152.55772.89842.81723.54082.50433.2500
O52.62841.46272.54152.55772.89843.54082.81723.25002.5043
H61.09362.34743.64392.89842.89841.79551.79554.26344.2634
H71.09232.36982.87932.81723.54081.79551.79883.31523.7428
H81.09232.36982.87933.54082.81721.79551.79883.74283.3152
H93.36652.23981.01952.50433.25004.26343.31523.74281.6776
H103.36652.23981.01953.25002.50434.26343.74283.31521.6776

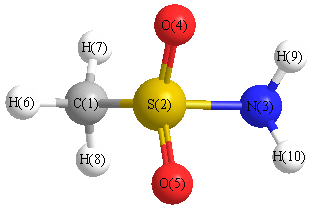
picture of methanesulfonamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 N3 102.173 C1 S2 O4 107.959
C1 S2 O5 107.959 S2 C1 H6 107.190
S2 C1 H7 108.895 S2 C1 H8 108.895
S2 N3 H9 109.430 S2 N3 H10 109.430
N3 S2 O4 107.541 N3 S2 O5 107.541
O4 S2 O5 121.927 H6 C1 H7 110.451
H6 C1 H8 110.451 H7 C1 H8 110.861
H9 N3 H10 110.720
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability