return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H7 (cycloheptatrienyl radical)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2A2
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-270.563150
Energy at 298.15K-270.569774
HF Energy-270.276138
Nuclear repulsion energy257.949181
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3228 3064 8.09      
2 A1 3219 3055 27.09      
3 A1 3202 3039 25.08      
4 A1 3177 3016 7.03      
5 A1 1675 1590 8.14      
6 A1 1630 1547 9.59      
7 A1 1512 1435 6.06      
8 A1 1312 1246 0.64      
9 A1 1277 1212 3.40      
10 A1 981 932 1.59      
11 A1 922 875 0.98      
12 A1 866 822 0.19      
13 A1 422 400 0.31      
14 A2 1013 961 0.00      
15 A2 973 924 0.00      
16 A2 771 732 0.00      
17 A2 565 537 0.00      
18 A2 292 277 0.00      
19 B1 977 928 0.52      
20 B1 838 796 0.01      
21 B1 748 710 2.20      
22 B1 660 626 57.72      
23 B1 510 484 38.14      
24 B1 160 152 0.01      
25 B2 3216 3053 55.29      
26 B2 3185 3023 2.51      
27 B2 3177 3016 9.89      
28 B2 1644 1561 0.56      
29 B2 1518 1441 3.67      
30 B2 1440 1367 0.00      
31 B2 1333 1265 0.35      
32 B2 1251 1188 0.07      
33 B2 993 943 1.10      
34 B2 923 876 4.47      
35 B2 462 439 13.89      
36 B2 420 399 19.38      

Unscaled Zero Point Vibrational Energy (zpe) 25246.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23964.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.12414 0.12401 0.06204

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.628
C2 0.000 1.265 0.995
C3 0.000 -1.265 0.995
C4 0.000 1.588 -0.343
C5 0.000 -1.588 -0.343
C6 0.000 0.683 -1.466
C7 0.000 -0.683 -1.466
H8 0.000 0.000 2.713
H9 0.000 2.120 1.668
H10 0.000 -2.120 1.668
H11 0.000 2.649 -0.572
H12 0.000 -2.649 -0.572
H13 0.000 1.142 -2.452
H14 0.000 -1.142 -2.452

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14
C11.41431.41432.53162.53163.16913.16911.08462.12062.12063.44413.44414.23704.2370
C21.41432.52901.37663.15082.52893.13862.13321.08873.45102.09144.21593.44894.2039
C31.41432.52903.15081.37663.13862.52892.13323.45101.08874.21592.09144.20393.4489
C42.53161.37663.15083.17611.44232.53373.44412.08044.21851.08564.24342.15523.4498
C52.53163.15081.37663.17612.53371.44233.44414.21852.08044.24341.08563.44982.1552
C63.16912.52893.13861.44232.53371.36634.23463.44804.20502.15953.45001.08732.0745
C73.16913.13862.52892.53371.44231.36634.23464.20503.44803.45002.15952.07451.0873
H81.08462.13322.13323.44413.44414.23464.23462.36372.36374.22054.22055.28955.2895
H92.12061.08873.45102.08044.21853.44804.20502.36374.24042.30225.26944.23435.2551
H102.12063.45101.08874.21852.08044.20503.44802.36374.24045.26942.30225.25514.2343
H113.44412.09144.21591.08564.24342.15953.45004.22052.30225.26945.29822.40874.2316
H123.44414.21592.09144.24341.08563.45002.15954.22055.26942.30225.29824.23162.4087
H134.23703.44894.20392.15523.44981.08732.07455.28954.23435.25512.40874.23162.2846
H144.23704.20393.44893.44982.15522.07451.08735.28955.25514.23434.23162.40872.2846

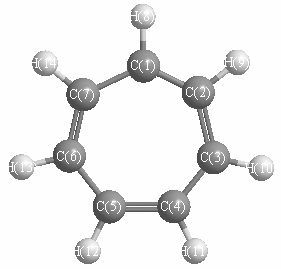
picture of cycloheptatrienyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 26.608 C1 C2 H9 115.202
C1 C7 C6 77.551 C1 C7 H14 167.470
C2 C1 C7 75.841 C2 C1 H8 116.608
C2 C3 C4 25.130 C2 C3 H10 141.810
C3 C2 H9 141.810 C3 C4 C5 25.130
C3 C4 H11 167.070 C4 C3 H10 166.940
C4 C5 C6 26.311 C4 C5 H12 167.800
C5 C4 H11 167.800 C5 C6 C7 26.311
C5 C6 H13 141.291 C6 C5 H12 141.489
C6 C7 H14 114.979 C7 C1 H8 167.551
C7 C6 H13 114.979
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.196      
2 C -0.117      
3 C -0.117      
4 C -0.182      
5 C -0.182      
6 C -0.148      
7 C -0.148      
8 H 0.153      
9 H 0.158      
10 H 0.158      
11 H 0.154      
12 H 0.154      
13 H 0.156      
14 H 0.156      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 3.557 0.000 0.000
y 0.000 13.156 0.000
z 0.000 0.000 13.319


<r2> (average value of r2) Å2
<r2> 186.068
(<r2>)1/2 13.641