return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSSCH3 (dimethyl trisulfide)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-1272.454312
Energy at 298.15K-1272.460558
Nuclear repulsion energy363.157799
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3188 3036 7.94      
2 A 3175 3023 0.18      
3 A 3084 2937 3.76      
4 A 1529 1456 7.83      
5 A 1509 1437 8.49      
6 A 1421 1353 0.19      
7 A 1027 978 0.03      
8 A 1024 975 4.78      
9 A 739 704 0.07      
10 A 495 472 0.39      
11 A 272 259 0.46      
12 A 192 183 1.74      
13 A 159 151 1.15      
14 A 63 60 2.36      
15 B 3188 3036 3.84      
16 B 3175 3023 7.59      
17 B 3084 2937 30.16      
18 B 1528 1455 12.46      
19 B 1509 1437 9.28      
20 B 1423 1356 5.18      
21 B 1026 977 2.85      
22 B 1020 971 15.09      
23 B 738 703 0.98      
24 B 499 475 10.73      
25 B 279 266 1.18      
26 B 182 173 0.67      
27 B 95 91 3.47      

Unscaled Zero Point Vibrational Energy (zpe) 17811.1 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 16961.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.14895 0.04553 0.04528

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 1.085
S2 0.000 1.661 -0.158
S3 0.000 -1.661 -0.158
C4 1.750 1.747 -0.647
C5 -1.750 -1.747 -0.647
H6 1.855 2.622 -1.298
H7 -1.855 -2.622 -1.298
H8 2.393 1.867 0.229
H9 2.036 0.850 -1.204
H10 -2.393 -1.867 0.229
H11 -2.036 -0.850 -1.204

Atom - Atom Distances (Å)
  S1 S2 S3 C4 C5 H6 H7 H8 H9 H10 H11
S12.07472.07473.01923.01923.99993.99993.15373.17963.15373.1796
S22.07473.32151.81883.86212.38044.80472.43302.42824.28043.3979
S32.07473.32153.86211.81884.80472.38044.28043.39792.43302.4282
C43.01921.81883.86214.94581.09575.70211.09331.09415.56734.6251
C53.01923.86211.81884.94585.70211.09575.56734.62511.09331.0941
H63.99992.38044.80471.09575.70216.42421.78651.78326.36665.2164
H73.99994.80472.38045.70211.09576.42426.36665.21641.78651.7832
H83.15372.43304.28041.09335.56731.78656.36661.79306.07055.3904
H93.17962.42823.39791.09414.62511.78325.21641.79305.39044.4130
H103.15374.28042.43305.56731.09336.36661.78656.07055.39041.7930
H113.17963.39792.42824.62511.09415.21641.78325.39044.41301.7930

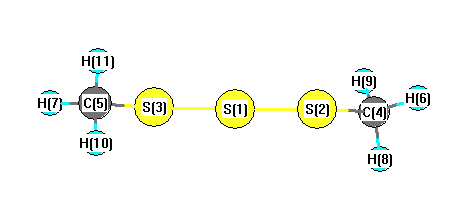
picture of dimethyl trisulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 C4 101.487 S1 S3 C5 101.487
S2 S1 S3 106.355 S2 C4 H6 106.889
S2 C4 H8 110.863 S2 C4 H9 110.455
S3 C5 H7 106.889 S3 C5 H10 110.863
S3 C5 H11 110.455 H6 C4 H8 109.406
H6 C4 H9 109.046 H7 C5 H10 109.406
H7 C5 H11 109.046 H8 C4 H9 110.107
H10 C5 H11 110.107
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability