return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4F2 (orthodifluorobenzene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-430.301922
Energy at 298.15K-430.306605
HF Energy-429.940638
Nuclear repulsion energy346.866968
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3255 3090 1.90      
2 A1 3241 3077 10.82      
3 A1 1677 1591 23.11      
4 A1 1573 1493 176.04      
5 A1 1382 1312 0.44      
6 A1 1325 1258 106.20      
7 A1 1196 1135 0.67      
8 A1 1060 1006 5.73      
9 A1 785 745 28.80      
10 A1 583 554 3.76      
11 A1 280 266 0.30      
12 A2 943 895 0.00      
13 A2 844 801 0.00      
14 A2 626 594 0.00      
15 A2 553 525 0.00      
16 A2 191 181 0.00      
17 B1 921 874 5.44      
18 B1 767 728 73.96      
19 B1 460 436 1.19      
20 B1 297 282 0.04      
21 B2 3251 3086 4.68      
22 B2 3229 3065 1.43      
23 B2 1676 1591 7.31      
24 B2 1514 1437 11.23      
25 B2 1308 1241 6.95      
26 B2 1247 1184 31.90      
27 B2 1138 1080 18.63      
28 B2 865 821 15.29      
29 B2 552 524 3.39      
30 B2 441 419 0.24      

Unscaled Zero Point Vibrational Energy (zpe) 18588.3 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 17644.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.10875 0.07386 0.04399

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.695 -0.535
C2 0.000 -0.695 -0.535
C3 0.000 -1.398 0.661
C4 0.000 -0.698 1.867
C5 0.000 0.698 1.867
C6 0.000 1.398 0.661
F7 0.000 1.350 -1.710
F8 0.000 -1.350 -1.710
H9 0.000 -2.480 0.627
H10 0.000 -1.245 2.803
H11 0.000 1.245 2.803
H12 0.000 2.480 0.627

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 H9 H10 H11 H12
C11.39002.41052.77712.40241.38741.34492.35853.38173.86093.38322.1307
C21.39001.38742.40242.77712.41052.35851.34492.13073.38323.86093.3817
C32.41051.38741.39442.41802.79543.62932.37131.08332.14733.40163.8783
C42.77712.40241.39441.39612.41804.12193.63592.17121.08382.15653.4117
C52.40242.77712.41801.39611.39443.63594.12193.41172.15651.08382.1712
C61.38742.41052.79542.41801.39442.37133.62933.87833.40162.14731.0833
F71.34492.35853.62934.12193.63592.37132.70044.48745.20574.51392.5962
F82.35851.34492.37133.63594.12193.62932.70042.59624.51395.20574.4874
H93.38172.13071.08332.17123.41173.87834.48742.59622.50194.31414.9610
H103.86093.38322.14731.08382.15653.40165.20574.51392.50192.48984.3141
H113.38323.86093.40162.15651.08382.14734.51395.20574.31412.48982.5019
H122.13073.38173.87833.41172.17121.08332.59624.48744.96104.31412.5019

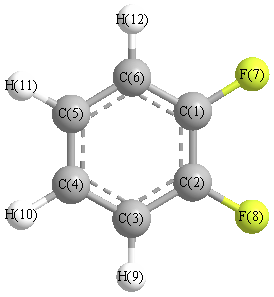
picture of orthodifluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.429 C1 C2 F8 119.153
C1 C6 C5 119.453 C1 C6 H12 118.650
C2 C1 C6 120.429 C2 C1 F7 119.153
C2 C3 C4 119.453 C2 C3 H9 118.650
C3 C2 F8 120.418 C3 C4 C5 120.118
C3 C4 H10 119.581 C4 C3 H9 121.896
C4 C5 C6 120.118 C4 C5 H11 120.301
C5 C4 H10 120.301 C5 C6 H12 121.896
C6 C1 F7 120.418 C6 C5 H11 119.581
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.352      
2 C 0.352      
3 C -0.228      
4 C -0.160      
5 C -0.160      
6 C -0.228      
7 F -0.330      
8 F -0.330      
9 H 0.190      
10 H 0.175      
11 H 0.175      
12 H 0.190      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 3.393 0.000 0.000
y 0.000 10.447 0.000
z 0.000 0.000 10.760


<r2> (average value of r2) Å2
<r2> 230.125
(<r2>)1/2 15.170