return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H11NO (Diethylhydroxylamine)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-288.618795
Energy at 298.15K-288.632286
HF Energy-288.330568
Nuclear repulsion energy263.717745
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3514 3336 4.96      
2 A' 3192 3029 37.17      
3 A' 3170 3009 36.74      
4 A' 3119 2961 39.24      
5 A' 3096 2939 2.35      
6 A' 2952 2802 145.38      
7 A' 1563 1484 8.06      
8 A' 1541 1463 0.19      
9 A' 1536 1458 16.31      
10 A' 1474 1399 16.30      
11 A' 1450 1376 38.68      
12 A' 1431 1358 2.06      
13 A' 1335 1267 3.84      
14 A' 1214 1153 0.26      
15 A' 1194 1133 12.33      
16 A' 1097 1042 2.79      
17 A' 964 915 11.96      
18 A' 848 805 2.08      
19 A' 785 745 18.95      
20 A' 471 447 4.10      
21 A' 431 409 0.08      
22 A' 222 210 0.12      
23 A' 213 202 2.29      
24 A' 112 107 0.43      
25 A" 3191 3029 0.14      
26 A" 3170 3009 13.22      
27 A" 3107 2949 0.00      
28 A" 3095 2938 42.91      
29 A" 2940 2791 4.88      
30 A" 1561 1481 0.47      
31 A" 1537 1459 1.08      
32 A" 1529 1451 2.18      
33 A" 1455 1381 21.01      
34 A" 1414 1343 32.31      
35 A" 1329 1261 1.71      
36 A" 1230 1167 0.55      
37 A" 1122 1065 1.32      
38 A" 1100 1045 23.10      
39 A" 973 924 0.05      
40 A" 826 784 0.01      
41 A" 556 528 27.67      
42 A" 431 410 94.82      
43 A" 351 333 3.85      
44 A" 256 243 0.02      
45 A" 141 133 0.17      

Unscaled Zero Point Vibrational Energy (zpe) 34117.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 32384.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.25711 0.06736 0.05905

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -0.184 -0.161 0.000
O2 1.011 -0.900 0.000
H3 1.782 -0.285 0.000
C4 -0.247 -0.185 2.493
C5 -0.247 -0.185 -2.493
C6 -0.247 0.664 1.219
C7 -0.247 0.664 -1.219
H8 0.665 -0.777 2.553
H9 0.665 -0.777 -2.553
H10 0.599 1.372 -1.233
H11 0.599 1.372 1.233
H12 -1.172 1.249 -1.168
H13 -1.172 1.249 1.168
H14 -1.102 -0.864 2.485
H15 -1.102 -0.864 -2.485
H16 -0.312 0.454 3.378
H17 -0.312 0.454 -3.378

Atom - Atom Distances (Å)
  N1 O2 H3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
N11.40461.97002.49422.49421.47311.47312.76052.76052.11732.11732.08012.08012.74042.74043.43553.4355
O21.40460.98632.88272.88272.34822.34822.57972.57972.61752.61753.27763.27763.26133.26133.87173.8717
H31.97000.98633.21653.21652.55082.55082.83032.83032.38092.38093.52743.52743.85033.85034.04224.0422
C42.49422.88273.21654.98651.53123.80801.08935.16264.12652.17414.03892.16031.09135.09601.09305.9059
C52.49422.88273.21654.98653.80801.53125.16261.08932.17414.12652.16034.03895.09601.09135.90591.0930
C61.47312.34822.55081.53123.80802.43782.16574.14012.68911.10322.62531.09502.16014.09632.16984.6018
C71.47312.34822.55083.80801.53122.43784.14012.16571.10322.68911.09502.62534.09632.16014.60182.1698
H82.76052.57972.83031.08935.16262.16574.14015.10694.35472.52304.61793.06561.77005.33941.77466.1358
H92.76052.57972.83035.16261.08934.14012.16575.10692.52304.35473.06564.61795.33941.77006.13581.7746
H102.11732.61752.38094.12652.17412.68911.10324.35472.52302.46691.77592.98584.65983.07494.78892.5037
H112.11732.61752.38092.17414.12651.10322.68912.52304.35472.46692.98581.77593.07494.65982.50374.7889
H122.08013.27763.52744.03892.16032.62531.09504.61793.06561.77592.98582.33554.21992.49054.69382.5009
H132.08013.27763.52742.16034.03891.09502.62533.06564.61792.98581.77592.33552.49054.21992.50094.6938
H142.74043.26133.85031.09135.09602.16014.09631.77005.33944.65983.07494.21992.49054.96911.77736.0602
H152.74043.26133.85035.09601.09134.09632.16015.33941.77003.07494.65982.49054.21994.96916.06021.7773
H163.43553.87174.04221.09305.90592.16984.60181.77466.13584.78892.50374.69382.50091.77736.06026.7552
H173.43553.87174.04225.90591.09304.60182.16986.13581.77462.50374.78892.50094.69386.06021.77736.7552

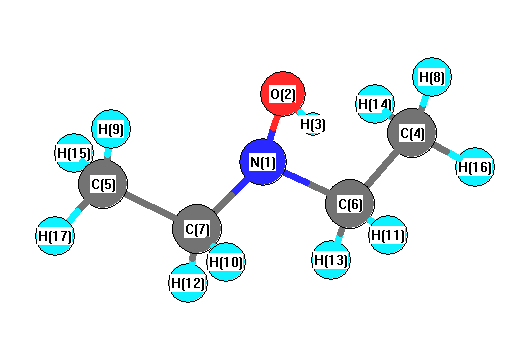
picture of Diethylhydroxylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 O2 H3 109.729 N1 C6 C4 112.221
N1 C6 H11 109.699 N1 C6 H13 107.267
N1 C7 C5 112.221 N1 C7 H10 109.699
N1 C7 H12 107.267 O2 N1 C6 109.345
O2 N1 C7 109.345 C4 C6 H11 110.167
C4 C6 H13 109.572 C5 C7 H10 110.167
C5 C7 H12 109.572 C6 N1 C7 111.669
C6 C4 H8 110.331 C6 C4 H14 109.774
C6 C4 H16 110.439 C7 C5 H9 110.331
C7 C5 H15 109.774 C7 C5 H17 110.439
H8 C4 H14 108.525 H8 C4 H16 108.812
H9 C5 H15 108.525 H9 C5 H17 108.812
H10 C7 H12 107.777 H11 C6 H13 107.777
H14 C4 H16 108.917 H15 C5 H17 108.917
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 N -0.191      
2 O -0.551      
3 H 0.388      
4 C -0.456      
5 C -0.456      
6 C -0.120      
7 C -0.120      
8 H 0.175      
9 H 0.175      
10 H 0.110      
11 H 0.110      
12 H 0.155      
13 H 0.155      
14 H 0.167      
15 H 0.167      
16 H 0.146      
17 H 0.146      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.802 -0.202 0.000
y -0.202 7.415 0.000
z 0.000 0.000 9.284


<r2> (average value of r2) Å2
<r2> 210.847
(<r2>)1/2 14.521