return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H6N2O2 ((E)-Azodioxymethane)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-338.634574
Energy at 298.15K-338.643799
HF Energy-337.646498
Nuclear repulsion energy259.916876
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3281 3079 0.00      
2 Ag 3143 2950 0.00      
3 Ag 1676 1573 0.00      
4 Ag 1524 1430 0.00      
5 Ag 1519 1426 0.00      
6 Ag 1470 1380 0.00      
7 Ag 1149 1078 0.00      
8 Ag 835 784 0.00      
9 Ag 651 611 0.00      
10 Ag 425 399 0.00      
11 Au 3225 3027 3.00      
12 Au 1522 1429 23.24      
13 Au 1171 1099 0.01      
14 Au 345 323 6.77      
15 Au 196 184 12.12      
16 Au 138 130 6.42      
17 Bg 3224 3026 0.00      
18 Bg 1520 1427 0.00      
19 Bg 1139 1069 0.00      
20 Bg 512 481 0.00      
21 Bg 176 165 0.00      
22 Bu 3281 3080 0.71      
23 Bu 3143 2950 7.04      
24 Bu 1539 1445 1.74      
25 Bu 1486 1394 40.08      
26 Bu 1441 1352 392.82      
27 Bu 1211 1137 30.25      
28 Bu 1011 949 42.65      
29 Bu 564 530 53.41      
30 Bu 317 297 30.19      

Unscaled Zero Point Vibrational Energy (zpe) 21415.9 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 20101.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.17511 0.13275 0.07771

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -0.057 0.645 0.000
N2 0.057 -0.645 0.000
O3 -1.194 1.205 0.000
O4 1.194 -1.205 0.000
C5 1.194 1.404 0.000
C6 -1.194 -1.404 0.000
H7 0.896 2.449 0.000
H8 1.773 1.138 0.886
H9 1.773 1.138 -0.886
H10 -0.896 -2.449 0.000
H11 -1.773 -1.138 0.886
H12 -1.773 -1.138 -0.886

Atom - Atom Distances (Å)
  N1 N2 O3 O4 C5 C6 H7 H8 H9 H10 H11 H12
N11.29441.26752.23331.46372.34292.04032.09212.09213.20502.62802.6280
N21.29442.23331.26752.34291.46373.20502.62802.62802.04032.09212.0921
O31.26752.23333.39312.39642.60922.43193.09693.09693.66602.57132.5713
O42.23331.26753.39312.60922.39643.66602.57132.57132.43193.09693.0969
C51.46372.34292.39642.60923.68621.08641.09121.09124.38304.00614.0061
C62.34291.46372.60922.39643.68624.38304.00614.00611.08641.09121.0912
H72.04033.20502.43193.66601.08644.38301.80861.80865.21484.55764.5576
H82.09212.62803.09692.57131.09124.00611.80861.77254.55764.21304.5707
H92.09212.62803.09692.57131.09124.00611.80861.77254.55764.57074.2130
H103.20502.04033.66602.43194.38301.08645.21484.55764.55761.80861.8086
H112.62802.09212.57133.09694.00611.09124.55764.21304.57071.80861.7725
H122.62802.09212.57133.09694.00611.09124.55764.57074.21301.80861.7725

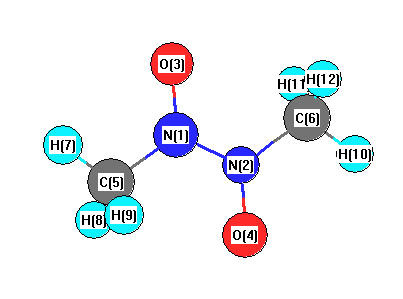
picture of (E)-Azodioxymethane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 O4 121.317 N1 N2 C6 116.180
N1 C5 H7 105.320 N1 C5 H8 109.077
N1 C5 H9 109.077 N2 N1 O3 121.317
N2 N1 C5 116.180 N2 C6 H10 105.320
N2 C6 H11 109.077 N2 C6 H12 109.077
O3 N1 C5 122.503 O4 N2 C6 122.503
H7 C5 H8 112.316 H7 C5 H9 112.316
H8 C5 H9 108.623 H10 C6 H11 112.316
H10 C6 H12 112.316 H11 C6 H12 108.623
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability