return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (benzocyclobutadiene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-307.150842
Energy at 298.15K-307.156651
HF Energy-306.335477
Nuclear repulsion energy314.238704
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3395 3136 10.09      
2 A1 3340 3085 14.29      
3 A1 3323 3070 14.45      
4 A1 1831 1691 3.40      
5 A1 1697 1568 0.72      
6 A1 1597 1475 4.46      
7 A1 1531 1414 13.86      
8 A1 1262 1166 0.00      
9 A1 1168 1079 0.27      
10 A1 1138 1051 0.22      
11 A1 1016 938 4.09      
12 A1 869 803 0.51      
13 A1 574 531 0.16      
14 A2 989 914 0.00      
15 A2 942 870 0.00      
16 A2 914 845 0.00      
17 A2 691 639 0.00      
18 A2 586 541 0.00      
19 A2 307 284 0.00      
20 B1 946 873 10.32      
21 B1 799 738 108.84      
22 B1 764 705 9.24      
23 B1 367 339 1.22      
24 B1 249 230 11.31      
25 B2 3364 3108 1.06      
26 B2 3333 3078 29.81      
27 B2 3309 3056 0.31      
28 B2 1796 1659 0.54      
29 B2 1552 1433 5.51      
30 B2 1390 1284 4.74      
31 B2 1336 1234 11.72      
32 B2 1158 1070 1.85      
33 B2 1112 1027 0.89      
34 B2 927 857 0.85      
35 B2 689 636 0.70      
36 B2 447 413 4.11      

Unscaled Zero Point Vibrational Energy (zpe) 25354.1 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 23419.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.16302 0.07413 0.05096

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.433 -0.612
C2 0.000 0.678 -1.835
C3 0.000 -0.678 -1.835
C4 0.000 -1.433 -0.612
C5 0.000 -0.667 2.031
C6 0.000 0.667 2.031
C7 0.000 0.712 0.520
C8 0.000 -0.712 0.520
H9 0.000 2.511 -0.639
H10 0.000 1.211 -2.772
H11 0.000 -1.211 -2.772
H12 0.000 -2.511 -0.639
H13 0.000 -1.428 2.791
H14 0.000 1.428 2.791

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.43662.43972.86583.37572.75221.34262.42501.07842.17093.41403.94394.44623.4033
C21.43661.35672.43974.09323.86592.35512.73452.18831.07802.10923.40625.08294.6862
C32.43971.35671.43663.86594.09322.73452.35513.40622.10921.07802.18834.68625.0829
C42.86582.43971.43662.75223.37572.42501.34263.94393.41402.17091.07843.40334.4462
C53.37574.09323.86592.75221.33342.04521.51174.15045.15704.83373.24501.07592.2285
C62.75223.86594.09323.37571.33341.51172.04523.24504.83375.15704.15042.22851.0759
C71.34262.35512.73452.42502.04521.51171.42312.14033.32963.81243.42463.12032.3814
C82.42502.73452.35511.34261.51172.04521.42313.42463.81243.32962.14032.38143.1203
H91.07842.18833.40623.94394.15043.24502.14033.42462.49774.29015.02195.22323.5967
H102.17091.07802.10923.41405.15704.83373.32963.81242.49772.42264.29016.15735.5671
H113.41402.10921.07802.17094.83375.15703.81243.32964.29012.42262.49775.56716.1573
H123.94393.40622.18831.07843.24504.15043.42462.14035.02194.29012.49773.59675.2232
H134.44625.08294.68623.40331.07592.22853.12032.38145.22326.15735.56713.59672.8565
H143.40334.68625.08294.44622.22851.07592.38143.12033.59675.56716.15735.22322.8565

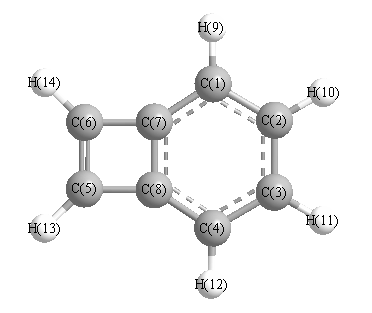
picture of benzocyclobutadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.682 C1 C2 H10 118.688
C1 C7 C6 149.203 C1 C7 N8 122.498
C2 C1 C7 115.820 C2 C1 H9 120.268
C2 C3 C4 121.682 C2 C3 H11 119.630
C3 C2 H10 119.630 C3 C4 N8 115.820
C3 C4 H12 120.268 C4 C3 H11 118.688
C4 N8 C5 149.203 C4 N8 C7 122.498
C5 C6 C7 91.701 C5 C6 H14 135.061
C5 N8 C7 88.299 C6 C5 N8 91.701
C6 C5 H13 135.061 C6 C7 N8 88.299
C7 C1 H9 123.911 C7 C6 H14 133.239
N8 C4 H12 123.911 N8 C5 H13 133.239
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability