return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (benzocyclobutadiene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-307.386276
Energy at 298.15K-307.391600
HF Energy-306.332937
Nuclear repulsion energy311.625633
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3271 3115 12.54      
2 A1 3220 3066 15.34      
3 A1 3205 3052 14.18      
4 A1 1749 1666 1.96      
5 A1 1590 1514 1.83      
6 A1 1537 1464 2.01      
7 A1 1467 1397 11.11      
8 A1 1219 1161 0.01      
9 A1 1118 1065 0.08      
10 A1 1090 1038 0.39      
11 A1 978 932 2.74      
12 A1 833 793 0.30      
13 A1 551 525 0.13      
14 A2 899 856 0.00      
15 A2 865 824 0.00      
16 A2 798 760 0.00      
17 A2 545 519 0.00      
18 A2 512 488 0.00      
19 A2 287 273 0.00      
20 B1 859 818 8.63      
21 B1 744 708 82.69      
22 B1 692 659 26.00      
23 B1 336 320 0.37      
24 B1 227 216 10.32      
25 B2 3243 3088 2.07      
26 B2 3214 3060 32.00      
27 B2 3191 3039 0.28      
28 B2 1714 1633 0.18      
29 B2 1489 1418 4.79      
30 B2 1333 1269 2.95      
31 B2 1277 1216 11.75      
32 B2 1115 1061 1.19      
33 B2 1066 1015 0.80      
34 B2 889 847 0.49      
35 B2 663 631 0.74      
36 B2 426 406 3.61      

Unscaled Zero Point Vibrational Energy (zpe) 24104.5 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 22954.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.15993 0.07307 0.05015

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.448 -0.619
C2 0.000 0.686 -1.850
C3 0.000 -0.686 -1.850
C4 0.000 -1.448 -0.619
C5 0.000 -0.678 2.047
C6 0.000 0.678 2.047
C7 0.000 0.720 0.527
C8 0.000 -0.720 0.527
H9 0.000 2.537 -0.648
H10 0.000 1.225 -2.796
H11 0.000 -1.225 -2.796
H12 0.000 -2.537 -0.648
H13 0.000 -1.449 2.812
H14 0.000 1.449 2.812

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.44772.46382.89533.40912.77421.35712.45211.08942.18923.44773.98454.48993.4309
C21.44771.37272.46384.12863.89672.37742.76222.20681.08912.13303.44015.12794.7242
C32.46381.37271.44773.89674.12862.76222.37743.44012.13301.08912.20684.72425.1279
C42.89532.46381.44772.77423.40912.45211.35713.98453.44772.18921.08943.43094.4899
C53.40914.12863.89672.77421.35592.06501.52024.19445.20374.87403.27321.08632.2602
C62.77423.89674.12863.40911.35591.52022.06503.27324.87405.20374.19442.26021.0863
C71.35712.37742.76222.45212.06501.52021.44072.16313.36173.85113.46243.15062.3984
C82.45212.76222.37741.35711.52022.06501.44073.46243.85113.36172.16312.39843.1506
H91.08942.20683.44013.98454.19443.27322.16313.46242.51764.33245.07345.27763.6269
H102.18921.08912.13303.44775.20374.87403.36173.85112.51762.45034.33246.21355.6133
H113.44772.13301.08912.18924.87405.20373.85113.36174.33242.45032.51765.61336.2135
H123.98453.44012.20681.08943.27324.19443.46242.16315.07344.33242.51763.62695.2776
H134.48995.12794.72423.43091.08632.26023.15062.39845.27766.21355.61333.62692.8972
H143.43094.72425.12794.48992.26021.08632.39843.15063.62695.61336.21355.27762.8972

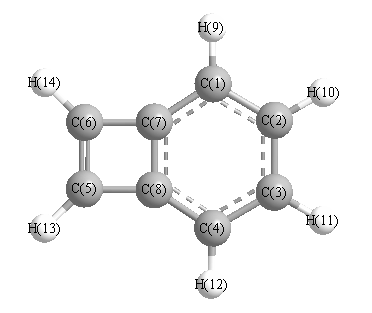
picture of benzocyclobutadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.727 C1 C2 H10 118.622
C1 C7 C6 149.189 C1 C7 N8 122.408
C2 C1 C7 115.866 C2 C1 H9 120.205
C2 C3 C4 121.727 C2 C3 H11 119.652
C3 C2 H10 119.652 C3 C4 N8 115.866
C3 C4 H12 120.205 C4 C3 H11 118.622
C4 N8 C5 149.189 C4 N8 C7 122.408
C5 C6 C7 91.597 C5 C6 H14 135.186
C5 N8 C7 88.403 C6 C5 N8 91.597
C6 C5 H13 135.186 C6 C7 N8 88.403
C7 C1 H9 123.929 C7 C6 H14 133.217
N8 C4 H12 123.929 N8 C5 H13 133.217
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability