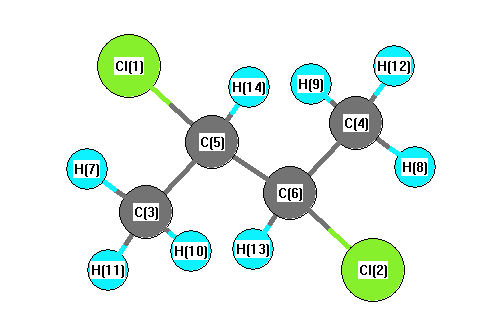
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3184 |
3028 |
0.00 |
|
|
|
| 2 |
Ag |
3176 |
3020 |
0.00 |
|
|
|
| 3 |
Ag |
3126 |
2973 |
0.00 |
|
|
|
| 4 |
Ag |
3091 |
2939 |
0.00 |
|
|
|
| 5 |
Ag |
1519 |
1444 |
0.00 |
|
|
|
| 6 |
Ag |
1518 |
1443 |
0.00 |
|
|
|
| 7 |
Ag |
1440 |
1370 |
0.00 |
|
|
|
| 8 |
Ag |
1412 |
1343 |
0.00 |
|
|
|
| 9 |
Ag |
1307 |
1243 |
0.00 |
|
|
|
| 10 |
Ag |
1198 |
1140 |
0.00 |
|
|
|
| 11 |
Ag |
1162 |
1105 |
0.00 |
|
|
|
| 12 |
Ag |
1048 |
997 |
0.00 |
|
|
|
| 13 |
Ag |
869 |
826 |
0.00 |
|
|
|
| 14 |
Ag |
719 |
683 |
0.00 |
|
|
|
| 15 |
Ag |
483 |
460 |
0.00 |
|
|
|
| 16 |
Ag |
351 |
334 |
0.00 |
|
|
|
| 17 |
Ag |
289 |
275 |
0.00 |
|
|
|
| 18 |
Ag |
237 |
226 |
0.00 |
|
|
|
| 19 |
Au |
3185 |
3029 |
14.85 |
|
|
|
| 20 |
Au |
3177 |
3021 |
25.98 |
|
|
|
| 21 |
Au |
3141 |
2987 |
8.00 |
|
|
|
| 22 |
Au |
3090 |
2938 |
16.81 |
|
|
|
| 23 |
Au |
1521 |
1446 |
20.16 |
|
|
|
| 24 |
Au |
1513 |
1439 |
10.36 |
|
|
|
| 25 |
Au |
1439 |
1369 |
25.70 |
|
|
|
| 26 |
Au |
1338 |
1272 |
5.03 |
|
|
|
| 27 |
Au |
1251 |
1190 |
39.39 |
|
|
|
| 28 |
Au |
1114 |
1060 |
9.73 |
|
|
|
| 29 |
Au |
1040 |
989 |
36.93 |
|
|
|
| 30 |
Au |
992 |
943 |
15.31 |
|
|
|
| 31 |
Au |
675 |
642 |
82.58 |
|
|
|
| 32 |
Au |
363 |
345 |
3.33 |
|
|
|
| 33 |
Au |
338 |
322 |
2.74 |
|
|
|
| 34 |
Au |
253 |
241 |
3.61 |
|
|
|
| 35 |
Au |
207 |
197 |
3.02 |
|
|
|
| 36 |
Au |
69 |
66 |
3.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25416.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 24171.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Cl |
-0.067 |
|
|
|
| 2 |
Cl |
-0.067 |
|
|
|
| 3 |
C |
-0.509 |
|
|
|
| 4 |
C |
-0.509 |
|
|
|
| 5 |
C |
-0.252 |
|
|
|
| 6 |
C |
-0.252 |
|
|
|
| 7 |
H |
0.199 |
|
|
|
| 8 |
H |
0.199 |
|
|
|
| 9 |
H |
0.210 |
|
|
|
| 10 |
H |
0.210 |
|
|
|
| 11 |
H |
0.191 |
|
|
|
| 12 |
H |
0.191 |
|
|
|
| 13 |
H |
0.229 |
|
|
|
| 14 |
H |
0.229 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.027 |
2.037 |
-3.685 |
| y |
2.037 |
-52.807 |
3.557 |
| z |
-3.685 |
3.557 |
-54.409 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.581 |
2.037 |
-3.685 |
| y |
2.037 |
-0.589 |
3.557 |
| z |
-3.685 |
3.557 |
-2.992 |
|
| Polar |
|---|
| 3z2-r2 | -5.984 |
|---|
| x2-y2 | 2.780 |
|---|
| xy | 2.037 |
|---|
| xz | -3.685 |
|---|
| yz | 3.557 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.075 |
-0.446 |
0.657 |
| y |
-0.446 |
8.434 |
-2.108 |
| z |
0.657 |
-2.108 |
10.172 |
<r2> (average value of r
2) Å
2
| <r2> |
290.244 |
| (<r2>)1/2 |
17.037 |