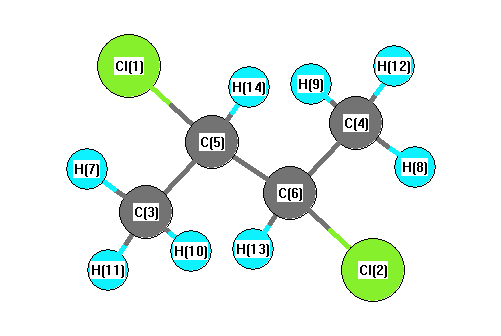
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3094 |
3043 |
0.00 |
|
|
|
| 2 |
Ag |
3086 |
3035 |
0.00 |
|
|
|
| 3 |
Ag |
3030 |
2980 |
0.00 |
|
|
|
| 4 |
Ag |
3005 |
2956 |
0.00 |
|
|
|
| 5 |
Ag |
1473 |
1448 |
0.00 |
|
|
|
| 6 |
Ag |
1472 |
1448 |
0.00 |
|
|
|
| 7 |
Ag |
1387 |
1364 |
0.00 |
|
|
|
| 8 |
Ag |
1359 |
1336 |
0.00 |
|
|
|
| 9 |
Ag |
1252 |
1232 |
0.00 |
|
|
|
| 10 |
Ag |
1152 |
1133 |
0.00 |
|
|
|
| 11 |
Ag |
1121 |
1102 |
0.00 |
|
|
|
| 12 |
Ag |
1008 |
991 |
0.00 |
|
|
|
| 13 |
Ag |
831 |
818 |
0.00 |
|
|
|
| 14 |
Ag |
676 |
665 |
0.00 |
|
|
|
| 15 |
Ag |
468 |
460 |
0.00 |
|
|
|
| 16 |
Ag |
335 |
330 |
0.00 |
|
|
|
| 17 |
Ag |
276 |
272 |
0.00 |
|
|
|
| 18 |
Ag |
233 |
230 |
0.00 |
|
|
|
| 19 |
Au |
3095 |
3044 |
16.74 |
|
|
|
| 20 |
Au |
3087 |
3036 |
24.10 |
|
|
|
| 21 |
Au |
3044 |
2994 |
10.01 |
|
|
|
| 22 |
Au |
3004 |
2955 |
16.59 |
|
|
|
| 23 |
Au |
1474 |
1450 |
17.61 |
|
|
|
| 24 |
Au |
1468 |
1444 |
10.99 |
|
|
|
| 25 |
Au |
1387 |
1364 |
25.17 |
|
|
|
| 26 |
Au |
1291 |
1270 |
5.04 |
|
|
|
| 27 |
Au |
1198 |
1178 |
38.92 |
|
|
|
| 28 |
Au |
1076 |
1058 |
10.40 |
|
|
|
| 29 |
Au |
1001 |
984 |
33.78 |
|
|
|
| 30 |
Au |
959 |
943 |
17.34 |
|
|
|
| 31 |
Au |
630 |
619 |
84.94 |
|
|
|
| 32 |
Au |
349 |
343 |
3.46 |
|
|
|
| 33 |
Au |
325 |
320 |
2.99 |
|
|
|
| 34 |
Au |
246 |
242 |
3.56 |
|
|
|
| 35 |
Au |
201 |
198 |
3.63 |
|
|
|
| 36 |
Au |
66 |
65 |
3.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24579.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 24174.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Cl |
-0.070 |
|
|
|
| 2 |
Cl |
-0.070 |
|
|
|
| 3 |
C |
-0.477 |
|
|
|
| 4 |
C |
-0.477 |
|
|
|
| 5 |
C |
-0.228 |
|
|
|
| 6 |
C |
-0.228 |
|
|
|
| 7 |
H |
0.186 |
|
|
|
| 8 |
H |
0.186 |
|
|
|
| 9 |
H |
0.197 |
|
|
|
| 10 |
H |
0.197 |
|
|
|
| 11 |
H |
0.180 |
|
|
|
| 12 |
H |
0.180 |
|
|
|
| 13 |
H |
0.212 |
|
|
|
| 14 |
H |
0.212 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.968 |
2.040 |
-3.677 |
| y |
2.040 |
-52.802 |
3.479 |
| z |
-3.677 |
3.479 |
-54.433 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.650 |
2.040 |
-3.677 |
| y |
2.040 |
-0.602 |
3.479 |
| z |
-3.677 |
3.479 |
-3.048 |
|
| Polar |
|---|
| 3z2-r2 | -6.097 |
|---|
| x2-y2 | 2.834 |
|---|
| xy | 2.040 |
|---|
| xz | -3.677 |
|---|
| yz | 3.479 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.516 |
-0.488 |
0.707 |
| y |
-0.488 |
8.742 |
-2.284 |
| z |
0.707 |
-2.284 |
10.537 |
<r2> (average value of r
2) Å
2
| <r2> |
295.013 |
| (<r2>)1/2 |
17.176 |