Jump to
S1C2
Vibrational Frequencies calculated at B3PW91/6-31G*
Geometric Data calculated at B3PW91/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -235.767792 |
| Energy at 298.15K | -235.779966 |
| HF Energy | -235.767792 |
| Nuclear repulsion energy | 226.222380 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3148 |
3012 |
57.72 |
|
|
|
| 2 |
A |
3134 |
2999 |
24.23 |
|
|
|
| 3 |
A |
3130 |
2994 |
36.28 |
|
|
|
| 4 |
A |
3129 |
2993 |
3.71 |
|
|
|
| 5 |
A |
3125 |
2989 |
51.69 |
|
|
|
| 6 |
A |
3093 |
2960 |
25.10 |
|
|
|
| 7 |
A |
3090 |
2956 |
21.17 |
|
|
|
| 8 |
A |
3068 |
2935 |
9.09 |
|
|
|
| 9 |
A |
3051 |
2918 |
37.10 |
|
|
|
| 10 |
A |
3046 |
2914 |
27.91 |
|
|
|
| 11 |
A |
3039 |
2907 |
39.14 |
|
|
|
| 12 |
A |
3018 |
2888 |
27.57 |
|
|
|
| 13 |
A |
1768 |
1691 |
0.42 |
|
|
|
| 14 |
A |
1533 |
1467 |
5.32 |
|
|
|
| 15 |
A |
1524 |
1458 |
6.39 |
|
|
|
| 16 |
A |
1518 |
1452 |
5.85 |
|
|
|
| 17 |
A |
1517 |
1452 |
4.23 |
|
|
|
| 18 |
A |
1504 |
1439 |
6.01 |
|
|
|
| 19 |
A |
1502 |
1437 |
2.35 |
|
|
|
| 20 |
A |
1436 |
1374 |
1.13 |
|
|
|
| 21 |
A |
1435 |
1373 |
2.74 |
|
|
|
| 22 |
A |
1404 |
1343 |
0.98 |
|
|
|
| 23 |
A |
1357 |
1299 |
0.90 |
|
|
|
| 24 |
A |
1348 |
1289 |
0.43 |
|
|
|
| 25 |
A |
1330 |
1272 |
0.05 |
|
|
|
| 26 |
A |
1301 |
1244 |
7.17 |
|
|
|
| 27 |
A |
1264 |
1209 |
0.48 |
|
|
|
| 28 |
A |
1190 |
1138 |
0.03 |
|
|
|
| 29 |
A |
1125 |
1076 |
1.18 |
|
|
|
| 30 |
A |
1108 |
1060 |
1.75 |
|
|
|
| 31 |
A |
1074 |
1027 |
0.18 |
|
|
|
| 32 |
A |
1069 |
1022 |
4.47 |
|
|
|
| 33 |
A |
1048 |
1003 |
6.10 |
|
|
|
| 34 |
A |
1012 |
968 |
28.78 |
|
|
|
| 35 |
A |
937 |
896 |
5.65 |
|
|
|
| 36 |
A |
911 |
871 |
2.76 |
|
|
|
| 37 |
A |
880 |
842 |
0.42 |
|
|
|
| 38 |
A |
781 |
747 |
0.76 |
|
|
|
| 39 |
A |
748 |
716 |
3.03 |
|
|
|
| 40 |
A |
531 |
508 |
0.17 |
|
|
|
| 41 |
A |
388 |
371 |
1.21 |
|
|
|
| 42 |
A |
310 |
297 |
0.78 |
|
|
|
| 43 |
A |
284 |
271 |
1.44 |
|
|
|
| 44 |
A |
247 |
236 |
0.05 |
|
|
|
| 45 |
A |
209 |
200 |
0.75 |
|
|
|
| 46 |
A |
142 |
136 |
0.29 |
|
|
|
| 47 |
A |
98 |
93 |
0.05 |
|
|
|
| 48 |
A |
77 |
74 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36487.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 34907.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
3.185 |
0.214 |
-0.159 |
| C2 |
1.806 |
-0.367 |
-0.216 |
| C3 |
0.741 |
0.108 |
0.434 |
| C4 |
-0.643 |
-0.468 |
0.373 |
| C5 |
-1.685 |
0.519 |
-0.172 |
| C6 |
-3.098 |
-0.057 |
-0.190 |
| H7 |
3.223 |
1.099 |
0.486 |
| H8 |
3.910 |
-0.516 |
0.226 |
| H9 |
3.537 |
0.508 |
-1.157 |
| H10 |
1.683 |
-1.253 |
-0.844 |
| H11 |
0.862 |
0.996 |
1.061 |
| H12 |
-0.955 |
-0.783 |
1.382 |
| H13 |
-0.640 |
-1.377 |
-0.245 |
| H14 |
-1.391 |
0.823 |
-1.185 |
| H15 |
-1.668 |
1.434 |
0.437 |
| H16 |
-3.821 |
0.667 |
-0.584 |
| H17 |
-3.153 |
-0.955 |
-0.817 |
| H18 |
-3.425 |
-0.339 |
0.818 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4979 | 2.5177 | 3.9249 | 4.8796 | 6.2896 | 1.0954 | 1.0985 | 1.0985 | 2.2086 | 2.7381 | 4.5291 | 4.1435 | 4.7294 | 5.0397 | 7.0334 | 6.4785 | 6.7054 |
C2 | 1.4979 | | 1.3354 | 2.5209 | 3.6016 | 4.9141 | 2.1563 | 2.1556 | 2.1557 | 1.0925 | 2.0926 | 3.2175 | 2.6460 | 3.5460 | 3.9672 | 5.7325 | 5.0294 | 5.3327 | C3 | 2.5177 | 1.3354 | | 1.5002 | 2.5334 | 3.8929 | 2.6732 | 3.2373 | 3.2422 | 2.0910 | 1.0933 | 2.1375 | 2.1380 | 2.7707 | 2.7496 | 4.7068 | 4.2255 | 4.2076 | C4 | 3.9249 | 2.5209 | 1.5002 | | 1.5356 | 2.5526 | 4.1731 | 4.5559 | 4.5573 | 2.7397 | 2.2092 | 1.1017 | 1.0987 | 2.1578 | 2.1623 | 3.5076 | 2.8201 | 2.8208 | C5 | 4.8796 | 3.6016 | 2.5334 | 1.5356 | | 1.5267 | 4.9854 | 5.7039 | 5.3142 | 3.8644 | 2.8687 | 2.1542 | 2.1662 | 1.0980 | 1.0992 | 2.1803 | 2.1786 | 2.1787 | C6 | 6.2896 | 4.9141 | 3.8929 | 2.5526 | 1.5267 | | 6.4616 | 7.0360 | 6.7295 | 4.9717 | 4.2842 | 2.7550 | 2.7910 | 2.1633 | 2.1595 | 1.0959 | 1.0970 | 1.0970 | H7 | 1.0954 | 2.1563 | 2.6732 | 4.1731 | 4.9854 | 6.4616 | | 1.7742 | 1.7743 | 3.1096 | 2.4324 | 4.6691 | 4.6457 | 4.9151 | 4.9028 | 7.1375 | 6.8241 | 6.8102 | H8 | 1.0985 | 2.1556 | 3.2373 | 4.5559 | 5.7039 | 7.0360 | 1.7742 | | 1.7606 | 2.5783 | 3.5039 | 5.0083 | 4.6545 | 5.6470 | 5.9133 | 7.8628 | 7.1531 | 7.3618 | H9 | 1.0985 | 2.1557 | 3.2422 | 4.5573 | 5.3142 | 6.7295 | 1.7743 | 1.7606 | | 2.5760 | 3.5093 | 5.3194 | 4.6722 | 4.9385 | 5.5222 | 7.3819 | 6.8565 | 7.2869 | H10 | 2.2086 | 1.0925 | 2.0910 | 2.7397 | 3.8644 | 4.9717 | 3.1096 | 2.5783 | 2.5760 | | 3.0590 | 3.4837 | 2.4017 | 3.7249 | 4.4822 | 5.8346 | 4.8449 | 5.4492 | H11 | 2.7381 | 2.0926 | 1.0933 | 2.2092 | 2.8687 | 4.2842 | 2.4324 | 3.5039 | 3.5093 | 3.0590 | | 2.5628 | 3.0964 | 3.1856 | 2.6420 | 4.9736 | 4.8424 | 4.4966 | H12 | 4.5291 | 3.2175 | 2.1375 | 1.1017 | 2.1542 | 2.7550 | 4.6691 | 5.0083 | 5.3194 | 3.4837 | 2.5628 | | 1.7606 | 3.0591 | 2.5130 | 3.7650 | 3.1136 | 2.5721 | H13 | 4.1435 | 2.6460 | 2.1380 | 1.0987 | 2.1662 | 2.7910 | 4.6457 | 4.6545 | 4.6722 | 2.4017 | 3.0964 | 1.7606 | | 2.5074 | 3.0699 | 3.7960 | 2.6115 | 3.1573 | H14 | 4.7294 | 3.5460 | 2.7707 | 2.1578 | 1.0980 | 2.1633 | 4.9151 | 5.6470 | 4.9385 | 3.7249 | 3.1856 | 3.0591 | 2.5074 | | 1.7556 | 2.5077 | 2.5302 | 3.0827 | H15 | 5.0397 | 3.9672 | 2.7496 | 2.1623 | 1.0992 | 2.1595 | 4.9028 | 5.9133 | 5.5222 | 4.4822 | 2.6420 | 2.5130 | 3.0699 | 1.7556 | | 2.5030 | 3.0804 | 2.5255 | H16 | 7.0334 | 5.7325 | 4.7068 | 3.5076 | 2.1803 | 1.0959 | 7.1375 | 7.8628 | 7.3819 | 5.8346 | 4.9736 | 3.7650 | 3.7960 | 2.5077 | 2.5030 | | 1.7700 | 1.7704 | H17 | 6.4785 | 5.0294 | 4.2255 | 2.8201 | 2.1786 | 1.0970 | 6.8241 | 7.1531 | 6.8565 | 4.8449 | 4.8424 | 3.1136 | 2.6115 | 2.5302 | 3.0804 | 1.7700 | | 1.7691 | H18 | 6.7054 | 5.3327 | 4.2076 | 2.8208 | 2.1787 | 1.0970 | 6.8102 | 7.3618 | 7.2869 | 5.4492 | 4.4966 | 2.5721 | 3.1573 | 3.0827 | 2.5255 | 1.7704 | 1.7691 | |
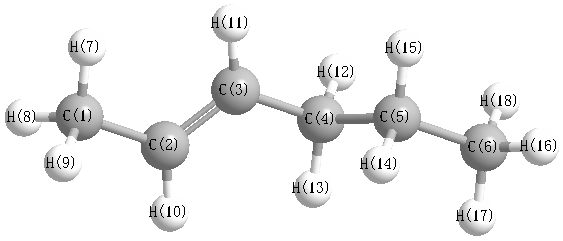 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.304 |
|
C1 |
C2 |
H10 |
116.119 |
| C2 |
C1 |
H7 |
111.567 |
|
C2 |
C1 |
H8 |
111.318 |
| C2 |
C1 |
H9 |
111.328 |
|
C2 |
C3 |
C4 |
125.402 |
| C2 |
C3 |
H11 |
118.663 |
|
C3 |
C2 |
H10 |
118.577 |
| C3 |
C4 |
C5 |
113.129 |
|
C3 |
C4 |
H12 |
109.521 |
| C3 |
C4 |
H13 |
109.736 |
|
C4 |
C3 |
H11 |
115.933 |
| C4 |
C5 |
C6 |
112.931 |
|
C4 |
C5 |
H14 |
108.905 |
| C4 |
C5 |
H15 |
109.185 |
|
C5 |
C4 |
H12 |
108.417 |
| C5 |
C4 |
H13 |
109.515 |
|
C5 |
C6 |
H16 |
111.419 |
| C5 |
C6 |
H17 |
111.219 |
|
C5 |
C6 |
H18 |
111.217 |
| C6 |
C5 |
H14 |
109.934 |
|
C6 |
C5 |
H15 |
109.576 |
| H7 |
C1 |
H8 |
107.935 |
|
H7 |
C1 |
H9 |
107.952 |
| H8 |
C1 |
H9 |
106.517 |
|
H12 |
C4 |
H13 |
106.287 |
| H14 |
C5 |
H15 |
106.075 |
|
H16 |
C6 |
H17 |
107.638 |
| H16 |
C6 |
H18 |
107.673 |
|
H17 |
C6 |
H18 |
107.483 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.555 |
|
|
|
| 2 |
C |
-0.125 |
|
|
|
| 3 |
C |
-0.113 |
|
|
|
| 4 |
C |
-0.352 |
|
|
|
| 5 |
C |
-0.305 |
|
|
|
| 6 |
C |
-0.509 |
|
|
|
| 7 |
H |
0.174 |
|
|
|
| 8 |
H |
0.176 |
|
|
|
| 9 |
H |
0.176 |
|
|
|
| 10 |
H |
0.145 |
|
|
|
| 11 |
H |
0.142 |
|
|
|
| 12 |
H |
0.163 |
|
|
|
| 13 |
H |
0.161 |
|
|
|
| 14 |
H |
0.166 |
|
|
|
| 15 |
H |
0.158 |
|
|
|
| 16 |
H |
0.168 |
|
|
|
| 17 |
H |
0.165 |
|
|
|
| 18 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.022 |
0.036 |
-0.045 |
0.062 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.291 |
-0.026 |
-0.653 |
| y |
-0.026 |
-39.165 |
1.211 |
| z |
-0.653 |
1.211 |
-39.397 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.990 |
-0.026 |
-0.653 |
| y |
-0.026 |
-0.322 |
1.211 |
| z |
-0.653 |
1.211 |
-0.669 |
|
| Polar |
|---|
| 3z2-r2 | -1.338 |
|---|
| x2-y2 | 0.875 |
|---|
| xy | -0.026 |
|---|
| xz | -0.653 |
|---|
| yz | 1.211 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.199 |
-0.225 |
-0.947 |
| y |
-0.225 |
8.207 |
0.708 |
| z |
-0.947 |
0.708 |
7.960 |
<r2> (average value of r
2) Å
2
| <r2> |
295.860 |
| (<r2>)1/2 |
17.201 |
