return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H4N4 (1H-Tetrazole, 5-methyl-)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-296.717329
Energy at 298.15K-296.724801
HF Energy-295.795490
Nuclear repulsion energy228.875965
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3723 3494 102.07      
2 A 3227 3029 0.94      
3 A 3118 2927 8.26      
4 A 1665 1563 49.85      
5 A 1568 1471 0.65      
6 A 1501 1409 31.98      
7 A 1479 1388 2.20      
8 A 1422 1335 1.32      
9 A 1344 1262 23.14      
10 A 1153 1083 3.90      
11 A 1148 1078 13.44      
12 A 1111 1042 20.37      
13 A 1042 978 0.36      
14 A 1031 968 2.18      
15 A 705 662 3.98      
16 A 345 324 4.47      
17 A 3188 2992 6.86      
18 A 1542 1447 11.21      
19 A 1105 1037 1.49      
20 A 743 698 9.37      
21 A 712 668 19.42      
22 A 574 539 86.34      
23 A 274 257 0.07      
24 A 84 79 0.77      

Unscaled Zero Point Vibrational Energy (zpe) 16901.5 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 15863.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.32822 0.12212 0.09052

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -1.145 -0.052 0.000
C2 0.000 0.597 0.000
C3 0.188 2.075 0.000
H4 0.736 2.404 0.888
H5 0.736 2.404 -0.888
H6 -0.797 2.544 0.000
N7 0.988 -0.322 0.000
N8 0.447 -1.556 0.000
N9 -0.832 -1.381 0.000
H10 1.991 -0.212 0.000

Atom - Atom Distances (Å)
  N1 C2 C3 H4 H5 H6 N7 N8 N9 H10
N11.31602.51043.21853.21852.61922.14952.18951.36473.1398
C21.31601.49022.14382.14382.10361.34912.19872.14562.1490
C32.51041.49021.09391.09391.09052.52723.64043.60332.9125
H43.21852.14381.09391.77561.77682.87814.06874.19193.0344
H53.21852.14381.09391.77561.77682.87814.06874.19193.0344
H62.61922.10361.09051.77681.77683.37604.28423.92463.9200
N72.14951.34912.52722.87812.87813.37601.34732.10551.0093
N82.18952.19873.64044.06874.06874.28421.34731.29102.0471
N91.36472.14563.60334.19194.19193.92462.10551.29103.0556
H103.13982.14902.91253.03443.03443.92001.00932.04713.0556

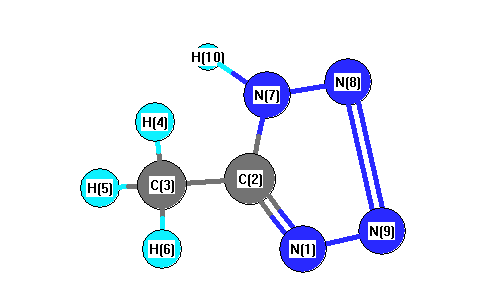
picture of 1H-Tetrazole, 5-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 126.797 N1 C2 N7 107.510
N1 N9 N8 111.040 C2 N1 N9 106.318
C2 C3 H4 111.188 C2 C3 H5 111.188
C2 C3 H6 108.204 C2 N7 N8 109.261
C2 N7 H10 130.808 C3 C2 N7 125.692
H4 C3 H5 108.498 H4 C3 H6 108.855
H5 C3 H6 108.855 N7 N8 N9 105.870
N8 N7 H10 119.931
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability