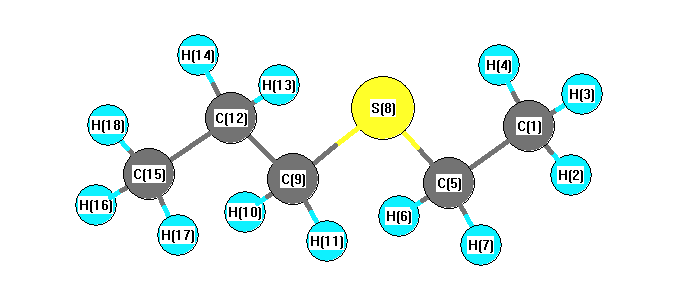
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3140 |
3004 |
23.60 |
|
|
|
| 2 |
A' |
3135 |
2999 |
33.54 |
|
|
|
| 3 |
A' |
3064 |
2931 |
25.79 |
|
|
|
| 4 |
A' |
3061 |
2929 |
35.81 |
|
|
|
| 5 |
A' |
3057 |
2924 |
32.06 |
|
|
|
| 6 |
A' |
3054 |
2922 |
27.59 |
|
|
|
| 7 |
A' |
3047 |
2915 |
14.96 |
|
|
|
| 8 |
A' |
1533 |
1466 |
4.54 |
|
|
|
| 9 |
A' |
1530 |
1463 |
2.56 |
|
|
|
| 10 |
A' |
1518 |
1452 |
0.79 |
|
|
|
| 11 |
A' |
1515 |
1450 |
1.15 |
|
|
|
| 12 |
A' |
1506 |
1441 |
10.59 |
|
|
|
| 13 |
A' |
1436 |
1374 |
1.88 |
|
|
|
| 14 |
A' |
1433 |
1371 |
3.76 |
|
|
|
| 15 |
A' |
1393 |
1332 |
1.96 |
|
|
|
| 16 |
A' |
1324 |
1266 |
20.42 |
|
|
|
| 17 |
A' |
1272 |
1217 |
32.58 |
|
|
|
| 18 |
A' |
1133 |
1084 |
3.27 |
|
|
|
| 19 |
A' |
1081 |
1034 |
6.67 |
|
|
|
| 20 |
A' |
1068 |
1022 |
0.17 |
|
|
|
| 21 |
A' |
1010 |
966 |
4.91 |
|
|
|
| 22 |
A' |
920 |
881 |
2.75 |
|
|
|
| 23 |
A' |
779 |
745 |
1.58 |
|
|
|
| 24 |
A' |
704 |
673 |
0.65 |
|
|
|
| 25 |
A' |
383 |
367 |
0.39 |
|
|
|
| 26 |
A' |
301 |
288 |
0.71 |
|
|
|
| 27 |
A' |
272 |
260 |
0.92 |
|
|
|
| 28 |
A' |
105 |
100 |
0.32 |
|
|
|
| 29 |
A" |
3149 |
3013 |
25.42 |
|
|
|
| 30 |
A" |
3131 |
2996 |
57.12 |
|
|
|
| 31 |
A" |
3108 |
2973 |
3.60 |
|
|
|
| 32 |
A" |
3102 |
2968 |
18.69 |
|
|
|
| 33 |
A" |
3087 |
2953 |
2.16 |
|
|
|
| 34 |
A" |
1526 |
1460 |
7.90 |
|
|
|
| 35 |
A" |
1518 |
1452 |
8.62 |
|
|
|
| 36 |
A" |
1336 |
1278 |
0.19 |
|
|
|
| 37 |
A" |
1280 |
1224 |
0.03 |
|
|
|
| 38 |
A" |
1254 |
1200 |
0.00 |
|
|
|
| 39 |
A" |
1083 |
1036 |
1.37 |
|
|
|
| 40 |
A" |
1055 |
1009 |
0.18 |
|
|
|
| 41 |
A" |
880 |
842 |
0.08 |
|
|
|
| 42 |
A" |
800 |
765 |
6.21 |
|
|
|
| 43 |
A" |
755 |
723 |
4.04 |
|
|
|
| 44 |
A" |
250 |
239 |
0.09 |
|
|
|
| 45 |
A" |
244 |
234 |
0.03 |
|
|
|
| 46 |
A" |
118 |
113 |
1.02 |
|
|
|
| 47 |
A" |
66 |
63 |
0.46 |
|
|
|
| 48 |
A" |
43 |
41 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35777.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 34227.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.522 |
|
|
-0.264 |
| 2 |
H |
0.176 |
|
|
0.061 |
| 3 |
H |
0.188 |
|
|
0.102 |
| 4 |
H |
0.188 |
|
|
0.102 |
| 5 |
C |
-0.458 |
|
|
0.076 |
| 6 |
H |
0.189 |
|
|
0.053 |
| 7 |
H |
0.189 |
|
|
0.053 |
| 8 |
S |
0.100 |
|
|
-0.323 |
| 9 |
C |
-0.459 |
|
|
-0.253 |
| 10 |
H |
0.186 |
|
|
0.118 |
| 11 |
H |
0.186 |
|
|
0.118 |
| 12 |
C |
-0.321 |
|
|
0.256 |
| 13 |
H |
0.180 |
|
|
0.012 |
| 14 |
H |
0.180 |
|
|
0.012 |
| 15 |
C |
-0.511 |
|
|
-0.464 |
| 16 |
H |
0.168 |
|
|
0.114 |
| 17 |
H |
0.168 |
|
|
0.114 |
| 18 |
H |
0.174 |
|
|
0.110 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.000 |
-1.342 |
0.000 |
1.674 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.002 |
-1.325 |
0.000 |
1.661 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.105 |
2.548 |
0.000 |
| y |
2.548 |
-46.729 |
0.000 |
| z |
0.000 |
0.000 |
-47.438 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.978 |
2.548 |
0.000 |
| y |
2.548 |
-1.458 |
0.000 |
| z |
0.000 |
0.000 |
-2.520 |
|
| Polar |
|---|
| 3z2-r2 | -5.040 |
|---|
| x2-y2 | 3.624 |
|---|
| xy | 2.548 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
354.321 |
| (<r2>)1/2 |
18.823 |