return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (Ethyl propyl sulfide)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-595.519972
Energy at 298.15K-595.532715
HF Energy-595.261897
Nuclear repulsion energy298.893333
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3161 3001 24.58      
2 A' 3156 2996 34.61      
3 A' 3088 2931 24.31      
4 A' 3085 2928 28.24      
5 A' 3083 2927 35.38      
6 A' 3078 2922 27.74      
7 A' 3071 2915 13.58      
8 A' 1559 1480 3.23      
9 A' 1555 1476 1.57      
10 A' 1544 1466 0.63      
11 A' 1542 1463 1.47      
12 A' 1533 1455 11.19      
13 A' 1460 1386 1.08      
14 A' 1459 1385 2.53      
15 A' 1416 1344 1.66      
16 A' 1348 1280 21.74      
17 A' 1298 1232 35.37      
18 A' 1146 1087 2.93      
19 A' 1093 1038 5.69      
20 A' 1072 1018 0.13      
21 A' 1017 966 3.45      
22 A' 928 880 2.04      
23 A' 785 745 1.63      
24 A' 707 671 0.86      
25 A' 390 370 0.34      
26 A' 305 289 0.70      
27 A' 277 263 0.83      
28 A' 108 103 0.29      
29 A" 3171 3010 26.74      
30 A" 3153 2993 58.98      
31 A" 3131 2972 0.02      
32 A" 3128 2969 22.40      
33 A" 3111 2953 1.75      
34 A" 1551 1472 6.62      
35 A" 1543 1465 7.12      
36 A" 1354 1285 0.20      
37 A" 1300 1234 0.00      
38 A" 1278 1213 0.02      
39 A" 1103 1047 1.20      
40 A" 1075 1020 0.08      
41 A" 895 849 0.09      
42 A" 812 771 4.71      
43 A" 768 729 3.04      
44 A" 254 241 0.08      
45 A" 249 236 0.04      
46 A" 120 114 0.97      
47 A" 68 65 0.42      
48 A" 46 43 0.27      

Unscaled Zero Point Vibrational Energy (zpe) 36186.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 34348.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.38224 0.03390 0.03207

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.718 2.548 0.000
H2 2.787 2.767 0.000
H3 1.262 3.000 0.854
H4 1.262 3.000 -0.854
C5 1.491 1.031 0.000
H6 1.963 0.594 -0.902
H7 1.963 0.594 0.902
S8 0.000 0.710 0.000
C9 -0.104 -0.806 0.000
H10 0.386 -1.231 -0.879
H11 0.386 -1.231 0.879
C12 -1.575 -1.260 0.000
H13 -2.068 -0.855 0.894
H14 -2.068 -0.855 -0.894
C15 -1.667 -2.788 0.000
H16 -1.170 -3.198 -0.853
H17 -1.170 -3.198 0.853
H18 -2.713 -3.097 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 S8 C9 H10 H11 C12 H13 H14 C15 H16 H17 H18
C11.09131.06851.06851.53342.16592.16592.51593.81704.10204.10205.03395.16815.16816.31866.48676.48677.1761
H21.09131.76371.76372.16602.49252.49253.46404.59614.74564.74565.93626.12256.12257.11977.20827.20828.0395
H31.06851.76371.70752.15803.06002.50662.75064.13304.65514.32085.18845.09365.38516.54236.87296.65757.3279
H41.06851.76371.70752.15802.50663.06002.75064.13304.32084.65515.18845.38515.09366.54236.65756.87297.3279
C51.53342.16602.15802.15801.10821.10821.52532.43322.66682.66683.82734.12574.12574.95555.06875.06875.8920
H62.16592.49253.06002.50661.10821.80412.16382.65492.41262.99854.09514.64504.28365.04284.91895.22256.0255
H72.16592.49252.50663.06001.10821.80412.16382.65492.99852.41264.09514.28364.64505.04285.22254.91896.0255
S82.51593.46402.75062.75061.52532.16382.16381.51972.16502.16502.52172.74282.74283.87444.16714.16714.6745
C93.81704.59614.13304.13302.43322.65492.65491.51971.09181.09181.53922.15842.15842.52372.75372.75373.4719
H104.10204.74564.65514.32082.66682.41262.99852.16501.09181.75722.14853.05022.48212.72192.50753.04733.7223
H114.10204.74564.32084.65512.66682.99852.41262.16501.09181.75722.14852.48213.05022.72193.04732.50753.7223
C125.03395.93625.18845.18843.82734.09514.09512.52171.53922.14852.14851.09861.09861.53062.15602.15602.1611
H135.16816.12255.09365.38514.12574.64504.28362.74282.15843.05022.48211.09861.78872.16703.05792.50972.4986
H145.16816.12255.38515.09364.12574.28364.64502.74282.15842.48213.05021.09861.78872.16702.50973.05792.4986
C156.31867.11976.54236.54234.95555.04285.04283.87442.52372.72192.72191.53062.16702.16701.06951.06951.0908
H166.48677.20826.87296.65755.06874.91895.22254.16712.75372.50753.04732.15603.05792.50971.06951.70631.7666
H176.48677.20826.65756.87295.06875.22254.91894.16712.75373.04732.50752.15602.50973.05791.06951.70631.7666
H187.17618.03957.32797.32795.89206.02556.02554.67453.47193.72233.72232.16112.49862.49861.09081.76661.7666

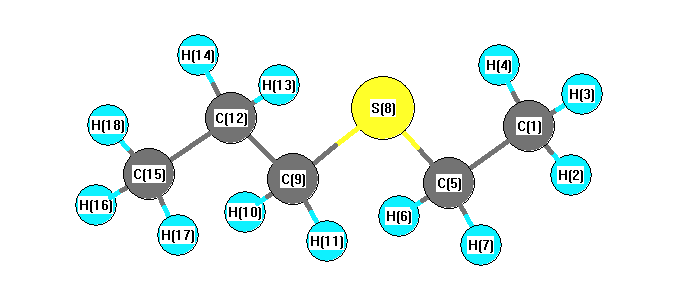
picture of Ethyl propyl sulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.091 C1 C5 H7 109.091
C1 C5 S8 110.683 H2 C1 H3 109.485
H2 C1 H4 109.485 H2 C1 C5 110.086
H3 C1 H4 106.076 H3 C1 C5 110.812
H4 C1 C5 110.812 C5 S8 C9 106.089
H6 C5 H7 108.976 H6 C5 S8 109.486
H7 C5 S8 109.486 S8 C9 H10 110.942
S8 C9 H11 110.942 S8 C9 C12 111.054
C9 C12 H13 108.673 C9 C12 H14 108.673
C9 C12 C15 110.590 H10 C9 H11 107.169
H10 C9 C12 108.296 H11 C9 C12 108.296
C12 C15 H16 110.784 C12 C15 H17 110.784
C12 C15 H18 109.924 H13 C12 H14 108.998
H13 C12 C15 109.934 H14 C12 C15 109.934
H16 C15 H17 105.828 H16 C15 H18 109.722
H17 C15 H18 109.722
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.475      
2 H 0.160      
3 H 0.171      
4 H 0.171      
5 C -0.422      
6 H 0.173      
7 H 0.173      
8 S 0.097      
9 C -0.425      
10 H 0.170      
11 H 0.170      
12 C -0.290      
13 H 0.163      
14 H 0.163      
15 C -0.462      
16 H 0.152      
17 H 0.152      
18 H 0.159      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 11.764 2.202 0.000
y 2.202 11.344 0.000
z 0.000 0.000 8.429


<r2> (average value of r2) Å2
<r2> 354.803
(<r2>)1/2 18.836