return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHFCH3 (2-Fluoropropane)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-217.726017
Energy at 298.15K-217.733935
HF Energy-217.117714
Nuclear repulsion energy132.345302
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3162 3011 31.58      
2 A' 3156 3005 47.44      
3 A' 3076 2929 4.58      
4 A' 3072 2926 37.88      
5 A' 1559 1484 6.33      
6 A' 1542 1468 4.52      
7 A' 1471 1401 17.73      
8 A' 1422 1354 18.55      
9 A' 1240 1181 9.19      
10 A' 1196 1139 41.96      
11 A' 993 946 35.02      
12 A' 851 810 9.05      
13 A' 485 462 3.33      
14 A' 358 341 0.93      
15 A' 273 260 0.15      
16 A" 3160 3009 16.61      
17 A" 3149 2999 0.06      
18 A" 3072 2925 13.16      
19 A" 1536 1463 0.05      
20 A" 1531 1458 0.03      
21 A" 1471 1401 29.67      
22 A" 1423 1355 1.86      
23 A" 1202 1145 10.35      
24 A" 976 930 0.01      
25 A" 956 910 0.48      
26 A" 411 392 7.13      
27 A" 231 220 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 21486.5 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 20461.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.29029 0.26894 0.15965

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.285 0.245 0.000
F2 -0.884 1.031 0.000
H3 1.131 0.948 0.000
C4 0.285 -0.581 1.272
C5 0.285 -0.581 -1.272
H6 1.198 -1.185 1.342
H7 1.198 -1.185 -1.342
H8 0.229 0.073 2.149
H9 0.229 0.073 -2.149
H10 -0.582 -1.252 1.281
H11 -0.582 -1.252 -1.281

Atom - Atom Distances (Å)
  C1 F2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.40921.09921.51671.51672.16292.16292.15702.15702.15262.1526
F21.40922.01672.36292.36293.32363.32362.60312.60312.63472.6347
H31.09922.01672.16092.16092.52052.52052.48942.48943.06783.0678
C41.51672.36292.16092.54471.09702.83411.09563.48421.09662.7789
C51.51672.36292.16092.54472.83411.09703.48421.09562.77891.0966
H62.16293.32362.52051.09702.83412.68351.78193.83551.78253.1706
H72.16293.32362.52052.83411.09702.68353.83551.78193.17061.7825
H82.15702.60312.48941.09563.48421.78193.83554.29881.77983.7659
H92.15702.60312.48943.48421.09563.83551.78194.29883.76591.7798
H102.15262.63473.06781.09662.77891.78253.17061.77983.76592.5622
H112.15262.63473.06782.77891.09663.17061.78253.76591.77982.5622

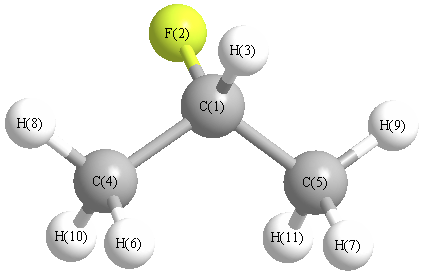
picture of 2-Fluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H8 110.280 C1 C4 H10 109.867
C1 C5 H7 110.663 C1 C5 H9 110.280
C1 C5 H11 109.867 F2 C1 H3 106.370
F2 C1 C4 107.664 F2 C1 C5 107.664
H3 C1 C4 110.370 H3 C1 C5 110.370
C4 C1 C5 114.040 H7 C5 H9 108.721
H7 C5 H11 108.699 H8 C4 H10 108.561
H9 C5 H11 108.561
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability