Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -376.458834 |
| Energy at 298.15K | |
| HF Energy | -375.785329 |
| Nuclear repulsion energy | 191.940444 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3281 |
3031 |
35.79 |
|
|
|
| 2 |
A |
3258 |
3009 |
21.66 |
|
|
|
| 3 |
A |
3213 |
2968 |
16.03 |
|
|
|
| 4 |
A |
1608 |
1485 |
4.37 |
|
|
|
| 5 |
A |
1571 |
1451 |
17.42 |
|
|
|
| 6 |
A |
1513 |
1398 |
34.48 |
|
|
|
| 7 |
A |
1433 |
1324 |
30.64 |
|
|
|
| 8 |
A |
1336 |
1234 |
15.97 |
|
|
|
| 9 |
A |
1254 |
1158 |
111.25 |
|
|
|
| 10 |
A |
1222 |
1129 |
82.74 |
|
|
|
| 11 |
A |
1209 |
1117 |
116.06 |
|
|
|
| 12 |
A |
1178 |
1088 |
48.79 |
|
|
|
| 13 |
A |
977 |
903 |
33.65 |
|
|
|
| 14 |
A |
603 |
557 |
5.33 |
|
|
|
| 15 |
A |
505 |
467 |
21.76 |
|
|
|
| 16 |
A |
447 |
413 |
6.77 |
|
|
|
| 17 |
A |
253 |
234 |
9.52 |
|
|
|
| 18 |
A |
129 |
119 |
8.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12494.5 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 11541.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.761 |
-0.603 |
-0.277 |
| C2 |
0.464 |
0.021 |
0.330 |
| F3 |
-1.861 |
0.104 |
0.147 |
| F4 |
1.534 |
-0.741 |
-0.010 |
| F5 |
0.650 |
1.257 |
-0.183 |
| H6 |
-0.690 |
-0.553 |
-1.360 |
| H7 |
-0.852 |
-1.637 |
0.045 |
| H8 |
0.414 |
0.100 |
1.412 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5030 | 1.3748 | 2.3143 | 2.3363 | 1.0866 | 1.0864 | 2.1740 |
C2 | 1.5030 | | 2.3343 | 1.3568 | 1.3513 | 2.1259 | 2.1359 | 1.0854 | F3 | 1.3748 | 2.3343 | | 3.5023 | 2.7823 | 2.0181 | 2.0149 | 2.6034 | F4 | 2.3143 | 1.3568 | 3.5023 | | 2.1920 | 2.6082 | 2.5491 | 1.9961 | F5 | 2.3363 | 1.3513 | 2.7823 | 2.1920 | | 2.5411 | 3.2682 | 1.9840 | H6 | 1.0866 | 2.1259 | 2.0181 | 2.6082 | 2.5411 | | 1.7817 | 3.0543 | H7 | 1.0864 | 2.1359 | 2.0149 | 2.5491 | 3.2682 | 1.7817 | | 2.5473 | H8 | 2.1740 | 1.0854 | 2.6034 | 1.9961 | 1.9840 | 3.0543 | 2.5473 | |
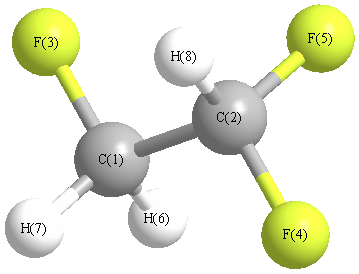 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.934 |
|
C1 |
C2 |
F5 |
109.758 |
| C1 |
C2 |
H8 |
113.275 |
|
C2 |
C1 |
F3 |
108.325 |
| C2 |
C1 |
H6 |
109.301 |
|
C2 |
C1 |
H7 |
110.106 |
| F3 |
C1 |
H6 |
109.593 |
|
F3 |
C1 |
H7 |
109.335 |
| F4 |
C2 |
F5 |
108.079 |
|
F4 |
C2 |
H8 |
109.137 |
| F5 |
C2 |
H8 |
108.533 |
|
H6 |
C1 |
H7 |
110.152 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -376.456080 |
| Energy at 298.15K | |
| HF Energy | -375.782293 |
| Nuclear repulsion energy | 195.011620 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3231 |
2985 |
56.71 |
|
|
|
| 2 |
A' |
3201 |
2957 |
10.22 |
|
|
|
| 3 |
A' |
1606 |
1483 |
5.86 |
|
|
|
| 4 |
A' |
1546 |
1428 |
35.07 |
|
|
|
| 5 |
A' |
1518 |
1402 |
26.32 |
|
|
|
| 6 |
A' |
1270 |
1173 |
118.62 |
|
|
|
| 7 |
A' |
1185 |
1095 |
16.45 |
|
|
|
| 8 |
A' |
947 |
875 |
40.30 |
|
|
|
| 9 |
A' |
801 |
740 |
56.54 |
|
|
|
| 10 |
A' |
533 |
492 |
14.65 |
|
|
|
| 11 |
A' |
245 |
227 |
2.08 |
|
|
|
| 12 |
A" |
3263 |
3014 |
24.51 |
|
|
|
| 13 |
A" |
1510 |
1395 |
45.48 |
|
|
|
| 14 |
A" |
1354 |
1251 |
21.92 |
|
|
|
| 15 |
A" |
1243 |
1148 |
96.19 |
|
|
|
| 16 |
A" |
1039 |
960 |
55.61 |
|
|
|
| 17 |
A" |
391 |
361 |
0.15 |
|
|
|
| 18 |
A" |
122 |
112 |
2.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12502.1 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 11548.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.746 |
-0.821 |
0.000 |
| C2 |
0.358 |
0.630 |
0.000 |
| F3 |
-0.389 |
-1.589 |
0.000 |
| F4 |
-0.389 |
0.903 |
1.093 |
| F5 |
-0.389 |
0.903 |
-1.093 |
| H6 |
1.330 |
-1.045 |
-0.890 |
| H7 |
1.330 |
-1.045 |
0.890 |
| H8 |
1.226 |
1.286 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5020 | 1.3710 | 2.3367 | 2.3367 | 1.0875 | 1.0875 | 2.1609 |
C2 | 1.5020 | | 2.3413 | 1.3525 | 1.3525 | 2.1312 | 2.1312 | 1.0877 | F3 | 1.3710 | 2.3413 | | 2.7219 | 2.7219 | 2.0107 | 2.0107 | 3.2976 | F4 | 2.3367 | 1.3525 | 2.7219 | | 2.1869 | 3.2692 | 2.6066 | 1.9874 | F5 | 2.3367 | 1.3525 | 2.7219 | 2.1869 | | 2.6066 | 3.2692 | 1.9874 | H6 | 1.0875 | 2.1312 | 2.0107 | 3.2692 | 2.6066 | | 1.7803 | 2.4976 | H7 | 1.0875 | 2.1312 | 2.0107 | 2.6066 | 3.2692 | 1.7803 | | 2.4976 | H8 | 2.1609 | 1.0877 | 3.2976 | 1.9874 | 1.9874 | 2.4976 | 2.4976 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.783 |
|
C1 |
C2 |
F5 |
109.783 |
| C1 |
C2 |
H8 |
112.124 |
|
C2 |
C1 |
F3 |
109.077 |
| C2 |
C1 |
H6 |
109.740 |
|
C2 |
C1 |
H7 |
109.740 |
| F3 |
C1 |
H6 |
109.197 |
|
F3 |
C1 |
H7 |
109.197 |
| F4 |
C2 |
F5 |
107.892 |
|
F4 |
C2 |
H8 |
108.576 |
| F5 |
C2 |
H8 |
108.576 |
|
H6 |
C1 |
H7 |
109.871 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
