Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -377.384118 |
| Energy at 298.15K | |
| HF Energy | -377.384118 |
| Nuclear repulsion energy | 190.731697 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3142 |
3006 |
39.14 |
|
|
|
| 2 |
A |
3107 |
2972 |
34.12 |
|
|
|
| 3 |
A |
3080 |
2947 |
20.05 |
|
|
|
| 4 |
A |
1529 |
1463 |
2.81 |
|
|
|
| 5 |
A |
1480 |
1416 |
13.98 |
|
|
|
| 6 |
A |
1438 |
1376 |
30.31 |
|
|
|
| 7 |
A |
1365 |
1305 |
26.06 |
|
|
|
| 8 |
A |
1278 |
1223 |
10.73 |
|
|
|
| 9 |
A |
1183 |
1132 |
99.67 |
|
|
|
| 10 |
A |
1153 |
1103 |
22.97 |
|
|
|
| 11 |
A |
1147 |
1097 |
201.43 |
|
|
|
| 12 |
A |
1124 |
1075 |
18.85 |
|
|
|
| 13 |
A |
924 |
884 |
28.14 |
|
|
|
| 14 |
A |
576 |
551 |
3.74 |
|
|
|
| 15 |
A |
479 |
458 |
16.99 |
|
|
|
| 16 |
A |
425 |
407 |
5.21 |
|
|
|
| 17 |
A |
238 |
227 |
7.40 |
|
|
|
| 18 |
A |
122 |
117 |
6.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11894.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 11379.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.771 |
-0.598 |
-0.275 |
| C2 |
0.471 |
0.022 |
0.329 |
| F3 |
-1.877 |
0.106 |
0.146 |
| F4 |
1.538 |
-0.755 |
-0.011 |
| F5 |
0.665 |
1.265 |
-0.182 |
| H6 |
-0.698 |
-0.556 |
-1.367 |
| H7 |
-0.855 |
-1.641 |
0.049 |
| H8 |
0.420 |
0.102 |
1.420 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5135 | 1.3771 | 2.3291 | 2.3541 | 1.0956 | 1.0955 | 2.1862 |
C2 | 1.5135 | | 2.3571 | 1.3632 | 1.3575 | 2.1391 | 2.1458 | 1.0956 | F3 | 1.3771 | 2.3571 | | 3.5258 | 2.8135 | 2.0291 | 2.0265 | 2.6274 | F4 | 2.3291 | 1.3632 | 3.5258 | | 2.2073 | 2.6223 | 2.5527 | 2.0081 | F5 | 2.3541 | 1.3575 | 2.8135 | 2.2073 | | 2.5650 | 3.2881 | 1.9950 | H6 | 1.0956 | 2.1391 | 2.0291 | 2.6223 | 2.5650 | | 1.7908 | 3.0741 | H7 | 1.0955 | 2.1458 | 2.0265 | 2.5527 | 3.2881 | 1.7908 | | 2.5581 | H8 | 2.1862 | 1.0956 | 2.6274 | 2.0081 | 1.9950 | 3.0741 | 2.5581 | |
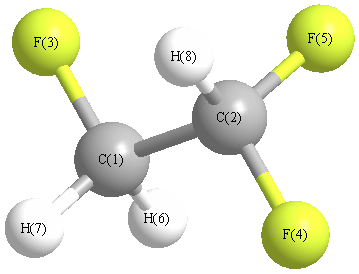 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.009 |
|
C1 |
C2 |
F5 |
110.043 |
| C1 |
C2 |
H8 |
112.864 |
|
C2 |
C1 |
F3 |
109.165 |
| C2 |
C1 |
H6 |
109.087 |
|
C2 |
C1 |
H7 |
109.620 |
| F3 |
C1 |
H6 |
109.766 |
|
F3 |
C1 |
H7 |
109.558 |
| F4 |
C2 |
F5 |
108.444 |
|
F4 |
C2 |
H8 |
109.027 |
| F5 |
C2 |
H8 |
108.363 |
|
H6 |
C1 |
H7 |
109.629 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.054 |
|
|
|
| 2 |
C |
0.404 |
|
|
|
| 3 |
F |
-0.299 |
|
|
|
| 4 |
F |
-0.286 |
|
|
|
| 5 |
F |
-0.277 |
|
|
|
| 6 |
H |
0.182 |
|
|
|
| 7 |
H |
0.172 |
|
|
|
| 8 |
H |
0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.177 |
-1.333 |
0.340 |
1.387 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.123 |
1.368 |
1.186 |
| y |
1.368 |
-27.006 |
0.468 |
| z |
1.186 |
0.468 |
-24.375 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.433 |
1.368 |
1.186 |
| y |
1.368 |
0.743 |
0.468 |
| z |
1.186 |
0.468 |
4.689 |
|
| Polar |
|---|
| 3z2-r2 | 9.379 |
|---|
| x2-y2 | -4.117 |
|---|
| xy | 1.368 |
|---|
| xz | 1.186 |
|---|
| yz | 0.468 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.485 |
-0.007 |
0.017 |
| y |
-0.007 |
3.518 |
0.033 |
| z |
0.017 |
0.033 |
3.450 |
<r2> (average value of r
2) Å
2
| <r2> |
110.738 |
| (<r2>)1/2 |
10.523 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -377.381681 |
| Energy at 298.15K | |
| HF Energy | -377.381681 |
| Nuclear repulsion energy | 193.512010 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3080 |
2946 |
73.38 |
|
|
|
| 2 |
A' |
3064 |
2931 |
9.21 |
|
|
|
| 3 |
A' |
1526 |
1460 |
4.48 |
|
|
|
| 4 |
A' |
1459 |
1396 |
19.50 |
|
|
|
| 5 |
A' |
1440 |
1377 |
36.17 |
|
|
|
| 6 |
A' |
1203 |
1151 |
117.44 |
|
|
|
| 7 |
A' |
1130 |
1081 |
18.66 |
|
|
|
| 8 |
A' |
889 |
851 |
30.86 |
|
|
|
| 9 |
A' |
769 |
736 |
45.58 |
|
|
|
| 10 |
A' |
511 |
488 |
11.21 |
|
|
|
| 11 |
A' |
227 |
218 |
1.63 |
|
|
|
| 12 |
A" |
3123 |
2988 |
30.08 |
|
|
|
| 13 |
A" |
1431 |
1369 |
38.82 |
|
|
|
| 14 |
A" |
1294 |
1238 |
14.61 |
|
|
|
| 15 |
A" |
1172 |
1121 |
96.98 |
|
|
|
| 16 |
A" |
980 |
938 |
55.74 |
|
|
|
| 17 |
A" |
370 |
354 |
0.18 |
|
|
|
| 18 |
A" |
109 |
105 |
1.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11889.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 11374.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.739 |
-0.831 |
0.000 |
| C2 |
0.355 |
0.631 |
0.000 |
| F3 |
-0.388 |
-1.617 |
0.000 |
| F4 |
-0.388 |
0.921 |
1.101 |
| F5 |
-0.388 |
0.921 |
-1.101 |
| H6 |
1.335 |
-1.049 |
-0.894 |
| H7 |
1.335 |
-1.049 |
0.894 |
| H8 |
1.243 |
1.277 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5124 | 1.3733 | 2.3560 | 2.3560 | 1.0967 | 1.0967 | 2.1677 |
C2 | 1.5124 | | 2.3676 | 1.3590 | 1.3590 | 2.1408 | 2.1408 | 1.0983 | F3 | 1.3733 | 2.3676 | | 2.7659 | 2.7659 | 2.0227 | 2.0227 | 3.3217 | F4 | 2.3560 | 1.3590 | 2.7659 | | 2.2015 | 3.2904 | 2.6247 | 1.9999 | F5 | 2.3560 | 1.3590 | 2.7659 | 2.2015 | | 2.6247 | 3.2904 | 1.9999 | H6 | 1.0967 | 2.1408 | 2.0227 | 3.2904 | 2.6247 | | 1.7888 | 2.4930 | H7 | 1.0967 | 2.1408 | 2.0227 | 2.6247 | 3.2904 | 1.7888 | | 2.4930 | H8 | 2.1677 | 1.0983 | 3.3217 | 1.9999 | 1.9999 | 2.4930 | 2.4930 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.161 |
|
C1 |
C2 |
F5 |
110.161 |
| C1 |
C2 |
H8 |
111.277 |
|
C2 |
C1 |
F3 |
110.173 |
| C2 |
C1 |
H6 |
109.241 |
|
C2 |
C1 |
H7 |
109.241 |
| F3 |
C1 |
H6 |
109.442 |
|
F3 |
C1 |
H7 |
109.442 |
| F4 |
C2 |
F5 |
108.186 |
|
F4 |
C2 |
H8 |
108.488 |
| F5 |
C2 |
H8 |
108.488 |
|
H6 |
C1 |
H7 |
109.284 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.042 |
|
|
|
| 2 |
C |
0.414 |
|
|
|
| 3 |
F |
-0.292 |
|
|
|
| 4 |
F |
-0.280 |
|
|
|
| 5 |
F |
-0.280 |
|
|
|
| 6 |
H |
0.170 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
| 8 |
H |
0.141 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.840 |
0.094 |
0.000 |
2.842 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.616 |
-0.299 |
0.000 |
| y |
-0.299 |
-28.072 |
0.000 |
| z |
0.000 |
0.000 |
-27.709 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.274 |
-0.299 |
0.000 |
| y |
-0.299 |
-1.909 |
0.000 |
| z |
0.000 |
0.000 |
-1.365 |
|
| Polar |
|---|
| 3z2-r2 | -2.730 |
|---|
| x2-y2 | 3.455 |
|---|
| xy | -0.299 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.506 |
0.015 |
0.000 |
| y |
0.015 |
3.503 |
0.000 |
| z |
0.000 |
0.000 |
3.510 |
<r2> (average value of r
2) Å
2
| <r2> |
102.502 |
| (<r2>)1/2 |
10.124 |
