Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -376.576899 |
| Energy at 298.15K | |
| HF Energy | -375.783383 |
| Nuclear repulsion energy | 190.399710 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3170 |
3019 |
35.08 |
|
|
|
| 2 |
A |
3144 |
2994 |
23.90 |
|
|
|
| 3 |
A |
3105 |
2957 |
16.83 |
|
|
|
| 4 |
A |
1558 |
1484 |
3.45 |
|
|
|
| 5 |
A |
1511 |
1439 |
16.10 |
|
|
|
| 6 |
A |
1460 |
1390 |
29.44 |
|
|
|
| 7 |
A |
1382 |
1316 |
26.43 |
|
|
|
| 8 |
A |
1287 |
1226 |
11.70 |
|
|
|
| 9 |
A |
1204 |
1146 |
94.21 |
|
|
|
| 10 |
A |
1180 |
1124 |
21.83 |
|
|
|
| 11 |
A |
1160 |
1104 |
160.12 |
|
|
|
| 12 |
A |
1125 |
1071 |
44.84 |
|
|
|
| 13 |
A |
944 |
899 |
32.74 |
|
|
|
| 14 |
A |
579 |
551 |
4.75 |
|
|
|
| 15 |
A |
489 |
466 |
20.10 |
|
|
|
| 16 |
A |
430 |
409 |
6.27 |
|
|
|
| 17 |
A |
243 |
232 |
8.95 |
|
|
|
| 18 |
A |
126 |
120 |
7.89 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12048.2 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 11473.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.759 |
-0.613 |
-0.277 |
| C2 |
0.466 |
0.022 |
0.335 |
| F3 |
-1.874 |
0.099 |
0.145 |
| F4 |
1.551 |
-0.741 |
-0.012 |
| F5 |
0.641 |
1.272 |
-0.184 |
| H6 |
-0.682 |
-0.564 |
-1.368 |
| H7 |
-0.848 |
-1.655 |
0.050 |
| H8 |
0.418 |
0.101 |
1.425 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5093 | 1.3891 | 2.3285 | 2.3502 | 1.0954 | 1.0954 | 2.1889 |
C2 | 1.5093 | | 2.3491 | 1.3708 | 1.3648 | 2.1359 | 2.1493 | 1.0945 | F3 | 1.3891 | 2.3491 | | 3.5302 | 2.7946 | 2.0376 | 2.0347 | 2.6248 | F4 | 2.3285 | 1.3708 | 3.5302 | | 2.2158 | 2.6186 | 2.5679 | 2.0152 | F5 | 2.3502 | 1.3648 | 2.7946 | 2.2158 | | 2.5545 | 3.2924 | 2.0025 | H6 | 1.0954 | 2.1359 | 2.0376 | 2.6186 | 2.5545 | | 1.7969 | 3.0751 | H7 | 1.0954 | 2.1493 | 2.0347 | 2.5679 | 3.2924 | 1.7969 | | 2.5647 | H8 | 2.1889 | 1.0945 | 2.6248 | 2.0152 | 2.0025 | 3.0751 | 2.5647 | |
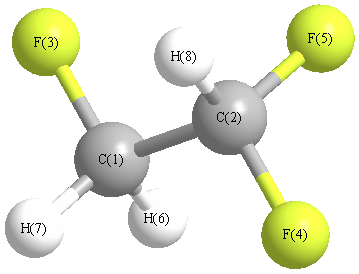 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.799 |
|
C1 |
C2 |
F5 |
109.611 |
| C1 |
C2 |
H8 |
113.462 |
|
C2 |
C1 |
F3 |
108.220 |
| C2 |
C1 |
H6 |
109.142 |
|
C2 |
C1 |
H7 |
110.201 |
| F3 |
C1 |
H6 |
109.631 |
|
F3 |
C1 |
H7 |
109.399 |
| F4 |
C2 |
F5 |
108.185 |
|
F4 |
C2 |
H8 |
109.136 |
| F5 |
C2 |
H8 |
108.525 |
|
H6 |
C1 |
H7 |
110.216 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -376.574278 |
| Energy at 298.15K | |
| HF Energy | -375.780333 |
| Nuclear repulsion energy | 193.441883 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3115 |
2967 |
58.90 |
|
|
|
| 2 |
A' |
3092 |
2945 |
9.76 |
|
|
|
| 3 |
A' |
1556 |
1482 |
4.50 |
|
|
|
| 4 |
A' |
1486 |
1416 |
30.49 |
|
|
|
| 5 |
A' |
1466 |
1396 |
23.93 |
|
|
|
| 6 |
A' |
1217 |
1159 |
100.52 |
|
|
|
| 7 |
A' |
1132 |
1078 |
13.48 |
|
|
|
| 8 |
A' |
915 |
872 |
38.07 |
|
|
|
| 9 |
A' |
773 |
736 |
51.41 |
|
|
|
| 10 |
A' |
511 |
486 |
13.05 |
|
|
|
| 11 |
A' |
237 |
226 |
1.88 |
|
|
|
| 12 |
A" |
3151 |
3001 |
25.20 |
|
|
|
| 13 |
A" |
1456 |
1387 |
39.52 |
|
|
|
| 14 |
A" |
1303 |
1241 |
17.32 |
|
|
|
| 15 |
A" |
1193 |
1136 |
86.07 |
|
|
|
| 16 |
A" |
1002 |
954 |
57.03 |
|
|
|
| 17 |
A" |
378 |
360 |
0.14 |
|
|
|
| 18 |
A" |
117 |
111 |
2.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12050.7 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 11475.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.755 |
-0.823 |
0.000 |
| C2 |
0.362 |
0.634 |
0.000 |
| F3 |
-0.393 |
-1.598 |
0.000 |
| F4 |
-0.393 |
0.907 |
1.105 |
| F5 |
-0.393 |
0.907 |
-1.105 |
| H6 |
1.342 |
-1.049 |
-0.898 |
| H7 |
1.342 |
-1.049 |
0.898 |
| H8 |
1.234 |
1.300 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5088 | 1.3853 | 2.3521 | 2.3521 | 1.0965 | 1.0965 | 2.1767 |
C2 | 1.5088 | | 2.3560 | 1.3662 | 1.3662 | 2.1445 | 2.1445 | 1.0972 | F3 | 1.3853 | 2.3560 | | 2.7376 | 2.7376 | 2.0299 | 2.0299 | 3.3237 | F4 | 2.3521 | 1.3662 | 2.7376 | | 2.2098 | 3.2940 | 2.6234 | 2.0058 | F5 | 2.3521 | 1.3662 | 2.7376 | 2.2098 | | 2.6234 | 3.2940 | 2.0058 | H6 | 1.0965 | 2.1445 | 2.0299 | 3.2940 | 2.6234 | | 1.7955 | 2.5175 | H7 | 1.0965 | 2.1445 | 2.0299 | 2.6234 | 3.2940 | 1.7955 | | 2.5175 | H8 | 2.1767 | 1.0972 | 3.3237 | 2.0058 | 2.0058 | 2.5175 | 2.5175 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.695 |
|
C1 |
C2 |
F5 |
109.695 |
| C1 |
C2 |
H8 |
112.325 |
|
C2 |
C1 |
F3 |
108.920 |
| C2 |
C1 |
H6 |
109.791 |
|
C2 |
C1 |
H7 |
109.791 |
| F3 |
C1 |
H6 |
109.197 |
|
F3 |
C1 |
H7 |
109.197 |
| F4 |
C2 |
F5 |
107.948 |
|
F4 |
C2 |
H8 |
108.534 |
| F5 |
C2 |
H8 |
108.534 |
|
H6 |
C1 |
H7 |
109.921 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
