Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -377.201260 |
| Energy at 298.15K | |
| HF Energy | -376.951571 |
| Nuclear repulsion energy | 190.567205 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3178 |
3016 |
37.43 |
33.16 |
0.66 |
0.80 |
| 2 |
A |
3148 |
2989 |
28.21 |
93.96 |
0.35 |
0.52 |
| 3 |
A |
3114 |
2955 |
18.38 |
93.57 |
0.09 |
0.17 |
| 4 |
A |
1555 |
1476 |
2.90 |
9.45 |
0.73 |
0.85 |
| 5 |
A |
1500 |
1424 |
13.51 |
3.12 |
0.66 |
0.79 |
| 6 |
A |
1455 |
1381 |
28.80 |
5.17 |
0.74 |
0.85 |
| 7 |
A |
1381 |
1310 |
27.78 |
3.37 |
0.75 |
0.86 |
| 8 |
A |
1288 |
1222 |
9.74 |
9.66 |
0.73 |
0.84 |
| 9 |
A |
1186 |
1126 |
92.89 |
5.01 |
0.69 |
0.82 |
| 10 |
A |
1170 |
1110 |
8.01 |
2.49 |
0.50 |
0.67 |
| 11 |
A |
1143 |
1085 |
200.86 |
1.44 |
0.45 |
0.63 |
| 12 |
A |
1115 |
1059 |
30.84 |
6.22 |
0.50 |
0.66 |
| 13 |
A |
934 |
887 |
32.94 |
5.43 |
0.34 |
0.50 |
| 14 |
A |
576 |
546 |
4.21 |
1.81 |
0.40 |
0.57 |
| 15 |
A |
485 |
460 |
18.37 |
0.78 |
0.72 |
0.84 |
| 16 |
A |
427 |
405 |
5.57 |
1.53 |
0.54 |
0.70 |
| 17 |
A |
241 |
228 |
8.07 |
0.05 |
0.54 |
0.70 |
| 18 |
A |
124 |
118 |
7.10 |
0.01 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 12009.1 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 11399.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.763 |
-0.608 |
-0.275 |
| C2 |
0.468 |
0.022 |
0.332 |
| F3 |
-1.875 |
0.101 |
0.145 |
| F4 |
1.547 |
-0.746 |
-0.012 |
| F5 |
0.649 |
1.270 |
-0.183 |
| H6 |
-0.687 |
-0.564 |
-1.364 |
| H7 |
-0.848 |
-1.647 |
0.051 |
| H8 |
0.420 |
0.101 |
1.420 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5104 | 1.3842 | 2.3287 | 2.3512 | 1.0927 | 1.0926 | 2.1850 |
C2 | 1.5104 | | 2.3527 | 1.3682 | 1.3621 | 2.1341 | 2.1447 | 1.0920 | F3 | 1.3842 | 2.3527 | | 3.5290 | 2.8011 | 2.0322 | 2.0294 | 2.6258 | F4 | 2.3287 | 1.3682 | 3.5290 | | 2.2139 | 2.6172 | 2.5600 | 2.0102 | F5 | 2.3512 | 1.3621 | 2.8011 | 2.2139 | | 2.5577 | 3.2874 | 1.9971 | H6 | 1.0927 | 2.1341 | 2.0322 | 2.6172 | 2.5577 | | 1.7897 | 3.0689 | H7 | 1.0926 | 2.1447 | 2.0294 | 2.5600 | 3.2874 | 1.7897 | | 2.5572 | H8 | 2.1850 | 1.0920 | 2.6258 | 2.0102 | 1.9971 | 3.0689 | 2.5572 | |
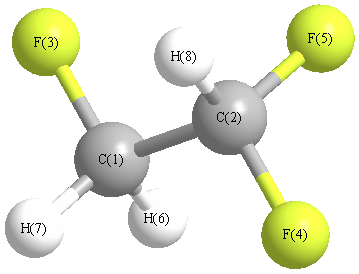 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.885 |
|
C1 |
C2 |
F5 |
109.765 |
| C1 |
C2 |
H8 |
113.216 |
|
C2 |
C1 |
F3 |
108.660 |
| C2 |
C1 |
H6 |
109.085 |
|
C2 |
C1 |
H7 |
109.921 |
| F3 |
C1 |
H6 |
109.704 |
|
F3 |
C1 |
H7 |
109.481 |
| F4 |
C2 |
F5 |
108.362 |
|
F4 |
C2 |
H8 |
109.072 |
| F5 |
C2 |
H8 |
108.436 |
|
H6 |
C1 |
H7 |
109.968 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.009 |
|
|
|
| 2 |
C |
0.445 |
|
|
|
| 3 |
F |
-0.317 |
|
|
|
| 4 |
F |
-0.303 |
|
|
|
| 5 |
F |
-0.293 |
|
|
|
| 6 |
H |
0.170 |
|
|
|
| 7 |
H |
0.160 |
|
|
|
| 8 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.169 |
-1.389 |
0.364 |
1.446 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.524 |
1.397 |
1.204 |
| y |
1.397 |
-27.238 |
0.482 |
| z |
1.204 |
0.482 |
-24.562 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.624 |
1.397 |
1.204 |
| y |
1.397 |
0.805 |
0.482 |
| z |
1.204 |
0.482 |
4.819 |
|
| Polar |
|---|
| 3z2-r2 | 9.639 |
|---|
| x2-y2 | -4.286 |
|---|
| xy | 1.397 |
|---|
| xz | 1.204 |
|---|
| yz | 0.482 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.440 |
-0.031 |
-0.003 |
| y |
-0.031 |
3.486 |
0.026 |
| z |
-0.003 |
0.026 |
3.392 |
<r2> (average value of r
2) Å
2
| <r2> |
110.870 |
| (<r2>)1/2 |
10.529 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -377.198717 |
| Energy at 298.15K | |
| HF Energy | -376.948918 |
| Nuclear repulsion energy | 193.526704 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3120 |
2961 |
66.77 |
101.08 |
0.09 |
0.16 |
| 2 |
A' |
3100 |
2942 |
8.94 |
66.20 |
0.31 |
0.48 |
| 3 |
A' |
1552 |
1473 |
4.07 |
8.44 |
0.73 |
0.85 |
| 4 |
A' |
1478 |
1403 |
18.17 |
2.75 |
0.57 |
0.73 |
| 5 |
A' |
1459 |
1385 |
36.59 |
4.72 |
0.69 |
0.81 |
| 6 |
A' |
1203 |
1142 |
108.95 |
5.63 |
0.42 |
0.59 |
| 7 |
A' |
1123 |
1066 |
15.42 |
3.11 |
0.75 |
0.86 |
| 8 |
A' |
902 |
856 |
35.18 |
7.25 |
0.18 |
0.30 |
| 9 |
A' |
770 |
731 |
48.44 |
2.89 |
0.53 |
0.69 |
| 10 |
A' |
509 |
483 |
11.71 |
1.55 |
0.75 |
0.86 |
| 11 |
A' |
233 |
221 |
1.76 |
0.11 |
0.39 |
0.57 |
| 12 |
A" |
3158 |
2998 |
27.63 |
53.53 |
0.75 |
0.86 |
| 13 |
A" |
1449 |
1375 |
37.43 |
2.02 |
0.75 |
0.86 |
| 14 |
A" |
1303 |
1237 |
13.69 |
16.68 |
0.75 |
0.86 |
| 15 |
A" |
1176 |
1116 |
87.61 |
0.90 |
0.75 |
0.86 |
| 16 |
A" |
992 |
941 |
65.22 |
4.08 |
0.75 |
0.86 |
| 17 |
A" |
376 |
356 |
0.13 |
0.16 |
0.75 |
0.86 |
| 18 |
A" |
112 |
106 |
1.93 |
0.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12005.9 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 11396.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.748 |
-0.826 |
0.000 |
| C2 |
0.359 |
0.633 |
0.000 |
| F3 |
-0.391 |
-1.605 |
0.000 |
| F4 |
-0.391 |
0.911 |
1.104 |
| F5 |
-0.391 |
0.911 |
-1.104 |
| H6 |
1.338 |
-1.047 |
-0.894 |
| H7 |
1.338 |
-1.047 |
0.894 |
| H8 |
1.236 |
1.289 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5097 | 1.3802 | 2.3527 | 2.3527 | 1.0937 | 1.0937 | 2.1703 |
C2 | 1.5097 | | 2.3601 | 1.3636 | 1.3636 | 2.1399 | 2.1399 | 1.0946 | F3 | 1.3802 | 2.3601 | | 2.7477 | 2.7477 | 2.0253 | 2.0253 | 3.3196 | F4 | 2.3527 | 1.3636 | 2.7477 | | 2.2080 | 3.2891 | 2.6211 | 2.0018 | F5 | 2.3527 | 1.3636 | 2.7477 | 2.2080 | | 2.6211 | 3.2891 | 2.0018 | H6 | 1.0937 | 2.1399 | 2.0253 | 3.2891 | 2.6211 | | 1.7880 | 2.5033 | H7 | 1.0937 | 2.1399 | 2.0253 | 2.6211 | 3.2891 | 1.7880 | | 2.5033 | H8 | 2.1703 | 1.0946 | 3.3196 | 2.0018 | 2.0018 | 2.5033 | 2.5033 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.826 |
|
C1 |
C2 |
F5 |
109.826 |
| C1 |
C2 |
H8 |
111.898 |
|
C2 |
C1 |
F3 |
109.420 |
| C2 |
C1 |
H6 |
109.528 |
|
C2 |
C1 |
H7 |
109.528 |
| F3 |
C1 |
H6 |
109.350 |
|
F3 |
C1 |
H7 |
109.350 |
| F4 |
C2 |
F5 |
108.112 |
|
F4 |
C2 |
H8 |
108.544 |
| F5 |
C2 |
H8 |
108.544 |
|
H6 |
C1 |
H7 |
109.650 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.002 |
|
|
|
| 2 |
C |
0.456 |
|
|
|
| 3 |
F |
-0.310 |
|
|
|
| 4 |
F |
-0.296 |
|
|
|
| 5 |
F |
-0.296 |
|
|
|
| 6 |
H |
0.157 |
|
|
|
| 7 |
H |
0.157 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.953 |
0.123 |
0.000 |
2.956 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.787 |
-0.296 |
-0.008 |
| y |
-0.296 |
-28.301 |
0.001 |
| z |
-0.008 |
0.001 |
-27.994 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.361 |
-0.296 |
-0.008 |
| y |
-0.296 |
-1.911 |
0.001 |
| z |
-0.008 |
0.001 |
-1.450 |
|
| Polar |
|---|
| 3z2-r2 | -2.899 |
|---|
| x2-y2 | 3.514 |
|---|
| xy | -0.296 |
|---|
| xz | -0.008 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.469 |
0.022 |
0.002 |
| y |
0.022 |
3.434 |
-0.001 |
| z |
0.002 |
-0.001 |
3.480 |
<r2> (average value of r
2) Å
2
| <r2> |
102.268 |
| (<r2>)1/2 |
10.113 |
