Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -377.181127 |
| Energy at 298.15K | |
| HF Energy | -377.181127 |
| Nuclear repulsion energy | 191.251767 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3156 |
3001 |
37.10 |
37.71 |
0.68 |
0.81 |
| 2 |
A |
3120 |
2967 |
33.88 |
93.56 |
0.35 |
0.52 |
| 3 |
A |
3093 |
2941 |
18.83 |
101.85 |
0.09 |
0.17 |
| 4 |
A |
1532 |
1457 |
3.02 |
9.47 |
0.73 |
0.84 |
| 5 |
A |
1489 |
1416 |
16.02 |
3.36 |
0.67 |
0.80 |
| 6 |
A |
1443 |
1373 |
31.77 |
5.17 |
0.74 |
0.85 |
| 7 |
A |
1368 |
1301 |
27.38 |
3.41 |
0.75 |
0.86 |
| 8 |
A |
1282 |
1219 |
12.03 |
9.55 |
0.73 |
0.84 |
| 9 |
A |
1195 |
1136 |
105.47 |
4.82 |
0.69 |
0.82 |
| 10 |
A |
1164 |
1107 |
138.57 |
2.54 |
0.75 |
0.86 |
| 11 |
A |
1159 |
1103 |
78.91 |
1.25 |
0.35 |
0.52 |
| 12 |
A |
1136 |
1080 |
17.82 |
5.91 |
0.47 |
0.64 |
| 13 |
A |
932 |
886 |
27.75 |
5.12 |
0.36 |
0.53 |
| 14 |
A |
581 |
552 |
3.89 |
1.71 |
0.43 |
0.60 |
| 15 |
A |
482 |
458 |
16.86 |
0.75 |
0.73 |
0.84 |
| 16 |
A |
429 |
408 |
5.48 |
1.48 |
0.53 |
0.69 |
| 17 |
A |
238 |
227 |
7.50 |
0.05 |
0.54 |
0.70 |
| 18 |
A |
122 |
116 |
6.63 |
0.01 |
0.70 |
0.82 |
Unscaled Zero Point Vibrational Energy (zpe) 11959.8 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 11373.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.767 |
-0.600 |
-0.272 |
| C2 |
0.470 |
0.022 |
0.329 |
| F3 |
-1.870 |
0.104 |
0.144 |
| F4 |
1.537 |
-0.748 |
-0.012 |
| F5 |
0.656 |
1.263 |
-0.182 |
| H6 |
-0.691 |
-0.561 |
-1.364 |
| H7 |
-0.851 |
-1.642 |
0.053 |
| H8 |
0.419 |
0.101 |
1.420 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5096 | 1.3737 | 2.3228 | 2.3460 | 1.0954 | 1.0950 | 2.1821 |
C2 | 1.5096 | | 2.3490 | 1.3596 | 1.3541 | 2.1339 | 2.1426 | 1.0954 | F3 | 1.3737 | 2.3490 | | 3.5155 | 2.7985 | 2.0271 | 2.0246 | 2.6207 | F4 | 2.3228 | 1.3596 | 3.5155 | | 2.2018 | 2.6119 | 2.5498 | 2.0061 | F5 | 2.3460 | 1.3541 | 2.7985 | 2.2018 | | 2.5568 | 3.2809 | 1.9931 | H6 | 1.0954 | 2.1339 | 2.0271 | 2.6119 | 2.5568 | | 1.7901 | 3.0694 | H7 | 1.0950 | 2.1426 | 2.0246 | 2.5498 | 3.2809 | 1.7901 | | 2.5531 | H8 | 2.1821 | 1.0954 | 2.6207 | 2.0061 | 1.9931 | 3.0694 | 2.5531 | |
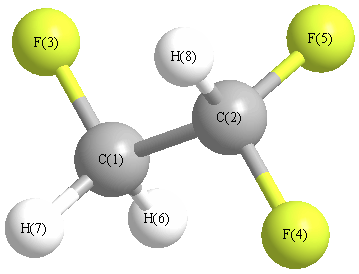 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.997 |
|
C1 |
C2 |
F5 |
109.892 |
| C1 |
C2 |
H8 |
112.825 |
|
C2 |
C1 |
F3 |
109.028 |
| C2 |
C1 |
H6 |
108.970 |
|
C2 |
C1 |
H7 |
109.671 |
| F3 |
C1 |
H6 |
109.856 |
|
F3 |
C1 |
H7 |
109.675 |
| F4 |
C2 |
F5 |
108.457 |
|
F4 |
C2 |
H8 |
109.133 |
| F5 |
C2 |
H8 |
108.450 |
|
H6 |
C1 |
H7 |
109.623 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.055 |
|
|
|
| 2 |
C |
0.405 |
|
|
|
| 3 |
F |
-0.300 |
|
|
|
| 4 |
F |
-0.287 |
|
|
|
| 5 |
F |
-0.278 |
|
|
|
| 6 |
H |
0.183 |
|
|
|
| 7 |
H |
0.174 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.170 |
-1.335 |
0.346 |
1.389 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.126 |
1.357 |
1.167 |
| y |
1.357 |
-27.031 |
0.463 |
| z |
1.167 |
0.463 |
-24.432 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.395 |
1.357 |
1.167 |
| y |
1.357 |
0.748 |
0.463 |
| z |
1.167 |
0.463 |
4.647 |
|
| Polar |
|---|
| 3z2-r2 | 9.293 |
|---|
| x2-y2 | -4.095 |
|---|
| xy | 1.357 |
|---|
| xz | 1.167 |
|---|
| yz | 0.463 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.457 |
-0.006 |
0.016 |
| y |
-0.006 |
3.498 |
0.033 |
| z |
0.016 |
0.033 |
3.440 |
<r2> (average value of r
2) Å
2
| <r2> |
110.167 |
| (<r2>)1/2 |
10.496 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -377.178700 |
| Energy at 298.15K | |
| HF Energy | -377.178700 |
| Nuclear repulsion energy | 194.110992 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3095 |
2944 |
71.55 |
115.96 |
0.05 |
0.09 |
| 2 |
A' |
3079 |
2928 |
8.29 |
58.81 |
0.49 |
0.66 |
| 3 |
A' |
1529 |
1454 |
4.86 |
8.38 |
0.73 |
0.84 |
| 4 |
A' |
1466 |
1395 |
26.17 |
3.07 |
0.58 |
0.73 |
| 5 |
A' |
1446 |
1375 |
32.22 |
4.49 |
0.71 |
0.83 |
| 6 |
A' |
1214 |
1155 |
116.97 |
5.35 |
0.39 |
0.57 |
| 7 |
A' |
1141 |
1085 |
17.89 |
3.09 |
0.74 |
0.85 |
| 8 |
A' |
900 |
856 |
30.79 |
6.87 |
0.21 |
0.35 |
| 9 |
A' |
775 |
737 |
46.23 |
2.70 |
0.54 |
0.70 |
| 10 |
A' |
514 |
489 |
11.52 |
1.50 |
0.75 |
0.86 |
| 11 |
A' |
229 |
218 |
1.65 |
0.10 |
0.38 |
0.55 |
| 12 |
A" |
3139 |
2985 |
28.48 |
55.64 |
0.75 |
0.86 |
| 13 |
A" |
1437 |
1366 |
41.61 |
1.78 |
0.75 |
0.86 |
| 14 |
A" |
1298 |
1235 |
16.34 |
16.87 |
0.75 |
0.86 |
| 15 |
A" |
1182 |
1124 |
96.99 |
1.02 |
0.75 |
0.86 |
| 16 |
A" |
987 |
938 |
52.12 |
3.80 |
0.75 |
0.86 |
| 17 |
A" |
373 |
354 |
0.16 |
0.15 |
0.75 |
0.86 |
| 18 |
A" |
110 |
104 |
1.83 |
0.01 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11956.8 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 11371.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.740 |
-0.828 |
0.000 |
| C2 |
0.355 |
0.631 |
0.000 |
| F3 |
-0.388 |
-1.606 |
0.000 |
| F4 |
-0.388 |
0.914 |
1.098 |
| F5 |
-0.388 |
0.914 |
-1.098 |
| H6 |
1.335 |
-1.046 |
-0.894 |
| H7 |
1.335 |
-1.046 |
0.894 |
| H8 |
1.241 |
1.279 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5086 | 1.3703 | 2.3478 | 2.3478 | 1.0962 | 1.0962 | 2.1658 |
C2 | 1.5086 | | 2.3573 | 1.3556 | 1.3556 | 2.1380 | 2.1380 | 1.0978 | F3 | 1.3703 | 2.3573 | | 2.7492 | 2.7492 | 2.0207 | 2.0207 | 3.3135 | F4 | 2.3478 | 1.3556 | 2.7492 | | 2.1955 | 3.2832 | 2.6178 | 1.9981 | F5 | 2.3478 | 1.3556 | 2.7492 | 2.1955 | | 2.6178 | 3.2832 | 1.9981 | H6 | 1.0962 | 2.1380 | 2.0207 | 3.2832 | 2.6178 | | 1.7883 | 2.4926 | H7 | 1.0962 | 2.1380 | 2.0207 | 2.6178 | 3.2832 | 1.7883 | | 2.4926 | H8 | 2.1658 | 1.0978 | 3.3135 | 1.9981 | 1.9981 | 2.4926 | 2.4926 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.997 |
|
C1 |
C2 |
F5 |
109.997 |
| C1 |
C2 |
H8 |
111.424 |
|
C2 |
C1 |
F3 |
109.844 |
| C2 |
C1 |
H6 |
109.308 |
|
C2 |
C1 |
H7 |
109.308 |
| F3 |
C1 |
H6 |
109.526 |
|
F3 |
C1 |
H7 |
109.526 |
| F4 |
C2 |
F5 |
108.142 |
|
F4 |
C2 |
H8 |
108.601 |
| F5 |
C2 |
H8 |
108.601 |
|
H6 |
C1 |
H7 |
109.313 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.044 |
|
|
|
| 2 |
C |
0.415 |
|
|
|
| 3 |
F |
-0.293 |
|
|
|
| 4 |
F |
-0.281 |
|
|
|
| 5 |
F |
-0.281 |
|
|
|
| 6 |
H |
0.171 |
|
|
|
| 7 |
H |
0.171 |
|
|
|
| 8 |
H |
0.142 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.837 |
0.099 |
0.000 |
2.839 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.661 |
-0.294 |
0.000 |
| y |
-0.294 |
-28.069 |
0.000 |
| z |
0.000 |
0.000 |
-27.739 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.243 |
-0.294 |
0.000 |
| y |
-0.294 |
-1.869 |
0.000 |
| z |
0.000 |
0.000 |
-1.374 |
|
| Polar |
|---|
| 3z2-r2 | -2.748 |
|---|
| x2-y2 | 3.408 |
|---|
| xy | -0.294 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.489 |
0.015 |
0.000 |
| y |
0.015 |
3.478 |
0.000 |
| z |
0.000 |
0.000 |
3.490 |
<r2> (average value of r
2) Å
2
| <r2> |
101.845 |
| (<r2>)1/2 |
10.092 |
