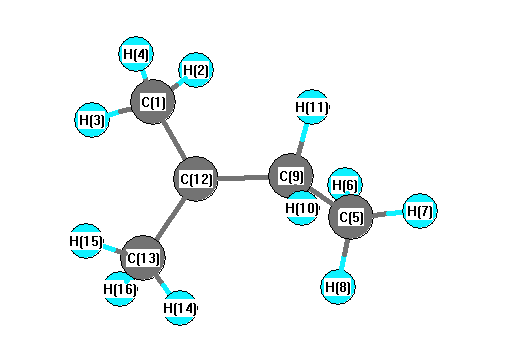
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3136 |
3000 |
40.58 |
|
|
|
| 2 |
A |
3127 |
2992 |
45.05 |
|
|
|
| 3 |
A |
3113 |
2978 |
40.71 |
|
|
|
| 4 |
A |
3110 |
2976 |
16.43 |
|
|
|
| 5 |
A |
3072 |
2939 |
19.90 |
|
|
|
| 6 |
A |
3061 |
2929 |
44.04 |
|
|
|
| 7 |
A |
3060 |
2928 |
11.91 |
|
|
|
| 8 |
A |
3052 |
2920 |
31.61 |
|
|
|
| 9 |
A |
3014 |
2883 |
44.86 |
|
|
|
| 10 |
A |
2972 |
2843 |
57.67 |
|
|
|
| 11 |
A |
2964 |
2836 |
40.02 |
|
|
|
| 12 |
A |
1531 |
1465 |
2.69 |
|
|
|
| 13 |
A |
1523 |
1457 |
5.80 |
|
|
|
| 14 |
A |
1520 |
1454 |
4.65 |
|
|
|
| 15 |
A |
1512 |
1446 |
2.93 |
|
|
|
| 16 |
A |
1503 |
1438 |
2.86 |
|
|
|
| 17 |
A |
1499 |
1434 |
8.03 |
|
|
|
| 18 |
A |
1494 |
1429 |
1.95 |
|
|
|
| 19 |
A |
1439 |
1377 |
1.10 |
|
|
|
| 20 |
A |
1425 |
1363 |
2.67 |
|
|
|
| 21 |
A |
1422 |
1361 |
3.16 |
|
|
|
| 22 |
A |
1373 |
1314 |
0.80 |
|
|
|
| 23 |
A |
1352 |
1294 |
1.78 |
|
|
|
| 24 |
A |
1298 |
1242 |
8.62 |
|
|
|
| 25 |
A |
1264 |
1210 |
0.81 |
|
|
|
| 26 |
A |
1079 |
1032 |
5.83 |
|
|
|
| 27 |
A |
1065 |
1019 |
1.55 |
|
|
|
| 28 |
A |
1033 |
989 |
2.30 |
|
|
|
| 29 |
A |
1027 |
982 |
1.79 |
|
|
|
| 30 |
A |
987 |
944 |
1.36 |
|
|
|
| 31 |
A |
953 |
911 |
0.84 |
|
|
|
| 32 |
A |
947 |
906 |
0.13 |
|
|
|
| 33 |
A |
789 |
755 |
3.07 |
|
|
|
| 34 |
A |
746 |
714 |
0.76 |
|
|
|
| 35 |
A |
417 |
399 |
1.26 |
|
|
|
| 36 |
A |
384 |
367 |
0.12 |
|
|
|
| 37 |
A |
344 |
329 |
1.24 |
|
|
|
| 38 |
A |
226 |
216 |
0.03 |
|
|
|
| 39 |
A |
170 |
163 |
1.53 |
|
|
|
| 40 |
A |
95 |
91 |
0.01 |
|
|
|
| 41 |
A |
74 |
71 |
0.19 |
|
|
|
| 42 |
A |
19 |
18 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32095.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30705.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.560 |
|
|
-0.367 |
| 2 |
H |
0.171 |
|
|
0.099 |
| 3 |
H |
0.169 |
|
|
0.099 |
| 4 |
H |
0.169 |
|
|
0.098 |
| 5 |
C |
-0.500 |
|
|
-0.195 |
| 6 |
H |
0.167 |
|
|
0.050 |
| 7 |
H |
0.162 |
|
|
0.054 |
| 8 |
H |
0.168 |
|
|
0.046 |
| 9 |
C |
-0.365 |
|
|
0.001 |
| 10 |
H |
0.161 |
|
|
0.023 |
| 11 |
H |
0.162 |
|
|
0.014 |
| 12 |
C |
0.148 |
|
|
0.168 |
| 13 |
C |
-0.561 |
|
|
-0.435 |
| 14 |
H |
0.171 |
|
|
0.115 |
| 15 |
H |
0.169 |
|
|
0.109 |
| 16 |
H |
0.169 |
|
|
0.121 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.070 |
-0.004 |
0.113 |
0.133 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.107 |
0.003 |
0.098 |
0.145 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.667 |
-0.036 |
0.372 |
| y |
-0.036 |
-33.255 |
0.043 |
| z |
0.372 |
0.043 |
-34.074 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.002 |
-0.036 |
0.372 |
| y |
-0.036 |
0.615 |
0.043 |
| z |
0.372 |
0.043 |
-0.613 |
|
| Polar |
|---|
| 3z2-r2 | -1.226 |
|---|
| x2-y2 | -0.412 |
|---|
| xy | -0.036 |
|---|
| xz | 0.372 |
|---|
| yz | 0.043 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
155.052 |
| (<r2>)1/2 |
12.452 |