return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H11 (2-Methylbut-2-yl radical)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-196.834439
Energy at 298.15K-196.845213
HF Energy-196.625243
Nuclear repulsion energy181.728324
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3160 3000 36.63      
2 A 3154 2994 30.46      
3 A 3147 2988 43.21      
4 A 3138 2979 24.47      
5 A 3103 2946 26.66      
6 A 3092 2935 29.37      
7 A 3083 2926 22.38      
8 A 3077 2921 24.28      
9 A 2994 2842 60.94      
10 A 2988 2837 28.57      
11 A 2964 2814 39.22      
12 A 1560 1481 1.82      
13 A 1552 1473 5.06      
14 A 1544 1466 2.61      
15 A 1536 1458 14.79      
16 A 1534 1456 1.53      
17 A 1524 1446 2.07      
18 A 1520 1442 0.53      
19 A 1464 1390 2.67      
20 A 1456 1382 1.86      
21 A 1446 1373 2.49      
22 A 1414 1342 0.21      
23 A 1337 1269 0.89      
24 A 1313 1246 7.39      
25 A 1276 1211 6.20      
26 A 1140 1082 0.79      
27 A 1090 1035 0.98      
28 A 1066 1012 3.99      
29 A 1020 969 0.72      
30 A 1009 958 1.57      
31 A 964 915 0.62      
32 A 959 910 0.28      
33 A 786 746 1.37      
34 A 750 712 1.37      
35 A 457 433 0.76      
36 A 384 364 0.08      
37 A 310 294 1.73      
38 A 282 268 0.09      
39 A 208 197 0.92      
40 A 135 128 0.01      
41 A 122 116 0.05      
42 A 41 39 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 32549.2 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30895.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.24531 0.11415 0.08466

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.615 1.481 0.069
H2 0.257 2.015 -0.319
H3 -1.504 1.876 -0.434
H4 -0.703 1.744 1.136
C5 2.060 -0.042 -0.191
H6 2.196 0.950 0.247
H7 2.927 -0.648 0.087
H8 2.059 0.069 -1.280
C9 0.763 -0.704 0.290
H10 0.728 -1.738 -0.077
H11 0.796 -0.782 1.391
C12 -0.504 -0.003 -0.120
C13 -1.772 -0.804 -0.118
H14 -1.617 -1.804 -0.536
H15 -2.162 -0.940 0.905
H16 -2.560 -0.309 -0.695

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 C13 H14 H15 H16
C11.09421.09441.10303.08912.86674.13283.31152.59293.49072.97681.50002.56753.48692.99202.7511
H21.09421.77001.76472.73792.28353.79252.82112.83233.78993.32262.16593.47914.25964.01053.6715
H31.09441.77001.76724.05393.87425.12564.08373.50984.26243.96032.15162.71173.68313.18692.4411
H41.10301.76471.76723.54773.13484.47224.03432.97673.95502.94852.16133.03414.02723.06373.3191
C53.08912.73794.05393.54771.09341.09401.09401.53372.15892.15632.56453.90744.09174.45324.6548
H62.86672.28353.87423.13481.09341.76441.76862.18963.07972.50402.88664.35404.76874.79585.0096
H74.13283.79255.12564.47221.09401.76441.77052.17472.45972.50223.49694.70634.73045.16285.5531
H83.31152.82114.08374.03431.09401.76861.77052.17762.54603.07472.81384.09764.19244.85894.6715
C92.59292.83233.50982.97671.53372.18962.17472.17761.09681.10471.50452.56932.74852.99803.4882
H103.49073.78994.26243.95502.15893.07972.45972.54601.09681.75272.12792.66942.39103.15483.6384
H112.97683.32263.96032.94852.15632.50402.50223.07471.10471.75272.14022.97873.25293.00193.9799
C121.50002.16592.15162.16132.56452.88663.49692.81381.50452.12792.14021.50012.15772.16332.1572
C132.56753.47912.71173.03413.90744.35404.70634.09762.56932.66942.97871.50011.09501.10301.0950
H143.48694.25963.68314.02724.09174.76874.73044.19242.74852.39103.25292.15771.09501.76601.7746
H152.99204.01053.18693.06374.45324.79585.16284.85892.99803.15483.00192.16331.10301.76601.7654
H162.75113.67152.44113.31914.65485.00965.55314.67153.48823.63843.97992.15721.09501.77461.7654

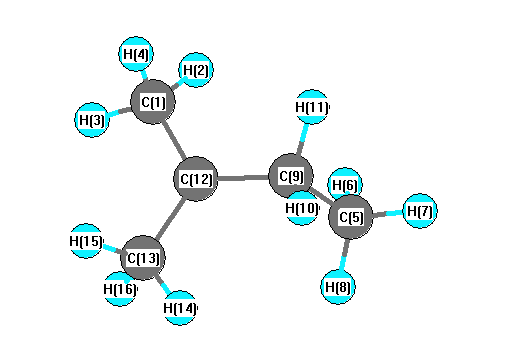
picture of 2-Methylbut-2-yl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C12 C9 119.312 C1 C12 C13 117.693
H2 C1 H3 107.950 H2 C1 H4 106.862
H2 C1 C12 112.264 H3 C1 H4 107.067
H3 C1 C12 111.098 H4 C1 C12 111.343
C5 C9 H10 109.190 C5 C9 H11 108.527
C5 C9 C12 115.140 H6 C5 H7 107.527
H6 C5 H8 107.905 H6 C5 C9 111.818
H7 C5 H8 108.032 H7 C5 C9 110.583
H8 C5 C9 110.823 C9 C12 C13 117.547
H10 C9 H11 105.524 H10 C9 C12 108.768
H11 C9 C12 109.259 C12 C13 H14 111.546
C12 C13 H15 111.501 C12 C13 H16 111.504
H14 C13 H15 106.920 H14 C13 H16 108.255
H15 C13 H16 106.870
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.501      
2 H 0.161      
3 H 0.154      
4 H 0.149      
5 C -0.465      
6 H 0.152      
7 H 0.150      
8 H 0.154      
9 C -0.303      
10 H 0.145      
11 H 0.141      
12 C 0.101      
13 C -0.497      
14 H 0.154      
15 H 0.150      
16 H 0.154      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.746 -0.026 0.095
y -0.026 8.004 0.001
z 0.095 0.001 6.708


<r2> (average value of r2) Å2
<r2> 156.109
(<r2>)1/2 12.494