return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CCNH2 (Diaminoacetylene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-187.381973
Energy at 298.15K-187.386106
HF Energy-186.870288
Nuclear repulsion energy101.939647
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3748 3462 21.47      
2 A 3656 3377 4.61      
3 A 2507 2315 0.03      
4 A 1753 1620 16.08      
5 A 1274 1177 0.01      
6 A 854 789 10.08      
7 A 685 633 145.52      
8 A 414 383 0.59      
9 A 359 332 39.12      
10 A 221 204 15.37      
11 B 3748 3462 20.44      
12 B 3655 3376 16.65      
13 B 1754 1620 41.79      
14 B 1401 1294 82.07      
15 B 1274 1176 0.03      
16 B 739 682 421.74      
17 B 398 368 19.50      
18 B 218 202 16.19      

Unscaled Zero Point Vibrational Energy (zpe) 14328.9 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 13235.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
5.12638 0.11942 0.11938

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.005 0.601 0.036
C2 -0.005 -0.601 0.036
N3 -0.005 1.958 -0.076
N4 0.005 -1.958 -0.076
H5 -0.327 2.444 0.745
H6 0.850 2.356 -0.429
H7 0.327 -2.444 0.745
H8 -0.850 -2.356 -0.429

Atom - Atom Distances (Å)
  C1 C2 N3 N4 H5 H6 H7 H8
C11.20251.36152.56172.00242.00213.14333.1133
C21.20252.56171.36153.14333.11332.00242.0021
N31.36152.56173.91611.00711.00734.49064.4099
N42.56171.36153.91614.49064.40991.00711.0073
H52.00243.14331.00714.49061.66474.93224.9690
H62.00213.11331.00734.40991.66474.96905.0092
H73.14332.00244.49061.00714.93224.96901.6647
H83.11332.00214.40991.00734.96905.00921.6647

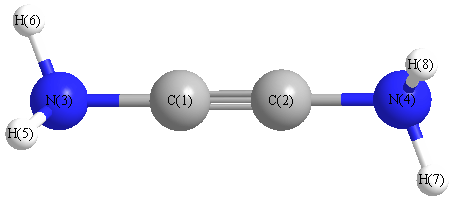
picture of Diaminoacetylene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N4 175.154 C1 N3 H5 114.597
C1 N3 H6 114.564 C2 C1 N3 175.154
C2 N4 H7 114.597 C2 N4 H8 114.564
H5 N3 H6 111.470 H7 N4 H8 111.470
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability