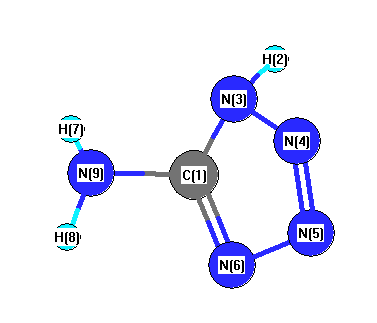
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3659 |
3514 |
67.23 |
|
|
|
| 2 |
A |
3642 |
3497 |
31.26 |
|
|
|
| 3 |
A |
3546 |
3405 |
26.63 |
|
|
|
| 4 |
A |
1691 |
1624 |
174.74 |
|
|
|
| 5 |
A |
1629 |
1565 |
108.05 |
|
|
|
| 6 |
A |
1501 |
1441 |
25.78 |
|
|
|
| 7 |
A |
1402 |
1347 |
12.59 |
|
|
|
| 8 |
A |
1299 |
1248 |
13.06 |
|
|
|
| 9 |
A |
1148 |
1102 |
8.62 |
|
|
|
| 10 |
A |
1112 |
1068 |
11.65 |
|
|
|
| 11 |
A |
1073 |
1030 |
15.66 |
|
|
|
| 12 |
A |
1023 |
982 |
16.78 |
|
|
|
| 13 |
A |
1010 |
970 |
10.52 |
|
|
|
| 14 |
A |
763 |
733 |
86.85 |
|
|
|
| 15 |
A |
734 |
705 |
0.56 |
|
|
|
| 16 |
A |
718 |
690 |
6.48 |
|
|
|
| 17 |
A |
662 |
636 |
237.58 |
|
|
|
| 18 |
A |
481 |
462 |
86.98 |
|
|
|
| 19 |
A |
384 |
369 |
8.56 |
|
|
|
| 20 |
A |
305 |
293 |
5.43 |
|
|
|
| 21 |
A |
268 |
258 |
51.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14025.7 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 13468.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.724 |
0.703 |
|
0.678 |
| 2 |
H |
0.354 |
0.255 |
|
0.272 |
| 3 |
N |
-0.492 |
-0.055 |
|
-0.069 |
| 4 |
N |
-0.048 |
-0.267 |
|
-0.263 |
| 5 |
N |
-0.086 |
-0.041 |
|
-0.047 |
| 6 |
N |
-0.383 |
-0.447 |
|
-0.437 |
| 7 |
H |
0.337 |
0.351 |
|
0.351 |
| 8 |
H |
0.357 |
0.388 |
|
0.391 |
| 9 |
N |
-0.763 |
-0.888 |
|
-0.874 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.935 |
3.500 |
1.016 |
6.135 |
| CHELPG |
-4.691 |
3.353 |
1.147 |
5.880 |
| AIM |
|
|
|
|
| ESP |
-4.793 |
3.409 |
1.135 |
5.990 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.414 |
0.703 |
-3.173 |
| y |
0.703 |
-31.675 |
0.480 |
| z |
-3.173 |
0.480 |
-34.473 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.339 |
0.703 |
-3.173 |
| y |
0.703 |
2.268 |
0.480 |
| z |
-3.173 |
0.480 |
-1.929 |
|
| Polar |
|---|
| 3z2-r2 | -3.858 |
|---|
| x2-y2 | -1.738 |
|---|
| xy | 0.703 |
|---|
| xz | -3.173 |
|---|
| yz | 0.480 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.803 |
-0.047 |
-0.060 |
| y |
-0.047 |
6.218 |
0.080 |
| z |
-0.060 |
0.080 |
2.788 |
<r2> (average value of r
2) Å
2
| <r2> |
120.729 |
| (<r2>)1/2 |
10.988 |