Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -153.839471 |
| Energy at 298.15K | |
| HF Energy | -153.440342 |
| Nuclear repulsion energy | 74.888573 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3886 |
3590 |
18.82 |
|
|
|
| 2 |
A' |
3267 |
3018 |
7.92 |
|
|
|
| 3 |
A' |
3107 |
2870 |
59.25 |
|
|
|
| 4 |
A' |
1615 |
1491 |
1.60 |
|
|
|
| 5 |
A' |
1562 |
1442 |
5.39 |
|
|
|
| 6 |
A' |
1508 |
1393 |
7.32 |
|
|
|
| 7 |
A' |
1296 |
1197 |
80.54 |
|
|
|
| 8 |
A' |
1130 |
1044 |
102.61 |
|
|
|
| 9 |
A' |
1056 |
975 |
9.27 |
|
|
|
| 10 |
A' |
632 |
584 |
16.69 |
|
|
|
| 11 |
A' |
388 |
359 |
35.72 |
|
|
|
| 12 |
A" |
3373 |
3116 |
12.54 |
|
|
|
| 13 |
A" |
3144 |
2904 |
61.87 |
|
|
|
| 14 |
A" |
1353 |
1250 |
0.32 |
|
|
|
| 15 |
A" |
1230 |
1136 |
1.80 |
|
|
|
| 16 |
A" |
844 |
780 |
0.45 |
|
|
|
| 17 |
A" |
291 |
269 |
146.43 |
|
|
|
| 18 |
A" |
40i |
37i |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14820.6 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 13689.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.106 |
-0.363 |
0.000 |
| C2 |
0.000 |
0.540 |
0.000 |
| C3 |
1.259 |
-0.256 |
0.000 |
| H4 |
-1.915 |
0.162 |
0.000 |
| H5 |
-0.035 |
1.185 |
0.884 |
| H6 |
-0.035 |
1.185 |
-0.884 |
| H7 |
1.639 |
-0.665 |
-0.925 |
| H8 |
1.639 |
-0.665 |
0.925 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4277 | 2.3682 | 0.9637 | 2.0797 | 2.0797 | 2.9128 | 2.9128 |
C2 | 1.4277 | | 1.4895 | 1.9518 | 1.0951 | 1.0951 | 2.2347 | 2.2347 | C3 | 2.3682 | 1.4895 | | 3.2017 | 2.1286 | 2.1286 | 1.0803 | 1.0803 | H4 | 0.9637 | 1.9518 | 3.2017 | | 2.3160 | 2.3160 | 3.7643 | 3.7643 | H5 | 2.0797 | 1.0951 | 2.1286 | 2.3160 | | 1.7687 | 3.0816 | 2.4950 | H6 | 2.0797 | 1.0951 | 2.1286 | 2.3160 | 1.7687 | | 2.4950 | 3.0816 | H7 | 2.9128 | 2.2347 | 1.0803 | 3.7643 | 3.0816 | 2.4950 | | 1.8494 | H8 | 2.9128 | 2.2347 | 1.0803 | 3.7643 | 2.4950 | 3.0816 | 1.8494 | |
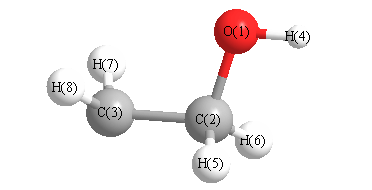 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.530 |
|
O1 |
C2 |
H5 |
110.352 |
| O1 |
C2 |
H6 |
110.352 |
|
C2 |
O1 |
H4 |
107.833 |
| C2 |
C3 |
H7 |
119.975 |
|
C2 |
C3 |
H8 |
119.975 |
| C3 |
C2 |
H5 |
109.947 |
|
C3 |
C2 |
H6 |
109.947 |
| H5 |
C2 |
H6 |
107.712 |
|
H7 |
C3 |
H8 |
117.733 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -153.840580 |
| Energy at 298.15K | -153.845307 |
| HF Energy | -153.441210 |
| Nuclear repulsion energy | 75.127150 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3906 |
3608 |
25.89 |
|
|
|
| 2 |
A |
3389 |
3130 |
6.74 |
|
|
|
| 3 |
A |
3276 |
3026 |
5.59 |
|
|
|
| 4 |
A |
3098 |
2861 |
65.50 |
|
|
|
| 5 |
A |
3044 |
2812 |
65.85 |
|
|
|
| 6 |
A |
1601 |
1479 |
3.01 |
|
|
|
| 7 |
A |
1554 |
1435 |
9.39 |
|
|
|
| 8 |
A |
1502 |
1387 |
2.15 |
|
|
|
| 9 |
A |
1337 |
1235 |
82.98 |
|
|
|
| 10 |
A |
1291 |
1192 |
10.10 |
|
|
|
| 11 |
A |
1185 |
1095 |
33.49 |
|
|
|
| 12 |
A |
1125 |
1039 |
35.17 |
|
|
|
| 13 |
A |
1030 |
951 |
7.68 |
|
|
|
| 14 |
A |
919 |
849 |
14.83 |
|
|
|
| 15 |
A |
466 |
430 |
18.45 |
|
|
|
| 16 |
A |
401 |
370 |
47.00 |
|
|
|
| 17 |
A |
293 |
271 |
122.97 |
|
|
|
| 18 |
A |
158 |
146 |
17.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14785.9 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 13657.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.101 |
-0.382 |
-0.050 |
| C2 |
-0.015 |
0.525 |
0.038 |
| C3 |
1.248 |
-0.255 |
-0.031 |
| H4 |
-1.918 |
0.115 |
0.061 |
| H5 |
-0.065 |
1.093 |
0.980 |
| H6 |
-0.048 |
1.261 |
-0.776 |
| H7 |
2.190 |
0.248 |
-0.193 |
| H8 |
1.249 |
-1.285 |
0.289 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4180 | 2.3533 | 0.9624 | 2.0763 | 2.0826 | 3.3543 | 2.5405 |
C2 | 1.4180 | | 1.4861 | 1.9474 | 1.1012 | 1.0976 | 2.2342 | 2.2219 | C3 | 2.3533 | 1.4861 | | 3.1896 | 2.1363 | 2.1289 | 1.0799 | 1.0790 | H4 | 0.9624 | 1.9474 | 3.1896 | | 2.2886 | 2.3482 | 4.1185 | 3.4703 | H5 | 2.0763 | 1.1012 | 2.1363 | 2.2886 | | 1.7633 | 2.6789 | 2.8039 | H6 | 2.0826 | 1.0976 | 2.1289 | 2.3482 | 1.7633 | | 2.5244 | 3.0493 | H7 | 3.3543 | 2.2342 | 1.0799 | 4.1185 | 2.6789 | 2.5244 | | 1.8628 | H8 | 2.5405 | 2.2219 | 1.0790 | 3.4703 | 2.8039 | 3.0493 | 1.8628 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.235 |
|
O1 |
C2 |
H5 |
110.374 |
| O1 |
C2 |
H6 |
111.118 |
|
C2 |
O1 |
H4 |
108.263 |
| C2 |
C3 |
H7 |
120.259 |
|
C2 |
C3 |
H8 |
119.192 |
| C3 |
C2 |
H5 |
110.435 |
|
C3 |
C2 |
H6 |
110.062 |
| H5 |
C2 |
H6 |
106.629 |
|
H7 |
C3 |
H8 |
119.278 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
