return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H7N (Cyclobutanecarbonitrile)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-249.170938
Energy at 298.15K-249.178858
HF Energy-248.903466
Nuclear repulsion energy210.524256
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3184 3023 42.47      
2 A 3177 3015 14.49      
3 A 3164 3003 9.50      
4 A 3121 2963 2.87      
5 A 3117 2959 41.67      
6 A 3115 2956 4.56      
7 A 3111 2953 25.85      
8 A 2294 2177 4.73      
9 A 1561 1482 0.78      
10 A 1536 1458 3.53      
11 A 1529 1451 1.26      
12 A 1393 1323 2.39      
13 A 1313 1246 1.47      
14 A 1311 1244 0.56      
15 A 1287 1221 0.28      
16 A 1275 1210 0.12      
17 A 1266 1201 0.14      
18 A 1233 1171 0.37      
19 A 1156 1098 2.20      
20 A 1093 1038 0.06      
21 A 1054 1000 0.04      
22 A 977 927 0.55      
23 A 968 919 0.40      
24 A 957 908 1.77      
25 A 920 873 2.05      
26 A 810 769 1.09      
27 A 768 729 1.45      
28 A 596 566 0.79      
29 A 534 507 0.01      
30 A 529 502 0.74      
31 A 272 258 1.57      
32 A 180 170 4.71      
33 A 136 129 3.13      

Unscaled Zero Point Vibrational Energy (zpe) 24467.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23224.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.32731 0.07888 0.06971

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.666 1.976 0.000
C2 0.685 1.343 0.000
C3 -0.469 0.455 0.000
C4 -0.510 -1.752 0.000
C5 -0.469 -0.654 1.098
C6 -0.469 -0.654 -1.098
H7 -1.377 1.064 0.000
H8 -1.415 -2.362 0.000
H9 0.351 -2.422 0.000
H10 0.427 -0.656 1.720
H11 -1.342 -0.616 1.751
H12 0.427 -0.656 -1.720
H13 -1.342 -0.616 -1.751

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13
N11.16802.62194.31703.56183.56183.17715.32154.59073.37994.34083.37994.3408
C21.16801.45613.31722.55452.55452.08094.25863.77912.64923.31912.64923.3191
C32.62191.45612.20721.56111.56111.09302.97182.99142.23502.23122.23502.2312
C44.31703.31722.20721.55331.55332.94601.09141.09112.24462.24692.24462.2469
C53.56182.55451.56111.55332.19682.23222.24022.23671.09041.09112.95742.9808
C63.56182.55451.56111.55332.19682.23222.24022.23672.95742.98081.09041.0911
H73.17712.08091.09302.94602.23222.23223.42633.89043.02812.42743.02812.4274
H85.32154.25862.97181.09142.24022.24023.42631.76693.04352.47393.04352.4739
H94.59073.77912.99141.09112.23672.23673.89041.76692.46643.03202.46643.0320
H103.37992.64922.23502.24461.09042.95743.02813.04352.46641.76983.44023.8965
H114.34083.31912.23122.24691.09112.98082.42742.47393.03201.76983.89653.5028
H123.37992.64922.23502.24462.95741.09043.02813.04352.46643.44023.89651.7698
H134.34083.31912.23122.24692.98081.09112.42742.47393.03203.89653.50281.7698

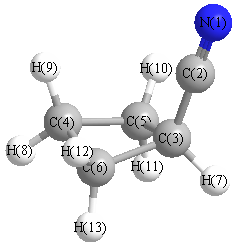
picture of Cyclobutanecarbonitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 175.324 C2 C3 C5 115.659
C2 C3 C6 115.659 C2 C3 H7 108.603
C3 C5 C4 90.261 C3 C5 H10 113.716
C3 C5 H11 113.360 C3 C6 C4 90.261
C3 C6 H12 113.716 C3 C6 H13 113.360
C4 C5 H10 115.088 C4 C5 H11 115.238
C4 C6 H12 115.088 C4 C6 H13 115.238
C5 C3 C6 89.433 C5 C3 H7 113.319
C5 C4 C6 90.004 C5 C4 H8 114.659
C5 C4 H9 114.389 C6 C3 H7 113.319
C6 C4 H8 114.659 C6 C4 H9 114.389
H8 C4 H9 108.107 H10 C5 H11 108.441
H12 C6 H13 108.441
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 N -0.478      
2 C 0.334      
3 C -0.197      
4 C -0.308      
5 C -0.291      
6 C -0.291      
7 H 0.203      
8 H 0.162      
9 H 0.167      
10 H 0.178      
11 H 0.171      
12 H 0.178      
13 H 0.171      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.791 -0.000 0.484
y -0.000 6.704 0.000
z 0.484 0.000 6.406


<r2> (average value of r2) Å2
<r2> 168.463
(<r2>)1/2 12.979