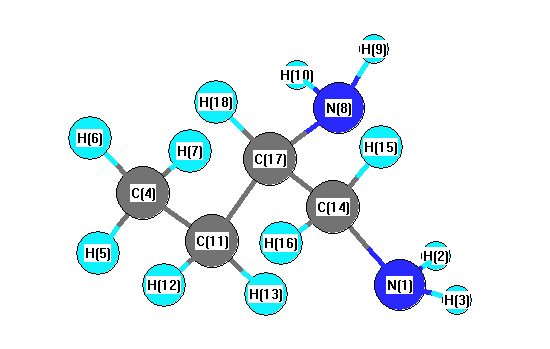
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3588 |
3433 |
1.74 |
|
|
|
| 2 |
A |
3585 |
3430 |
0.19 |
|
|
|
| 3 |
A |
3494 |
3343 |
1.63 |
|
|
|
| 4 |
A |
3493 |
3341 |
6.77 |
|
|
|
| 5 |
A |
3135 |
2999 |
39.98 |
|
|
|
| 6 |
A |
3126 |
2991 |
37.12 |
|
|
|
| 7 |
A |
3107 |
2972 |
14.92 |
|
|
|
| 8 |
A |
3071 |
2938 |
56.69 |
|
|
|
| 9 |
A |
3063 |
2931 |
7.38 |
|
|
|
| 10 |
A |
3046 |
2914 |
29.74 |
|
|
|
| 11 |
A |
2943 |
2816 |
124.98 |
|
|
|
| 12 |
A |
2921 |
2795 |
80.13 |
|
|
|
| 13 |
A |
1694 |
1621 |
40.76 |
|
|
|
| 14 |
A |
1675 |
1602 |
49.03 |
|
|
|
| 15 |
A |
1538 |
1471 |
4.42 |
|
|
|
| 16 |
A |
1534 |
1467 |
8.36 |
|
|
|
| 17 |
A |
1524 |
1458 |
4.77 |
|
|
|
| 18 |
A |
1504 |
1439 |
2.44 |
|
|
|
| 19 |
A |
1447 |
1384 |
1.85 |
|
|
|
| 20 |
A |
1443 |
1380 |
11.80 |
|
|
|
| 21 |
A |
1434 |
1372 |
2.27 |
|
|
|
| 22 |
A |
1404 |
1343 |
14.28 |
|
|
|
| 23 |
A |
1357 |
1298 |
1.76 |
|
|
|
| 24 |
A |
1317 |
1260 |
0.97 |
|
|
|
| 25 |
A |
1300 |
1244 |
2.31 |
|
|
|
| 26 |
A |
1275 |
1219 |
1.51 |
|
|
|
| 27 |
A |
1205 |
1153 |
5.45 |
|
|
|
| 28 |
A |
1168 |
1117 |
7.85 |
|
|
|
| 29 |
A |
1121 |
1073 |
2.07 |
|
|
|
| 30 |
A |
1085 |
1038 |
5.50 |
|
|
|
| 31 |
A |
1076 |
1029 |
2.00 |
|
|
|
| 32 |
A |
1024 |
980 |
1.14 |
|
|
|
| 33 |
A |
993 |
950 |
30.37 |
|
|
|
| 34 |
A |
939 |
898 |
41.19 |
|
|
|
| 35 |
A |
929 |
889 |
50.28 |
|
|
|
| 36 |
A |
879 |
841 |
149.10 |
|
|
|
| 37 |
A |
827 |
791 |
21.61 |
|
|
|
| 38 |
A |
771 |
738 |
0.94 |
|
|
|
| 39 |
A |
626 |
599 |
19.54 |
|
|
|
| 40 |
A |
446 |
427 |
6.02 |
|
|
|
| 41 |
A |
385 |
368 |
6.48 |
|
|
|
| 42 |
A |
345 |
330 |
47.13 |
|
|
|
| 43 |
A |
284 |
272 |
37.27 |
|
|
|
| 44 |
A |
278 |
266 |
23.99 |
|
|
|
| 45 |
A |
238 |
228 |
9.55 |
|
|
|
| 46 |
A |
215 |
205 |
3.22 |
|
|
|
| 47 |
A |
146 |
139 |
6.44 |
|
|
|
| 48 |
A |
85 |
81 |
0.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37039.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 35435.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.755 |
|
|
-0.975 |
| 2 |
H |
0.332 |
|
|
0.366 |
| 3 |
H |
0.310 |
|
|
0.379 |
| 4 |
C |
-0.526 |
|
|
-0.335 |
| 5 |
H |
0.173 |
|
|
0.083 |
| 6 |
H |
0.161 |
|
|
0.075 |
| 7 |
H |
0.160 |
|
|
0.083 |
| 8 |
N |
-0.765 |
|
|
-0.939 |
| 9 |
H |
0.311 |
|
|
0.361 |
| 10 |
H |
0.321 |
|
|
0.352 |
| 11 |
C |
-0.290 |
|
|
0.157 |
| 12 |
H |
0.157 |
|
|
-0.020 |
| 13 |
H |
0.193 |
|
|
0.023 |
| 14 |
C |
-0.172 |
|
|
0.222 |
| 15 |
H |
0.122 |
|
|
-0.023 |
| 16 |
H |
0.159 |
|
|
0.014 |
| 17 |
C |
-0.019 |
|
|
0.232 |
| 18 |
H |
0.129 |
|
|
-0.052 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.034 |
1.531 |
1.249 |
1.976 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.065 |
1.561 |
1.243 |
1.996 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.079 |
0.459 |
0.811 |
| y |
0.459 |
-38.121 |
-0.333 |
| z |
0.811 |
-0.333 |
-40.430 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.197 |
0.459 |
0.811 |
| y |
0.459 |
-0.366 |
-0.333 |
| z |
0.811 |
-0.333 |
-3.830 |
|
| Polar |
|---|
| 3z2-r2 | -7.661 |
|---|
| x2-y2 | 3.042 |
|---|
| xy | 0.459 |
|---|
| xz | 0.811 |
|---|
| yz | -0.333 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
205.129 |
| (<r2>)1/2 |
14.322 |