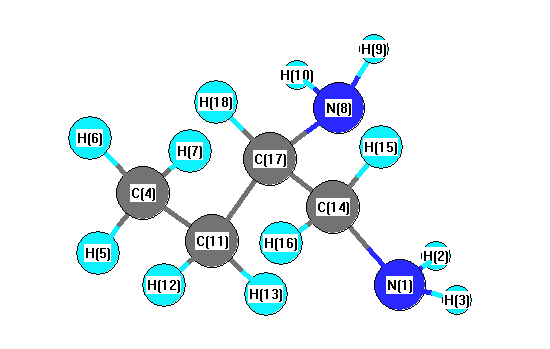
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3467 |
3410 |
0.35 |
|
|
|
| 2 |
A |
3466 |
3409 |
0.70 |
|
|
|
| 3 |
A |
3376 |
3320 |
4.22 |
|
|
|
| 4 |
A |
3368 |
3312 |
10.40 |
|
|
|
| 5 |
A |
3056 |
3005 |
37.29 |
|
|
|
| 6 |
A |
3052 |
3002 |
39.58 |
|
|
|
| 7 |
A |
3030 |
2980 |
9.05 |
|
|
|
| 8 |
A |
2990 |
2941 |
56.16 |
|
|
|
| 9 |
A |
2985 |
2936 |
7.05 |
|
|
|
| 10 |
A |
2972 |
2923 |
27.23 |
|
|
|
| 11 |
A |
2858 |
2811 |
116.68 |
|
|
|
| 12 |
A |
2832 |
2786 |
90.19 |
|
|
|
| 13 |
A |
1644 |
1617 |
34.32 |
|
|
|
| 14 |
A |
1626 |
1599 |
45.11 |
|
|
|
| 15 |
A |
1492 |
1467 |
3.62 |
|
|
|
| 16 |
A |
1491 |
1466 |
9.12 |
|
|
|
| 17 |
A |
1481 |
1457 |
4.40 |
|
|
|
| 18 |
A |
1458 |
1434 |
2.26 |
|
|
|
| 19 |
A |
1397 |
1374 |
2.44 |
|
|
|
| 20 |
A |
1395 |
1372 |
12.46 |
|
|
|
| 21 |
A |
1389 |
1366 |
1.23 |
|
|
|
| 22 |
A |
1355 |
1332 |
13.41 |
|
|
|
| 23 |
A |
1313 |
1292 |
1.77 |
|
|
|
| 24 |
A |
1277 |
1256 |
0.55 |
|
|
|
| 25 |
A |
1263 |
1242 |
2.61 |
|
|
|
| 26 |
A |
1237 |
1216 |
0.97 |
|
|
|
| 27 |
A |
1168 |
1149 |
4.43 |
|
|
|
| 28 |
A |
1131 |
1112 |
6.57 |
|
|
|
| 29 |
A |
1092 |
1074 |
1.97 |
|
|
|
| 30 |
A |
1054 |
1037 |
5.23 |
|
|
|
| 31 |
A |
1048 |
1031 |
1.66 |
|
|
|
| 32 |
A |
996 |
979 |
0.46 |
|
|
|
| 33 |
A |
962 |
946 |
27.17 |
|
|
|
| 34 |
A |
918 |
903 |
49.34 |
|
|
|
| 35 |
A |
907 |
892 |
38.87 |
|
|
|
| 36 |
A |
856 |
842 |
134.31 |
|
|
|
| 37 |
A |
806 |
793 |
17.94 |
|
|
|
| 38 |
A |
750 |
738 |
1.42 |
|
|
|
| 39 |
A |
610 |
600 |
18.29 |
|
|
|
| 40 |
A |
438 |
431 |
6.48 |
|
|
|
| 41 |
A |
376 |
370 |
7.24 |
|
|
|
| 42 |
A |
341 |
336 |
51.85 |
|
|
|
| 43 |
A |
281 |
277 |
25.11 |
|
|
|
| 44 |
A |
280 |
276 |
32.20 |
|
|
|
| 45 |
A |
239 |
235 |
7.93 |
|
|
|
| 46 |
A |
213 |
210 |
2.95 |
|
|
|
| 47 |
A |
145 |
143 |
6.22 |
|
|
|
| 48 |
A |
83 |
81 |
0.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35980.7 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 35387.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.705 |
|
|
|
| 2 |
H |
0.313 |
|
|
|
| 3 |
H |
0.292 |
|
|
|
| 4 |
C |
-0.501 |
|
|
|
| 5 |
H |
0.163 |
|
|
|
| 6 |
H |
0.152 |
|
|
|
| 7 |
H |
0.153 |
|
|
|
| 8 |
N |
-0.716 |
|
|
|
| 9 |
H |
0.295 |
|
|
|
| 10 |
H |
0.304 |
|
|
|
| 11 |
C |
-0.264 |
|
|
|
| 12 |
H |
0.145 |
|
|
|
| 13 |
H |
0.178 |
|
|
|
| 14 |
C |
-0.166 |
|
|
|
| 15 |
H |
0.110 |
|
|
|
| 16 |
H |
0.147 |
|
|
|
| 17 |
C |
-0.014 |
|
|
|
| 18 |
H |
0.114 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.007 |
1.543 |
1.204 |
1.957 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.411 |
0.515 |
0.766 |
| y |
0.515 |
-38.382 |
-0.332 |
| z |
0.766 |
-0.332 |
-40.764 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.162 |
0.515 |
0.766 |
| y |
0.515 |
-0.294 |
-0.332 |
| z |
0.766 |
-0.332 |
-3.868 |
|
| Polar |
|---|
| 3z2-r2 | -7.736 |
|---|
| x2-y2 | 2.971 |
|---|
| xy | 0.515 |
|---|
| xz | 0.766 |
|---|
| yz | -0.332 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.991 |
-0.020 |
0.118 |
| y |
-0.020 |
8.450 |
-0.070 |
| z |
0.118 |
-0.070 |
8.072 |
<r2> (average value of r
2) Å
2
| <r2> |
206.630 |
| (<r2>)1/2 |
14.375 |