Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -218.291804 |
| Energy at 298.15K | -218.299631 |
| HF Energy | -218.291804 |
| Nuclear repulsion energy | 127.954501 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3142 |
3006 |
27.71 |
|
|
|
| 2 |
A' |
3066 |
2933 |
40.06 |
|
|
|
| 3 |
A' |
3056 |
2924 |
21.41 |
|
|
|
| 4 |
A' |
3040 |
2909 |
33.54 |
|
|
|
| 5 |
A' |
1550 |
1483 |
2.11 |
|
|
|
| 6 |
A' |
1533 |
1467 |
5.58 |
|
|
|
| 7 |
A' |
1519 |
1454 |
0.42 |
|
|
|
| 8 |
A' |
1454 |
1391 |
20.24 |
|
|
|
| 9 |
A' |
1438 |
1375 |
1.76 |
|
|
|
| 10 |
A' |
1346 |
1287 |
1.83 |
|
|
|
| 11 |
A' |
1152 |
1102 |
3.77 |
|
|
|
| 12 |
A' |
1098 |
1051 |
94.38 |
|
|
|
| 13 |
A' |
1058 |
1012 |
4.20 |
|
|
|
| 14 |
A' |
911 |
872 |
9.43 |
|
|
|
| 15 |
A' |
452 |
432 |
5.80 |
|
|
|
| 16 |
A' |
265 |
254 |
2.88 |
|
|
|
| 17 |
A" |
3132 |
2996 |
71.74 |
|
|
|
| 18 |
A" |
3108 |
2973 |
11.54 |
|
|
|
| 19 |
A" |
3081 |
2948 |
22.37 |
|
|
|
| 20 |
A" |
1527 |
1461 |
8.25 |
|
|
|
| 21 |
A" |
1329 |
1272 |
0.25 |
|
|
|
| 22 |
A" |
1282 |
1226 |
0.26 |
|
|
|
| 23 |
A" |
1206 |
1154 |
1.13 |
|
|
|
| 24 |
A" |
903 |
864 |
2.10 |
|
|
|
| 25 |
A" |
772 |
738 |
2.38 |
|
|
|
| 26 |
A" |
235 |
225 |
0.00 |
|
|
|
| 27 |
A" |
131 |
126 |
2.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21392.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 20466.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.124 |
-0.788 |
0.000 |
| C2 |
0.000 |
0.723 |
0.000 |
| C3 |
-1.456 |
1.184 |
0.000 |
| F4 |
1.461 |
-1.157 |
0.000 |
| H5 |
-0.351 |
-1.223 |
0.890 |
| H6 |
-0.351 |
-1.223 |
-0.890 |
| H7 |
0.524 |
1.115 |
-0.880 |
| H8 |
0.524 |
1.115 |
0.880 |
| H9 |
-1.521 |
2.276 |
0.000 |
| H10 |
-1.992 |
0.821 |
-0.885 |
| H11 |
-1.992 |
0.821 |
0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5160 | 2.5268 | 1.3867 | 1.0986 | 1.0986 | 2.1346 | 2.1346 | 3.4780 | 2.8019 | 2.8019 |
C2 | 1.5160 | | 1.5271 | 2.3811 | 2.1680 | 2.1680 | 1.0971 | 1.0971 | 2.1739 | 2.1821 | 2.1821 | C3 | 2.5268 | 1.5271 | | 3.7401 | 2.7932 | 2.7932 | 2.1682 | 2.1682 | 1.0947 | 1.0968 | 1.0968 | F4 | 1.3867 | 2.3811 | 3.7401 | | 2.0201 | 2.0201 | 2.6107 | 2.6107 | 4.5476 | 4.0767 | 4.0767 | H5 | 1.0986 | 2.1680 | 2.7932 | 2.0201 | | 1.7795 | 3.0602 | 2.4964 | 3.7950 | 3.1650 | 2.6204 | H6 | 1.0986 | 2.1680 | 2.7932 | 2.0201 | 1.7795 | | 2.4964 | 3.0602 | 3.7950 | 2.6204 | 3.1650 | H7 | 2.1346 | 1.0971 | 2.1682 | 2.6107 | 3.0602 | 2.4964 | | 1.7606 | 2.5111 | 2.5336 | 3.0881 | H8 | 2.1346 | 1.0971 | 2.1682 | 2.6107 | 2.4964 | 3.0602 | 1.7606 | | 2.5111 | 3.0881 | 2.5336 | H9 | 3.4780 | 2.1739 | 1.0947 | 4.5476 | 3.7950 | 3.7950 | 2.5111 | 2.5111 | | 1.7679 | 1.7679 | H10 | 2.8019 | 2.1821 | 1.0968 | 4.0767 | 3.1650 | 2.6204 | 2.5336 | 3.0881 | 1.7679 | | 1.7705 | H11 | 2.8019 | 2.1821 | 1.0968 | 4.0767 | 2.6204 | 3.1650 | 3.0881 | 2.5336 | 1.7679 | 1.7705 | |
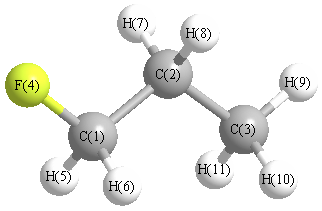 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.263 |
|
C1 |
C2 |
H7 |
108.485 |
| C1 |
C2 |
H8 |
108.485 |
|
C2 |
C1 |
F4 |
110.149 |
| C2 |
C1 |
H5 |
111.025 |
|
C2 |
C1 |
H6 |
111.025 |
| C2 |
C3 |
H9 |
110.947 |
|
C2 |
C3 |
H10 |
111.480 |
| C2 |
C3 |
H11 |
111.480 |
|
C3 |
C2 |
H7 |
110.354 |
| C3 |
C2 |
H8 |
110.354 |
|
F4 |
C1 |
H5 |
108.183 |
| F4 |
C1 |
H6 |
108.183 |
|
H5 |
C1 |
H6 |
108.173 |
| H7 |
C2 |
H8 |
106.717 |
|
H9 |
C3 |
H10 |
107.552 |
| H9 |
C3 |
H11 |
107.552 |
|
H10 |
C3 |
H11 |
107.635 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.025 |
|
|
|
| 2 |
C |
-0.351 |
|
|
|
| 3 |
C |
-0.526 |
|
|
|
| 4 |
F |
-0.318 |
|
|
|
| 5 |
H |
0.148 |
|
|
|
| 6 |
H |
0.148 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.177 |
|
|
|
| 9 |
H |
0.180 |
|
|
|
| 10 |
H |
0.171 |
|
|
|
| 11 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.608 |
0.736 |
0.000 |
1.768 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.541 |
2.008 |
0.000 |
| y |
2.008 |
-25.458 |
0.000 |
| z |
0.000 |
0.000 |
-24.314 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.655 |
2.008 |
0.000 |
| y |
2.008 |
0.469 |
0.000 |
| z |
0.000 |
0.000 |
2.186 |
|
| Polar |
|---|
| 3z2-r2 | 4.372 |
|---|
| x2-y2 | -2.083 |
|---|
| xy | 2.008 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.100 |
-0.230 |
0.000 |
| y |
-0.230 |
5.104 |
0.000 |
| z |
0.000 |
0.000 |
4.744 |
<r2> (average value of r
2) Å
2
| <r2> |
103.191 |
| (<r2>)1/2 |
10.158 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -218.292211 |
| Energy at 298.15K | |
| HF Energy | -218.292211 |
| Nuclear repulsion energy | 130.403271 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3153 |
3017 |
23.78 |
|
|
|
| 2 |
A |
3135 |
2999 |
43.66 |
|
|
|
| 3 |
A |
3103 |
2968 |
49.62 |
|
|
|
| 4 |
A |
3091 |
2957 |
31.14 |
|
|
|
| 5 |
A |
3061 |
2928 |
20.64 |
|
|
|
| 6 |
A |
3055 |
2923 |
22.45 |
|
|
|
| 7 |
A |
3044 |
2913 |
41.39 |
|
|
|
| 8 |
A |
1544 |
1477 |
0.68 |
|
|
|
| 9 |
A |
1535 |
1468 |
8.55 |
|
|
|
| 10 |
A |
1521 |
1455 |
7.53 |
|
|
|
| 11 |
A |
1503 |
1437 |
2.31 |
|
|
|
| 12 |
A |
1451 |
1388 |
15.88 |
|
|
|
| 13 |
A |
1432 |
1370 |
6.68 |
|
|
|
| 14 |
A |
1396 |
1336 |
0.67 |
|
|
|
| 15 |
A |
1317 |
1260 |
1.10 |
|
|
|
| 16 |
A |
1285 |
1230 |
0.83 |
|
|
|
| 17 |
A |
1191 |
1139 |
1.37 |
|
|
|
| 18 |
A |
1140 |
1090 |
19.58 |
|
|
|
| 19 |
A |
1109 |
1061 |
41.76 |
|
|
|
| 20 |
A |
1002 |
959 |
36.85 |
|
|
|
| 21 |
A |
931 |
891 |
3.43 |
|
|
|
| 22 |
A |
889 |
850 |
3.70 |
|
|
|
| 23 |
A |
778 |
744 |
0.64 |
|
|
|
| 24 |
A |
481 |
460 |
3.09 |
|
|
|
| 25 |
A |
316 |
302 |
1.15 |
|
|
|
| 26 |
A |
221 |
212 |
1.77 |
|
|
|
| 27 |
A |
145 |
139 |
2.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21413.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 20486.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.803 |
0.512 |
0.292 |
| C2 |
-0.594 |
0.661 |
-0.278 |
| C3 |
-1.523 |
-0.484 |
0.118 |
| F4 |
1.385 |
-0.662 |
-0.170 |
| H5 |
1.446 |
1.349 |
-0.009 |
| H6 |
0.779 |
0.467 |
1.389 |
| H7 |
-0.520 |
0.724 |
-1.371 |
| H8 |
-1.000 |
1.622 |
0.066 |
| H9 |
-2.514 |
-0.366 |
-0.333 |
| H10 |
-1.111 |
-1.443 |
-0.211 |
| H11 |
-1.654 |
-0.531 |
1.206 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5160 | 2.5362 | 1.3895 | 1.0976 | 1.0986 | 2.1349 | 2.1291 | 3.4873 | 2.7814 | 2.8210 |
C2 | 1.5160 | | 1.5262 | 2.3829 | 2.1704 | 2.1683 | 1.0976 | 1.0988 | 2.1773 | 2.1667 | 2.1780 | C3 | 2.5362 | 1.5262 | | 2.9274 | 3.4919 | 2.7963 | 2.1641 | 2.1708 | 1.0951 | 1.0942 | 1.0969 | F4 | 1.3895 | 2.3829 | 2.9274 | | 2.0188 | 2.0178 | 2.6443 | 3.3107 | 3.9131 | 2.6150 | 3.3377 | H5 | 1.0976 | 2.1704 | 3.4919 | 2.0188 | | 1.7823 | 2.4721 | 2.4626 | 4.3276 | 3.7913 | 3.8235 | H6 | 1.0986 | 2.1683 | 2.7963 | 2.0178 | 1.7823 | | 3.0611 | 2.5000 | 3.8082 | 3.1272 | 2.6357 | H7 | 2.1349 | 1.0976 | 2.1641 | 2.6443 | 2.4721 | 3.0611 | | 1.7611 | 2.4982 | 2.5276 | 3.0826 | H8 | 2.1291 | 1.0988 | 2.1708 | 3.3107 | 2.4626 | 2.5000 | 1.7611 | | 2.5304 | 3.0793 | 2.5227 | H9 | 3.4873 | 2.1773 | 1.0951 | 3.9131 | 4.3276 | 3.8082 | 2.4982 | 2.5304 | | 1.7727 | 1.7713 | H10 | 2.7814 | 2.1667 | 1.0942 | 2.6150 | 3.7913 | 3.1272 | 2.5276 | 3.0793 | 1.7727 | | 1.7705 | H11 | 2.8210 | 2.1780 | 1.0969 | 3.3377 | 3.8235 | 2.6357 | 3.0826 | 2.5227 | 1.7713 | 1.7705 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.956 |
|
C1 |
C2 |
H7 |
108.485 |
| C1 |
C2 |
H8 |
107.969 |
|
C2 |
C1 |
F4 |
110.123 |
| C2 |
C1 |
H5 |
111.279 |
|
C2 |
C1 |
H6 |
111.054 |
| C2 |
C3 |
H9 |
111.266 |
|
C2 |
C3 |
H10 |
110.468 |
| C2 |
C3 |
H11 |
111.209 |
|
C3 |
C2 |
H7 |
110.068 |
| C3 |
C2 |
H8 |
110.521 |
|
F4 |
C1 |
H5 |
107.956 |
| F4 |
C1 |
H6 |
107.809 |
|
H5 |
C1 |
H6 |
108.497 |
| H7 |
C2 |
H8 |
106.609 |
|
H9 |
C3 |
H10 |
108.137 |
| H9 |
C3 |
H11 |
107.813 |
|
H10 |
C3 |
H11 |
107.809 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.017 |
|
|
|
| 2 |
C |
-0.349 |
|
|
|
| 3 |
C |
-0.512 |
|
|
|
| 4 |
F |
-0.317 |
|
|
|
| 5 |
H |
0.152 |
|
|
|
| 6 |
H |
0.148 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.162 |
|
|
|
| 9 |
H |
0.171 |
|
|
|
| 10 |
H |
0.188 |
|
|
|
| 11 |
H |
0.165 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.872 |
1.339 |
0.414 |
1.651 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.384 |
1.584 |
0.584 |
| y |
1.584 |
-24.954 |
0.051 |
| z |
0.584 |
0.051 |
-24.419 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.698 |
1.584 |
0.584 |
| y |
1.584 |
0.447 |
0.051 |
| z |
0.584 |
0.051 |
1.251 |
|
| Polar |
|---|
| 3z2-r2 | 2.501 |
|---|
| x2-y2 | -1.430 |
|---|
| xy | 1.584 |
|---|
| xz | 0.584 |
|---|
| yz | 0.051 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.256 |
0.223 |
0.036 |
| y |
0.223 |
4.903 |
-0.023 |
| z |
0.036 |
-0.023 |
4.795 |
<r2> (average value of r
2) Å
2
| <r2> |
91.165 |
| (<r2>)1/2 |
9.548 |
