Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -217.717696 |
| Energy at 298.15K | |
| HF Energy | -217.110882 |
| Nuclear repulsion energy | 127.709452 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -217.718589 |
| Energy at 298.15K | |
| HF Energy | -217.111254 |
| Nuclear repulsion energy | 130.295565 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3163 |
3012 |
23.54 |
|
|
|
| 2 |
A |
3141 |
2991 |
47.45 |
|
|
|
| 3 |
A |
3124 |
2975 |
49.61 |
|
|
|
| 4 |
A |
3106 |
2958 |
16.36 |
|
|
|
| 5 |
A |
3072 |
2926 |
23.70 |
|
|
|
| 6 |
A |
3069 |
2922 |
33.29 |
|
|
|
| 7 |
A |
3062 |
2916 |
19.42 |
|
|
|
| 8 |
A |
1573 |
1498 |
0.31 |
|
|
|
| 9 |
A |
1559 |
1485 |
7.92 |
|
|
|
| 10 |
A |
1547 |
1473 |
6.45 |
|
|
|
| 11 |
A |
1534 |
1461 |
1.47 |
|
|
|
| 12 |
A |
1475 |
1404 |
10.29 |
|
|
|
| 13 |
A |
1463 |
1393 |
10.66 |
|
|
|
| 14 |
A |
1424 |
1356 |
1.23 |
|
|
|
| 15 |
A |
1336 |
1272 |
0.71 |
|
|
|
| 16 |
A |
1299 |
1237 |
0.68 |
|
|
|
| 17 |
A |
1216 |
1158 |
1.86 |
|
|
|
| 18 |
A |
1158 |
1103 |
11.50 |
|
|
|
| 19 |
A |
1127 |
1073 |
38.49 |
|
|
|
| 20 |
A |
1012 |
963 |
39.24 |
|
|
|
| 21 |
A |
947 |
902 |
3.27 |
|
|
|
| 22 |
A |
905 |
862 |
3.86 |
|
|
|
| 23 |
A |
792 |
755 |
0.47 |
|
|
|
| 24 |
A |
491 |
468 |
3.76 |
|
|
|
| 25 |
A |
324 |
309 |
1.38 |
|
|
|
| 26 |
A |
236 |
225 |
2.20 |
|
|
|
| 27 |
A |
149 |
142 |
2.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21650.9 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 20618.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.801 |
0.523 |
0.294 |
| C2 |
-0.595 |
0.667 |
-0.281 |
| C3 |
-1.512 |
-0.489 |
0.120 |
| F4 |
1.373 |
-0.670 |
-0.170 |
| H5 |
1.456 |
1.348 |
-0.014 |
| H6 |
0.780 |
0.475 |
1.391 |
| H7 |
-0.515 |
0.724 |
-1.375 |
| H8 |
-1.010 |
1.627 |
0.060 |
| H9 |
-2.508 |
-0.377 |
-0.324 |
| H10 |
-1.090 |
-1.442 |
-0.215 |
| H11 |
-1.631 |
-0.535 |
1.211 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5164 | 2.5305 | 1.4018 | 1.0976 | 1.0985 | 2.1349 | 2.1334 | 3.4841 | 2.7744 | 2.8064 |
C2 | 1.5164 | | 1.5288 | 2.3814 | 2.1776 | 2.1729 | 1.0985 | 1.1001 | 2.1793 | 2.1673 | 2.1779 | C3 | 2.5305 | 1.5288 | | 2.9051 | 3.4930 | 2.7918 | 2.1681 | 2.1751 | 1.0964 | 1.0950 | 1.0981 | F4 | 1.4018 | 2.3814 | 2.9051 | | 2.0257 | 2.0246 | 2.6382 | 3.3173 | 3.8950 | 2.5819 | 3.3092 | H5 | 1.0976 | 2.1776 | 3.4930 | 2.0257 | | 1.7873 | 2.4754 | 2.4826 | 4.3339 | 3.7829 | 3.8180 | H6 | 1.0985 | 2.1729 | 2.7918 | 2.0246 | 1.7873 | | 3.0643 | 2.5100 | 3.8044 | 3.1225 | 2.6199 | H7 | 2.1349 | 1.0985 | 2.1681 | 2.6382 | 2.4754 | 3.0643 | | 1.7662 | 2.5072 | 2.5238 | 3.0850 | H8 | 2.1334 | 1.1001 | 2.1751 | 3.3173 | 2.4826 | 2.5100 | 1.7662 | | 2.5307 | 3.0823 | 2.5263 | H9 | 3.4841 | 2.1793 | 1.0964 | 3.8950 | 4.3339 | 3.8044 | 2.5072 | 2.5307 | | 1.7770 | 1.7748 | H10 | 2.7744 | 2.1673 | 1.0950 | 2.5819 | 3.7829 | 3.1225 | 2.5238 | 3.0823 | 1.7770 | | 1.7745 | H11 | 2.8064 | 2.1779 | 1.0981 | 3.3092 | 3.8180 | 2.6199 | 3.0850 | 2.5263 | 1.7748 | 1.7745 | |
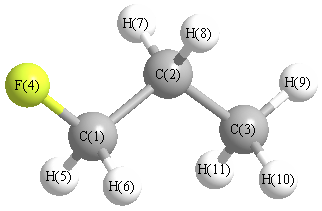 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.398 |
|
C1 |
C2 |
H7 |
108.400 |
| C1 |
C2 |
H8 |
108.199 |
|
C2 |
C1 |
F4 |
109.320 |
| C2 |
C1 |
H5 |
111.828 |
|
C2 |
C1 |
H6 |
111.398 |
| C2 |
C3 |
H9 |
111.162 |
|
C2 |
C3 |
H10 |
110.290 |
| C2 |
C3 |
H11 |
110.947 |
|
C3 |
C2 |
H7 |
110.143 |
| C3 |
C2 |
H8 |
110.608 |
|
F4 |
C1 |
H5 |
107.665 |
| F4 |
C1 |
H6 |
107.521 |
|
H5 |
C1 |
H6 |
108.947 |
| H7 |
C2 |
H8 |
106.904 |
|
H9 |
C3 |
H10 |
108.361 |
| H9 |
C3 |
H11 |
107.948 |
|
H10 |
C3 |
H11 |
108.021 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
