Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -218.128955 |
| Energy at 298.15K | -218.136821 |
| HF Energy | -217.944885 |
| Nuclear repulsion energy | 127.878975 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3163 |
3003 |
28.40 |
|
|
|
| 2 |
A' |
3091 |
2934 |
39.92 |
|
|
|
| 3 |
A' |
3081 |
2924 |
24.94 |
|
|
|
| 4 |
A' |
3074 |
2918 |
22.50 |
|
|
|
| 5 |
A' |
1576 |
1496 |
1.40 |
|
|
|
| 6 |
A' |
1559 |
1480 |
4.75 |
|
|
|
| 7 |
A' |
1545 |
1466 |
0.48 |
|
|
|
| 8 |
A' |
1475 |
1401 |
19.93 |
|
|
|
| 9 |
A' |
1463 |
1388 |
1.06 |
|
|
|
| 10 |
A' |
1367 |
1297 |
2.87 |
|
|
|
| 11 |
A' |
1165 |
1106 |
1.82 |
|
|
|
| 12 |
A' |
1093 |
1038 |
83.17 |
|
|
|
| 13 |
A' |
1061 |
1007 |
13.62 |
|
|
|
| 14 |
A' |
920 |
874 |
8.38 |
|
|
|
| 15 |
A' |
454 |
431 |
5.93 |
|
|
|
| 16 |
A' |
270 |
256 |
3.26 |
|
|
|
| 17 |
A" |
3155 |
2995 |
80.81 |
|
|
|
| 18 |
A" |
3134 |
2975 |
10.30 |
|
|
|
| 19 |
A" |
3115 |
2957 |
12.68 |
|
|
|
| 20 |
A" |
1553 |
1474 |
6.99 |
|
|
|
| 21 |
A" |
1347 |
1279 |
0.22 |
|
|
|
| 22 |
A" |
1299 |
1233 |
0.15 |
|
|
|
| 23 |
A" |
1225 |
1163 |
1.51 |
|
|
|
| 24 |
A" |
916 |
870 |
1.92 |
|
|
|
| 25 |
A" |
783 |
743 |
1.54 |
|
|
|
| 26 |
A" |
239 |
227 |
0.00 |
|
|
|
| 27 |
A" |
135 |
128 |
2.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21629.5 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 20530.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.120 |
-0.789 |
0.000 |
| C2 |
0.000 |
0.723 |
0.000 |
| C3 |
-1.458 |
1.184 |
0.000 |
| F4 |
1.465 |
-1.155 |
0.000 |
| H5 |
-0.350 |
-1.223 |
0.889 |
| H6 |
-0.350 |
-1.223 |
-0.889 |
| H7 |
0.524 |
1.111 |
-0.879 |
| H8 |
0.524 |
1.111 |
0.879 |
| H9 |
-1.522 |
2.275 |
0.000 |
| H10 |
-1.991 |
0.820 |
-0.884 |
| H11 |
-1.991 |
0.820 |
0.884 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5160 | 2.5260 | 1.3940 | 1.0956 | 1.0956 | 2.1320 | 2.1320 | 3.4756 | 2.7971 | 2.7971 |
C2 | 1.5160 | | 1.5292 | 2.3816 | 2.1679 | 2.1679 | 1.0947 | 1.0947 | 2.1737 | 2.1804 | 2.1804 | C3 | 2.5260 | 1.5292 | | 3.7436 | 2.7951 | 2.7951 | 2.1692 | 2.1692 | 1.0926 | 1.0945 | 1.0945 | F4 | 1.3940 | 2.3816 | 3.7436 | | 2.0223 | 2.0223 | 2.6068 | 2.6068 | 4.5481 | 4.0772 | 4.0772 | H5 | 1.0956 | 2.1679 | 2.7951 | 2.0223 | | 1.7778 | 3.0562 | 2.4927 | 3.7946 | 3.1637 | 2.6203 | H6 | 1.0956 | 2.1679 | 2.7951 | 2.0223 | 1.7778 | | 2.4927 | 3.0562 | 3.7946 | 2.6203 | 3.1637 | H7 | 2.1320 | 1.0947 | 2.1692 | 2.6068 | 3.0562 | 2.4927 | | 1.7587 | 2.5123 | 2.5312 | 3.0849 | H8 | 2.1320 | 1.0947 | 2.1692 | 2.6068 | 2.4927 | 3.0562 | 1.7587 | | 2.5123 | 3.0849 | 2.5312 | H9 | 3.4756 | 2.1737 | 1.0926 | 4.5481 | 3.7946 | 3.7946 | 2.5123 | 2.5123 | | 1.7657 | 1.7657 | H10 | 2.7971 | 2.1804 | 1.0945 | 4.0772 | 3.1637 | 2.6203 | 2.5312 | 3.0849 | 1.7657 | | 1.7682 | H11 | 2.7971 | 2.1804 | 1.0945 | 4.0772 | 2.6203 | 3.1637 | 3.0849 | 2.5312 | 1.7657 | 1.7682 | |
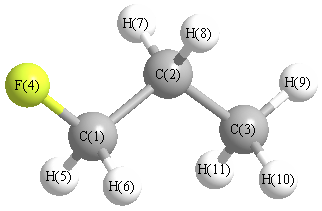 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.096 |
|
C1 |
C2 |
H7 |
108.415 |
| C1 |
C2 |
H8 |
108.415 |
|
C2 |
C1 |
F4 |
109.784 |
| C2 |
C1 |
H5 |
111.200 |
|
C2 |
C1 |
H6 |
111.200 |
| C2 |
C3 |
H9 |
110.918 |
|
C2 |
C3 |
H10 |
111.327 |
| C2 |
C3 |
H11 |
111.327 |
|
C3 |
C2 |
H7 |
110.427 |
| C3 |
C2 |
H8 |
110.427 |
|
F4 |
C1 |
H5 |
108.044 |
| F4 |
C1 |
H6 |
108.044 |
|
H5 |
C1 |
H6 |
108.458 |
| H7 |
C2 |
H8 |
106.888 |
|
H9 |
C3 |
H10 |
107.672 |
| H9 |
C3 |
H11 |
107.672 |
|
H10 |
C3 |
H11 |
107.749 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.066 |
|
|
|
| 2 |
C |
-0.309 |
|
|
|
| 3 |
C |
-0.475 |
|
|
|
| 4 |
F |
-0.337 |
|
|
|
| 5 |
H |
0.134 |
|
|
|
| 6 |
H |
0.134 |
|
|
|
| 7 |
H |
0.159 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
H |
0.163 |
|
|
|
| 10 |
H |
0.153 |
|
|
|
| 11 |
H |
0.153 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.665 |
0.742 |
0.000 |
1.823 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.947 |
2.082 |
0.000 |
| y |
2.082 |
-25.782 |
0.000 |
| z |
0.000 |
0.000 |
-24.591 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.760 |
2.082 |
0.000 |
| y |
2.082 |
0.487 |
0.000 |
| z |
0.000 |
0.000 |
2.273 |
|
| Polar |
|---|
| 3z2-r2 | 4.547 |
|---|
| x2-y2 | -2.165 |
|---|
| xy | 2.082 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.024 |
-0.225 |
0.000 |
| y |
-0.225 |
5.004 |
0.000 |
| z |
0.000 |
0.000 |
4.679 |
<r2> (average value of r
2) Å
2
| <r2> |
103.452 |
| (<r2>)1/2 |
10.171 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -218.129699 |
| Energy at 298.15K | |
| HF Energy | -217.945404 |
| Nuclear repulsion energy | 130.459774 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3177 |
3015 |
23.70 |
|
|
|
| 2 |
A |
3156 |
2996 |
46.48 |
|
|
|
| 3 |
A |
3134 |
2975 |
55.74 |
|
|
|
| 4 |
A |
3118 |
2960 |
18.08 |
|
|
|
| 5 |
A |
3085 |
2929 |
14.81 |
|
|
|
| 6 |
A |
3082 |
2925 |
41.85 |
|
|
|
| 7 |
A |
3075 |
2919 |
23.31 |
|
|
|
| 8 |
A |
1570 |
1490 |
0.37 |
|
|
|
| 9 |
A |
1561 |
1481 |
7.18 |
|
|
|
| 10 |
A |
1547 |
1468 |
6.34 |
|
|
|
| 11 |
A |
1529 |
1452 |
1.61 |
|
|
|
| 12 |
A |
1472 |
1397 |
13.83 |
|
|
|
| 13 |
A |
1454 |
1380 |
8.33 |
|
|
|
| 14 |
A |
1419 |
1347 |
1.05 |
|
|
|
| 15 |
A |
1334 |
1266 |
0.77 |
|
|
|
| 16 |
A |
1298 |
1232 |
0.74 |
|
|
|
| 17 |
A |
1210 |
1148 |
1.74 |
|
|
|
| 18 |
A |
1150 |
1092 |
11.28 |
|
|
|
| 19 |
A |
1111 |
1054 |
41.11 |
|
|
|
| 20 |
A |
1005 |
954 |
40.48 |
|
|
|
| 21 |
A |
944 |
896 |
3.64 |
|
|
|
| 22 |
A |
895 |
850 |
3.68 |
|
|
|
| 23 |
A |
790 |
749 |
0.48 |
|
|
|
| 24 |
A |
488 |
463 |
3.40 |
|
|
|
| 25 |
A |
321 |
305 |
1.25 |
|
|
|
| 26 |
A |
231 |
219 |
2.00 |
|
|
|
| 27 |
A |
148 |
141 |
2.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21652.1 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 20552.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.802 |
0.520 |
0.292 |
| C2 |
-0.595 |
0.666 |
-0.279 |
| C3 |
-1.514 |
-0.488 |
0.119 |
| F4 |
1.375 |
-0.668 |
-0.170 |
| H5 |
1.451 |
1.346 |
-0.014 |
| H6 |
0.780 |
0.475 |
1.386 |
| H7 |
-0.519 |
0.727 |
-1.370 |
| H8 |
-1.006 |
1.623 |
0.065 |
| H9 |
-2.505 |
-0.378 |
-0.329 |
| H10 |
-1.092 |
-1.439 |
-0.211 |
| H11 |
-1.639 |
-0.534 |
1.206 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5161 | 2.5315 | 1.3971 | 1.0946 | 1.0956 | 2.1327 | 2.1298 | 3.4819 | 2.7708 | 2.8112 |
C2 | 1.5161 | | 1.5280 | 2.3812 | 2.1723 | 2.1683 | 1.0952 | 1.0966 | 2.1769 | 2.1641 | 2.1759 | C3 | 2.5315 | 1.5280 | | 2.9085 | 3.4890 | 2.7922 | 2.1643 | 2.1718 | 1.0929 | 1.0919 | 1.0947 | F4 | 1.3971 | 2.3812 | 2.9085 | | 2.0214 | 2.0201 | 2.6401 | 3.3119 | 3.8933 | 2.5846 | 3.3153 | H5 | 1.0946 | 2.1723 | 3.4890 | 2.0214 | | 1.7807 | 2.4700 | 2.4737 | 4.3265 | 3.7767 | 3.8173 | H6 | 1.0956 | 2.1683 | 2.7922 | 2.0201 | 1.7807 | | 3.0574 | 2.5009 | 3.8026 | 3.1180 | 2.6273 | H7 | 2.1327 | 1.0952 | 2.1643 | 2.6401 | 2.4700 | 3.0574 | | 1.7601 | 2.4996 | 2.5225 | 3.0788 | H8 | 2.1298 | 1.0966 | 2.1718 | 3.3119 | 2.4737 | 2.5009 | 1.7601 | | 2.5306 | 3.0757 | 2.5209 | H9 | 3.4819 | 2.1769 | 1.0929 | 3.8933 | 4.3265 | 3.8026 | 2.4996 | 2.5306 | | 1.7710 | 1.7690 | H10 | 2.7708 | 2.1641 | 1.0919 | 2.5846 | 3.7767 | 3.1180 | 2.5225 | 3.0757 | 1.7710 | | 1.7683 | H11 | 2.8112 | 2.1759 | 1.0947 | 3.3153 | 3.8173 | 2.6273 | 3.0788 | 2.5209 | 1.7690 | 1.7683 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.527 |
|
C1 |
C2 |
H7 |
108.441 |
| C1 |
C2 |
H8 |
108.137 |
|
C2 |
C1 |
F4 |
109.587 |
| C2 |
C1 |
H5 |
111.612 |
|
C2 |
C1 |
H6 |
111.225 |
| C2 |
C3 |
H9 |
111.231 |
|
C2 |
C3 |
H10 |
110.279 |
| C2 |
C3 |
H11 |
111.040 |
|
C3 |
C2 |
H7 |
110.090 |
| C3 |
C2 |
H8 |
110.607 |
|
F4 |
C1 |
H5 |
107.822 |
| F4 |
C1 |
H6 |
107.663 |
|
H5 |
C1 |
H6 |
108.793 |
| H7 |
C2 |
H8 |
106.843 |
|
H9 |
C3 |
H10 |
108.312 |
| H9 |
C3 |
H11 |
107.921 |
|
H10 |
C3 |
H11 |
107.940 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.059 |
|
|
|
| 2 |
C |
-0.308 |
|
|
|
| 3 |
C |
-0.461 |
|
|
|
| 4 |
F |
-0.336 |
|
|
|
| 5 |
H |
0.139 |
|
|
|
| 6 |
H |
0.134 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
| 8 |
H |
0.143 |
|
|
|
| 9 |
H |
0.153 |
|
|
|
| 10 |
H |
0.171 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.871 |
1.392 |
0.432 |
1.698 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.711 |
1.630 |
0.595 |
| y |
1.630 |
-25.267 |
0.071 |
| z |
0.595 |
0.071 |
-24.702 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.726 |
1.630 |
0.595 |
| y |
1.630 |
0.440 |
0.071 |
| z |
0.595 |
0.071 |
1.287 |
|
| Polar |
|---|
| 3z2-r2 | 2.574 |
|---|
| x2-y2 | -1.444 |
|---|
| xy | 1.630 |
|---|
| xz | 0.595 |
|---|
| yz | 0.071 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.151 |
0.201 |
0.026 |
| y |
0.201 |
4.831 |
-0.013 |
| z |
0.026 |
-0.013 |
4.724 |
<r2> (average value of r
2) Å
2
| <r2> |
91.008 |
| (<r2>)1/2 |
9.540 |
