Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -218.095636 |
| Energy at 298.15K | -218.103382 |
| HF Energy | -218.095636 |
| Nuclear repulsion energy | 127.198176 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3068 |
3018 |
25.56 |
|
|
|
| 2 |
A' |
2989 |
2940 |
43.75 |
|
|
|
| 3 |
A' |
2983 |
2934 |
11.39 |
|
|
|
| 4 |
A' |
2947 |
2899 |
42.04 |
|
|
|
| 5 |
A' |
1504 |
1480 |
2.41 |
|
|
|
| 6 |
A' |
1489 |
1464 |
4.56 |
|
|
|
| 7 |
A' |
1475 |
1451 |
0.45 |
|
|
|
| 8 |
A' |
1408 |
1385 |
20.09 |
|
|
|
| 9 |
A' |
1391 |
1368 |
0.77 |
|
|
|
| 10 |
A' |
1303 |
1282 |
2.01 |
|
|
|
| 11 |
A' |
1118 |
1100 |
2.51 |
|
|
|
| 12 |
A' |
1059 |
1041 |
88.05 |
|
|
|
| 13 |
A' |
1033 |
1016 |
6.27 |
|
|
|
| 14 |
A' |
886 |
871 |
9.58 |
|
|
|
| 15 |
A' |
440 |
433 |
5.39 |
|
|
|
| 16 |
A' |
257 |
253 |
2.46 |
|
|
|
| 17 |
A" |
3056 |
3006 |
59.57 |
|
|
|
| 18 |
A" |
3029 |
2979 |
11.82 |
|
|
|
| 19 |
A" |
2986 |
2937 |
33.84 |
|
|
|
| 20 |
A" |
1484 |
1459 |
7.99 |
|
|
|
| 21 |
A" |
1290 |
1269 |
0.31 |
|
|
|
| 22 |
A" |
1242 |
1221 |
0.10 |
|
|
|
| 23 |
A" |
1170 |
1150 |
0.80 |
|
|
|
| 24 |
A" |
878 |
864 |
2.13 |
|
|
|
| 25 |
A" |
754 |
742 |
2.60 |
|
|
|
| 26 |
A" |
230 |
226 |
0.00 |
|
|
|
| 27 |
A" |
133 |
131 |
2.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20801.3 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 20458.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.123 |
-0.791 |
0.000 |
| C2 |
0.000 |
0.727 |
0.000 |
| C3 |
-1.461 |
1.191 |
0.000 |
| F4 |
1.469 |
-1.166 |
0.000 |
| H5 |
-0.361 |
-1.226 |
0.897 |
| H6 |
-0.361 |
-1.226 |
-0.897 |
| H7 |
0.529 |
1.121 |
-0.887 |
| H8 |
0.529 |
1.121 |
0.887 |
| H9 |
-1.526 |
2.291 |
0.000 |
| H10 |
-2.001 |
0.825 |
-0.891 |
| H11 |
-2.001 |
0.825 |
0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5227 | 2.5373 | 1.3970 | 1.1084 | 1.1084 | 2.1459 | 2.1459 | 3.4959 | 2.8142 | 2.8142 |
C2 | 1.5227 | | 1.5331 | 2.3956 | 2.1791 | 2.1791 | 1.1053 | 1.1053 | 2.1858 | 2.1931 | 2.1931 | C3 | 2.5373 | 1.5331 | | 3.7602 | 2.8027 | 2.8027 | 2.1802 | 2.1802 | 1.1024 | 1.1046 | 1.1046 | F4 | 1.3970 | 2.3956 | 3.7602 | | 2.0392 | 2.0392 | 2.6260 | 2.6260 | 4.5741 | 4.0988 | 4.0988 | H5 | 1.1084 | 2.1791 | 2.8027 | 2.0392 | | 1.7936 | 3.0792 | 2.5100 | 3.8123 | 3.1770 | 2.6261 | H6 | 1.1084 | 2.1791 | 2.8027 | 2.0392 | 1.7936 | | 2.5100 | 3.0792 | 3.8123 | 2.6261 | 3.1770 | H7 | 2.1459 | 1.1053 | 2.1802 | 2.6260 | 3.0792 | 2.5100 | | 1.7736 | 2.5264 | 2.5479 | 3.1070 | H8 | 2.1459 | 1.1053 | 2.1802 | 2.6260 | 2.5100 | 3.0792 | 1.7736 | | 2.5264 | 3.1070 | 2.5479 | H9 | 3.4959 | 2.1858 | 1.1024 | 4.5741 | 3.8123 | 3.8123 | 2.5264 | 2.5264 | | 1.7806 | 1.7806 | H10 | 2.8142 | 2.1931 | 1.1046 | 4.0988 | 3.1770 | 2.6261 | 2.5479 | 3.1070 | 1.7806 | | 1.7826 | H11 | 2.8142 | 2.1931 | 1.1046 | 4.0988 | 2.6261 | 3.1770 | 3.1070 | 2.5479 | 1.7806 | 1.7826 | |
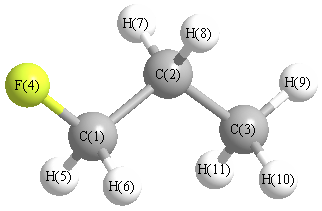 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.264 |
|
C1 |
C2 |
H7 |
108.444 |
| C1 |
C2 |
H8 |
108.444 |
|
C2 |
C1 |
F4 |
110.198 |
| C2 |
C1 |
H5 |
110.849 |
|
C2 |
C1 |
H6 |
110.849 |
| C2 |
C3 |
H9 |
111.011 |
|
C2 |
C3 |
H10 |
111.458 |
| C2 |
C3 |
H11 |
111.458 |
|
C3 |
C2 |
H7 |
110.398 |
| C3 |
C2 |
H8 |
110.398 |
|
F4 |
C1 |
H5 |
108.420 |
| F4 |
C1 |
H6 |
108.420 |
|
H5 |
C1 |
H6 |
108.019 |
| H7 |
C2 |
H8 |
106.706 |
|
H9 |
C3 |
H10 |
107.565 |
| H9 |
C3 |
H11 |
107.565 |
|
H10 |
C3 |
H11 |
107.586 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.011 |
|
|
|
| 2 |
C |
-0.320 |
|
|
|
| 3 |
C |
-0.498 |
|
|
|
| 4 |
F |
-0.285 |
|
|
|
| 5 |
H |
0.136 |
|
|
|
| 6 |
H |
0.136 |
|
|
|
| 7 |
H |
0.164 |
|
|
|
| 8 |
H |
0.164 |
|
|
|
| 9 |
H |
0.169 |
|
|
|
| 10 |
H |
0.162 |
|
|
|
| 11 |
H |
0.162 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.492 |
0.713 |
0.000 |
1.653 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.452 |
1.834 |
0.000 |
| y |
1.834 |
-25.578 |
0.000 |
| z |
0.000 |
0.000 |
-24.511 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.407 |
1.834 |
0.000 |
| y |
1.834 |
0.403 |
0.000 |
| z |
0.000 |
0.000 |
2.004 |
|
| Polar |
|---|
| 3z2-r2 | 4.008 |
|---|
| x2-y2 | -1.874 |
|---|
| xy | 1.834 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.279 |
-0.274 |
0.000 |
| y |
-0.274 |
5.282 |
0.000 |
| z |
0.000 |
0.000 |
4.845 |
<r2> (average value of r
2) Å
2
| <r2> |
104.188 |
| (<r2>)1/2 |
10.207 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -218.096277 |
| Energy at 298.15K | |
| HF Energy | -218.096277 |
| Nuclear repulsion energy | 129.710978 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3079 |
3028 |
21.55 |
|
|
|
| 2 |
A |
3061 |
3011 |
38.84 |
|
|
|
| 3 |
A |
3021 |
2971 |
32.11 |
|
|
|
| 4 |
A |
3001 |
2951 |
57.08 |
|
|
|
| 5 |
A |
2988 |
2939 |
21.69 |
|
|
|
| 6 |
A |
2976 |
2927 |
11.69 |
|
|
|
| 7 |
A |
2953 |
2904 |
47.61 |
|
|
|
| 8 |
A |
1496 |
1472 |
0.66 |
|
|
|
| 9 |
A |
1493 |
1468 |
7.96 |
|
|
|
| 10 |
A |
1477 |
1453 |
7.27 |
|
|
|
| 11 |
A |
1456 |
1432 |
2.36 |
|
|
|
| 12 |
A |
1404 |
1381 |
15.10 |
|
|
|
| 13 |
A |
1382 |
1360 |
7.15 |
|
|
|
| 14 |
A |
1350 |
1328 |
0.83 |
|
|
|
| 15 |
A |
1277 |
1256 |
0.92 |
|
|
|
| 16 |
A |
1243 |
1222 |
0.50 |
|
|
|
| 17 |
A |
1155 |
1136 |
1.10 |
|
|
|
| 18 |
A |
1103 |
1085 |
11.09 |
|
|
|
| 19 |
A |
1073 |
1055 |
42.21 |
|
|
|
| 20 |
A |
971 |
955 |
39.12 |
|
|
|
| 21 |
A |
904 |
889 |
3.29 |
|
|
|
| 22 |
A |
868 |
854 |
3.67 |
|
|
|
| 23 |
A |
756 |
744 |
0.69 |
|
|
|
| 24 |
A |
469 |
461 |
2.52 |
|
|
|
| 25 |
A |
311 |
306 |
1.05 |
|
|
|
| 26 |
A |
222 |
218 |
1.77 |
|
|
|
| 27 |
A |
148 |
146 |
1.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20818.7 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 20475.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.805 |
0.519 |
0.291 |
| C2 |
-0.599 |
0.666 |
-0.279 |
| C3 |
-1.524 |
-0.489 |
0.119 |
| F4 |
1.387 |
-0.668 |
-0.170 |
| H5 |
1.453 |
1.363 |
-0.014 |
| H6 |
0.779 |
0.480 |
1.399 |
| H7 |
-0.524 |
0.731 |
-1.380 |
| H8 |
-1.010 |
1.634 |
0.067 |
| H9 |
-2.521 |
-0.384 |
-0.340 |
| H10 |
-1.094 |
-1.451 |
-0.206 |
| H11 |
-1.658 |
-0.533 |
1.215 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5225 | 2.5433 | 1.4007 | 1.1070 | 1.1083 | 2.1462 | 2.1417 | 3.5033 | 2.7809 | 2.8331 |
C2 | 1.5225 | | 1.5322 | 2.3953 | 2.1835 | 2.1794 | 1.1058 | 1.1069 | 2.1906 | 2.1749 | 2.1888 | C3 | 2.5433 | 1.5322 | | 2.9306 | 3.5085 | 2.8073 | 2.1760 | 2.1844 | 1.1026 | 1.1021 | 1.1048 | F4 | 1.4007 | 2.3953 | 2.9306 | | 2.0382 | 2.0366 | 2.6602 | 3.3321 | 3.9218 | 2.6019 | 3.3479 | H5 | 1.1070 | 2.1835 | 3.5085 | 2.0382 | | 1.7972 | 2.4854 | 2.4797 | 4.3531 | 3.8001 | 3.8452 | H6 | 1.1083 | 2.1794 | 2.8073 | 2.0366 | 1.7972 | | 3.0800 | 2.5114 | 3.8289 | 3.1323 | 2.6461 | H7 | 2.1462 | 1.1058 | 2.1760 | 2.6602 | 2.4854 | 3.0800 | | 1.7739 | 2.5120 | 2.5421 | 3.1012 | H8 | 2.1417 | 1.1069 | 2.1844 | 3.3321 | 2.4797 | 2.5114 | 1.7739 | | 2.5529 | 3.0973 | 2.5359 | H9 | 3.5033 | 2.1906 | 1.1026 | 3.9218 | 4.3531 | 3.8289 | 2.5120 | 2.5529 | | 1.7863 | 1.7843 | H10 | 2.7809 | 2.1749 | 1.1021 | 2.6019 | 3.8001 | 3.1323 | 2.5421 | 3.0973 | 1.7863 | | 1.7827 | H11 | 2.8331 | 2.1888 | 1.1048 | 3.3479 | 3.8452 | 2.6461 | 3.1012 | 2.5359 | 1.7843 | 1.7827 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.735 |
|
C1 |
C2 |
H7 |
108.453 |
| C1 |
C2 |
H8 |
108.037 |
|
C2 |
C1 |
F4 |
109.979 |
| C2 |
C1 |
H5 |
111.300 |
|
C2 |
C1 |
H6 |
110.895 |
| C2 |
C3 |
H9 |
111.446 |
|
C2 |
C3 |
H10 |
110.234 |
| C2 |
C3 |
H11 |
111.173 |
|
C3 |
C2 |
H7 |
110.105 |
| C3 |
C2 |
H8 |
110.699 |
|
F4 |
C1 |
H5 |
108.166 |
| F4 |
C1 |
H6 |
107.957 |
|
H5 |
C1 |
H6 |
108.436 |
| H7 |
C2 |
H8 |
106.587 |
|
H9 |
C3 |
H10 |
108.233 |
| H9 |
C3 |
H11 |
107.861 |
|
H10 |
C3 |
H11 |
107.758 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.006 |
|
|
|
| 2 |
C |
-0.319 |
|
|
|
| 3 |
C |
-0.485 |
|
|
|
| 4 |
F |
-0.284 |
|
|
|
| 5 |
H |
0.140 |
|
|
|
| 6 |
H |
0.137 |
|
|
|
| 7 |
H |
0.164 |
|
|
|
| 8 |
H |
0.150 |
|
|
|
| 9 |
H |
0.160 |
|
|
|
| 10 |
H |
0.176 |
|
|
|
| 11 |
H |
0.156 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.816 |
1.242 |
0.370 |
1.531 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.415 |
1.441 |
0.522 |
| y |
1.441 |
-25.089 |
0.056 |
| z |
0.522 |
0.056 |
-24.612 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.565 |
1.441 |
0.522 |
| y |
1.441 |
0.425 |
0.056 |
| z |
0.522 |
0.056 |
1.140 |
|
| Polar |
|---|
| 3z2-r2 | 2.280 |
|---|
| x2-y2 | -1.326 |
|---|
| xy | 1.441 |
|---|
| xz | 0.522 |
|---|
| yz | 0.056 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.455 |
0.242 |
0.044 |
| y |
0.242 |
5.054 |
-0.021 |
| z |
0.044 |
-0.021 |
4.909 |
<r2> (average value of r
2) Å
2
| <r2> |
91.830 |
| (<r2>)1/2 |
9.583 |
