return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5F (Fluorobenzene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-330.323029
Energy at 298.15K-330.328776
HF Energy-329.554482
Nuclear repulsion energy270.810595
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3352 3096 0.08      
2 A1 3338 3084 18.07      
3 A1 3315 3062 0.05      
4 A1 1772 1637 54.49      
5 A1 1632 1508 92.00      
6 A1 1351 1248 85.63      
7 A1 1248 1153 6.38      
8 A1 1102 1018 4.13      
9 A1 1070 988 0.30      
10 A1 869 802 22.94      
11 A1 548 506 4.03      
12 A2 1008 931 0.00      
13 A2 886 819 0.00      
14 A2 441 407 0.00      
15 B1 1003 927 0.39      
16 B1 941 869 7.44      
17 B1 804 742 99.40      
18 B1 669 618 1.01      
19 B1 532 491 5.59      
20 B1 254 235 0.17      
21 B2 3350 3094 6.01      
22 B2 3325 3071 10.46      
23 B2 1769 1634 10.40      
24 B2 1582 1461 2.55      
25 B2 1410 1302 0.80      
26 B2 1349 1246 0.19      
27 B2 1233 1139 0.57      
28 B2 1154 1066 6.75      
29 B2 655 605 0.09      
30 B2 425 392 2.73      

Unscaled Zero Point Vibrational Energy (zpe) 21192.6 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 19575.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.19162 0.08658 0.05963

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
F1 0.000 0.000 2.266
C2 0.000 0.000 0.924
C3 0.000 1.209 0.259
C4 0.000 -1.209 0.259
C5 0.000 1.200 -1.127
C6 0.000 -1.200 -1.127
C7 0.000 0.000 -1.823
H8 0.000 2.128 0.824
H9 0.000 -2.128 0.824
H10 0.000 2.137 -1.664
H11 0.000 -2.137 -1.664
H12 0.000 0.000 -2.902

Atom - Atom Distances (Å)
  F1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
F11.34242.34282.34283.59903.59904.08892.57032.57034.47294.47295.1679
C21.34241.37961.37962.37602.37602.74652.13002.13003.35553.35553.8255
C32.34281.37962.41821.38642.77952.40781.07823.38422.13503.85903.3846
C42.34281.37962.41822.77951.38642.40783.38421.07823.85902.13503.3846
C53.59902.37601.38642.77952.40001.38712.16023.85741.07953.37952.1425
C63.59902.37602.77951.38642.40001.38713.85742.16023.37951.07952.1425
C74.08892.74652.40782.40781.38711.38713.39593.39592.14262.14261.0791
H82.57032.13001.07823.38422.16023.85743.39594.25532.48754.93684.2905
H92.57032.13003.38421.07823.85742.16023.39594.25534.93682.48754.2905
H104.47293.35552.13503.85901.07953.37952.14262.48754.93684.27342.4696
H114.47293.35553.85902.13503.37951.07952.14264.93682.48754.27342.4696
H125.16793.82553.38463.38462.14252.14251.07914.29054.29052.46962.4696

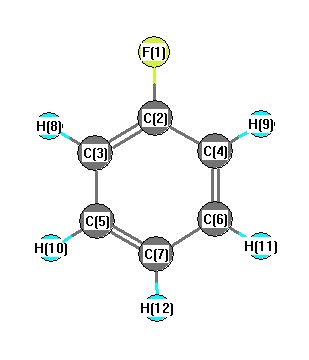
picture of Fluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
F1 C2 C3 118.785 F1 C2 C4 118.785
C2 C3 C5 118.409 C2 C3 H8 119.642
C2 C4 C6 118.409 C2 C4 H9 119.642
C3 C2 C4 122.431 C3 C5 C7 120.484
C3 C5 H10 119.429 C4 C6 C7 120.484
C4 C6 H11 119.429 C5 C3 H8 121.949
C5 C7 C6 119.784 C5 C7 H12 120.108
C6 C4 H9 121.949 C6 C7 H12 120.108
C7 C5 H10 120.087 C7 C6 H11 120.087
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability