return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5F (Fluorobenzene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-330.513432
Energy at 298.15K-330.518874
HF Energy-329.552490
Nuclear repulsion energy268.566430
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3237 3083 0.00      
2 A1 3223 3069 19.88      
3 A1 3200 3048 0.03      
4 A1 1696 1615 45.59      
5 A1 1568 1493 77.67      
6 A1 1293 1231 75.16      
7 A1 1205 1148 6.37      
8 A1 1060 1010 3.83      
9 A1 1030 981 0.20      
10 A1 833 794 20.71      
11 A1 526 501 3.66      
12 A2 923 879 0.00      
13 A2 829 789 0.00      
14 A2 414 395 0.00      
15 B1 916 872 0.93      
16 B1 868 827 3.80      
17 B1 753 717 93.72      
18 B1 576 549 0.06      
19 B1 495 472 5.48      
20 B1 240 229 0.11      
21 B2 3236 3082 7.07      
22 B2 3210 3057 11.54      
23 B2 1695 1614 7.85      
24 B2 1521 1448 2.15      
25 B2 1355 1290 0.59      
26 B2 1314 1251 0.15      
27 B2 1200 1143 0.45      
28 B2 1113 1060 6.23      
29 B2 631 601 0.07      
30 B2 406 386 2.60      

Unscaled Zero Point Vibrational Energy (zpe) 20282.5 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 19315.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.18850 0.08515 0.05865

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
F1 0.000 0.000 2.286
C2 0.000 0.000 0.928
C3 0.000 1.219 0.261
C4 0.000 -1.219 0.261
C5 0.000 1.210 -1.136
C6 0.000 -1.210 -1.136
C7 0.000 0.000 -1.837
H8 0.000 2.145 0.831
H9 0.000 -2.145 0.831
H10 0.000 2.154 -1.678
H11 0.000 -2.154 -1.678
H12 0.000 0.000 -2.925

Atom - Atom Distances (Å)
  F1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
F11.35802.36422.36423.63043.63044.12332.59242.59244.51194.51195.2117
C21.35801.39001.39002.39322.39322.76532.14732.14733.38143.38143.8537
C32.36421.39002.43851.39742.80222.42631.08723.41232.15253.89103.4114
C42.36421.39002.43852.80221.39742.42633.41231.08723.89102.15253.4114
C53.63042.39321.39742.80222.41951.39792.17833.88911.08873.40742.1594
C63.63042.39322.80221.39742.41951.39793.88912.17833.40741.08872.1594
C74.12332.76532.42632.42631.39791.39793.42313.42312.16022.16021.0883
H82.59242.14731.08723.41232.17833.88913.42314.29022.50874.97784.3254
H92.59242.14733.41231.08723.88912.17833.42314.29024.97782.50874.3254
H104.51193.38142.15253.89101.08873.40742.16022.50874.97784.30862.4893
H114.51193.38143.89102.15253.40741.08872.16024.97782.50874.30862.4893
H125.21173.85373.41143.41142.15942.15941.08834.32544.32542.48932.4893

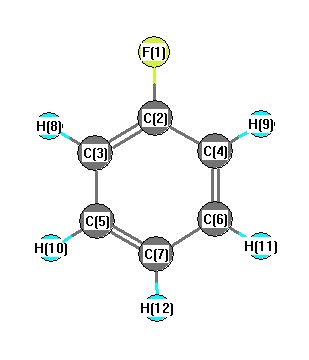
picture of Fluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
F1 C2 C3 118.703 F1 C2 C4 118.703
C2 C3 C5 118.314 C2 C3 H8 119.684
C2 C4 C6 118.314 C2 C4 H9 119.684
C3 C2 C4 122.595 C3 C5 C7 120.457
C3 C5 H10 119.434 C4 C6 C7 120.457
C4 C6 H11 119.434 C5 C3 H8 122.002
C5 C7 C6 119.862 C5 C7 H12 120.069
C6 C4 H9 122.002 C6 C7 H12 120.069
C7 C5 H10 120.109 C7 C6 H11 120.109
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability