return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2FCH2CH2F (1,3-difluoropropane)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-316.731591
Energy at 298.15K 
HF Energy-315.959530
Nuclear repulsion energy188.361648
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3154 3003 32.25      
2 A 3097 2949 5.80      
3 A 3075 2928 7.13      
4 A 1572 1497 0.37      
5 A 1521 1449 7.54      
6 A 1477 1406 13.33      
7 A 1338 1274 0.00      
8 A 1269 1209 0.81      
9 A 1152 1097 20.24      
10 A 1039 989 38.09      
11 A 906 863 0.11      
12 A 560 533 3.99      
13 A 257 245 4.19      
14 A 89 85 3.45      
15 B 3155 3004 36.14      
16 B 3123 2974 18.10      
17 B 3093 2946 64.66      
18 B 1575 1500 2.20      
19 B 1459 1390 12.89      
20 B 1431 1362 7.05      
21 B 1280 1219 6.88      
22 B 1164 1108 13.84      
23 B 1126 1073 77.33      
24 B 994 947 25.16      
25 B 803 765 2.30      
26 B 422 402 7.75      
27 B 211 201 12.15      

Unscaled Zero Point Vibrational Energy (zpe) 20170.1 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 19208.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.29448 0.10235 0.09481

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.001
C2 0.000 1.255 0.149
C3 0.000 -1.255 0.149
F4 1.157 1.259 -0.643
F5 -1.157 -1.259 -0.643
H6 0.887 -0.002 1.649
H7 -0.887 0.002 1.649
H8 -0.866 1.275 -0.521
H9 0.009 2.163 0.764
H10 0.866 -1.275 -0.521
H11 -0.009 -2.163 0.764

Atom - Atom Distances (Å)
  C1 C2 C3 F4 F5 H6 H7 H8 H9 H10 H11
C11.51651.51652.37242.37241.09861.09862.16592.17592.16592.1759
C21.51652.50941.40242.87872.14842.14581.09501.09672.75653.4724
C31.51652.50942.87871.40242.14582.14842.75653.47241.09501.0967
F42.37241.40242.87873.42012.63023.31852.02672.02832.55403.8796
F52.37242.87871.40243.42013.31852.63022.55403.87962.02672.0283
H61.09862.14842.14582.63023.31851.77483.06792.49852.51522.5010
H71.09862.14582.14843.31852.63021.77482.51522.50103.06792.4985
H82.16591.09502.75652.02672.55403.06792.51521.79003.08273.7688
H92.17591.09673.47242.02833.87962.49852.50101.79003.76884.3259
H102.16592.75651.09502.55402.02672.51523.06793.08273.76881.7900
H112.17593.47241.09673.87962.02832.50102.49853.76884.32591.7900

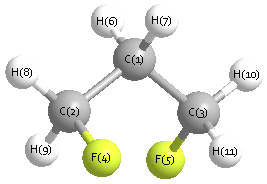
picture of 1,3-difluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 F4 108.673 C1 C2 H8 111.036
C1 C2 H9 111.743 C1 C3 F5 108.673
C1 C3 H10 111.036 C1 C3 H11 111.743
C2 C1 C3 111.661 C2 C1 H6 109.441
C2 C1 H7 109.233 C3 C1 H6 109.233
C3 C1 H7 109.441 F4 C2 H8 107.856
F4 C2 H9 107.888 F5 C3 H10 107.856
F5 C3 H11 107.888 H6 C1 H7 107.748
H8 C2 H9 109.512 H10 C3 H11 109.512
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability