return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CClCHCH3 (2-Butene, 2-chloro-, (Z)-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-616.458645
Energy at 298.15K-616.465558
HF Energy-616.243618
Nuclear repulsion energy213.397775
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3196 3034 19.01      
2 A' 3182 3020 4.72      
3 A' 3170 3009 11.75      
4 A' 3082 2926 19.26      
5 A' 3077 2920 24.32      
6 A' 1759 1670 10.86      
7 A' 1544 1465 8.10      
8 A' 1535 1457 0.72      
9 A' 1463 1389 1.26      
10 A' 1459 1385 2.35      
11 A' 1353 1284 26.00      
12 A' 1193 1133 27.92      
13 A' 1122 1065 3.33      
14 A' 1068 1014 15.54      
15 A' 929 882 28.16      
16 A' 618 587 26.00      
17 A' 531 504 0.20      
18 A' 346 328 0.48      
19 A' 239 227 1.35      
20 A" 3150 2990 12.78      
21 A" 3129 2970 20.16      
22 A" 1537 1459 6.26      
23 A" 1520 1443 8.56      
24 A" 1101 1046 0.39      
25 A" 1082 1027 0.92      
26 A" 834 792 12.42      
27 A" 471 448 1.00      
28 A" 254 241 1.68      
29 A" 193 184 1.43      
30 A" 106 100 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 22121.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 20998.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.16310 0.11039 0.06748

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.593 2.516 0.000
Cl2 -1.348 -0.554 0.000
C3 1.773 -1.343 0.000
C4 -0.325 1.929 0.000
H5 -0.906 2.202 0.881
H6 -0.906 2.202 -0.881
C7 0.000 0.486 0.000
C8 1.280 0.060 0.000
H9 2.014 0.815 0.000
H10 0.964 -2.059 0.000
H11 2.393 -1.526 -0.879
H12 2.393 -1.526 0.879

Atom - Atom Distances (Å)
  H1 Cl2 C3 C4 H5 H6 C7 C8 H9 H10 H11 H12
H13.63214.03581.09021.76671.76672.11542.55112.21704.59044.51174.5117
Cl23.63213.21942.68472.92672.92671.70212.69803.62982.75893.96363.9636
C34.03583.21943.88674.52984.52982.54741.48682.17111.08041.09101.0910
C41.09022.68473.88671.09001.09001.47902.46352.59094.19094.48294.4829
H51.76672.92674.52981.09001.76242.13113.18463.35044.73595.27994.9778
H61.76672.92674.52981.09001.76242.13113.18463.35044.73594.97785.2799
C72.11541.70212.54741.47902.13112.13111.34882.04082.72143.24763.2476
C82.55112.69801.48682.46353.18463.18461.34881.05362.14182.12762.1276
H92.21703.62982.17112.59093.35043.35042.04081.05363.05952.52922.5292
H104.59042.75891.08044.19094.73594.73592.72142.14183.05951.75991.7599
H114.51173.96361.09104.48295.27994.97783.24762.12762.52921.75991.7585
H124.51173.96361.09104.48294.97785.27993.24762.12762.52921.75991.7585

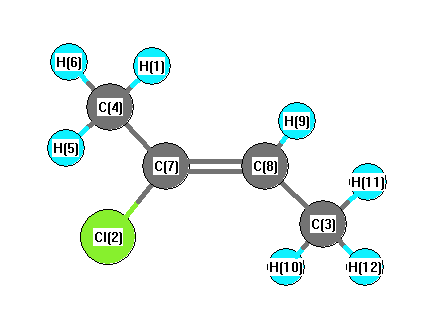
picture of 2-Butene, 2-chloro-, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C4 H5 108.255 H1 C4 H6 108.255
H1 C4 C7 109.929 Cl2 C7 C4 114.942
Cl2 C7 C8 123.930 C3 C8 C7 127.822
C3 C8 H9 116.406 C4 C7 C8 121.127
H5 C4 H6 107.886 H5 C4 C7 111.205
H6 C4 C7 111.205 C7 C8 H9 115.772
C8 C3 H10 112.117 C8 C3 H11 110.306
C8 C3 H12 110.306 H10 C3 H11 108.283
H10 C3 H12 108.283 H11 C3 H12 107.396
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.170      
2 Cl -0.052      
3 C -0.506      
4 C -0.489      
5 H 0.182      
6 H 0.182      
7 C -0.055      
8 C -0.122      
9 H 0.162      
10 H 0.191      
11 H 0.168      
12 H 0.168      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.050 -0.407 0.000
y -0.407 8.120 0.000
z 0.000 0.000 4.899


<r2> (average value of r2) Å2
<r2> 173.025
(<r2>)1/2 13.154