Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -357.326768 |
| Energy at 298.15K | -357.331918 |
| HF Energy | -356.543739 |
| Nuclear repulsion energy | 233.937907 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3896 |
3598 |
118.08 |
|
|
|
| 2 |
A' |
3893 |
3596 |
91.93 |
|
|
|
| 3 |
A' |
3759 |
3472 |
92.17 |
|
|
|
| 4 |
A' |
1978 |
1827 |
0.37 |
|
|
|
| 5 |
A' |
1950 |
1801 |
705.65 |
|
|
|
| 6 |
A' |
1715 |
1584 |
130.55 |
|
|
|
| 7 |
A' |
1548 |
1430 |
25.07 |
|
|
|
| 8 |
A' |
1411 |
1304 |
86.49 |
|
|
|
| 9 |
A' |
1289 |
1190 |
328.43 |
|
|
|
| 10 |
A' |
1172 |
1083 |
3.31 |
|
|
|
| 11 |
A' |
838 |
774 |
9.66 |
|
|
|
| 12 |
A' |
645 |
595 |
89.23 |
|
|
|
| 13 |
A' |
563 |
520 |
0.15 |
|
|
|
| 14 |
A' |
441 |
408 |
3.71 |
|
|
|
| 15 |
A' |
289 |
267 |
15.35 |
|
|
|
| 16 |
A" |
868 |
802 |
5.43 |
|
|
|
| 17 |
A" |
705 |
652 |
188.06 |
|
|
|
| 18 |
A" |
650 |
600 |
16.76 |
|
|
|
| 19 |
A" |
459 |
424 |
11.50 |
|
|
|
| 20 |
A" |
318 |
294 |
299.76 |
|
|
|
| 21 |
A" |
53 |
49 |
1.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14218.9 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 13134.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.749 |
0.000 |
| C2 |
-0.053 |
-0.782 |
0.000 |
| O3 |
-1.093 |
-1.388 |
0.000 |
| O4 |
1.031 |
1.365 |
0.000 |
| O5 |
-1.203 |
1.296 |
0.000 |
| N6 |
1.180 |
-1.314 |
0.000 |
| H7 |
1.278 |
-2.311 |
0.000 |
| H8 |
1.990 |
-0.726 |
0.000 |
| H9 |
-1.085 |
2.254 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5322 | 2.4012 | 1.2008 | 1.3213 | 2.3772 | 3.3163 | 2.4769 | 1.8549 |
C2 | 1.5322 | | 1.2038 | 2.4052 | 2.3744 | 1.3434 | 2.0274 | 2.0439 | 3.2062 | O3 | 2.4012 | 1.2038 | | 3.4776 | 2.6864 | 2.2744 | 2.5445 | 3.1534 | 3.6423 | O4 | 1.2008 | 2.4052 | 3.4776 | | 2.2353 | 2.6832 | 3.6837 | 2.2998 | 2.2954 | O5 | 1.3213 | 2.3744 | 2.6864 | 2.2353 | | 3.5342 | 4.3775 | 3.7789 | 0.9654 | N6 | 2.3772 | 1.3434 | 2.2744 | 2.6832 | 3.5342 | | 1.0012 | 1.0010 | 4.2263 | H7 | 3.3163 | 2.0274 | 2.5445 | 3.6837 | 4.3775 | 1.0012 | | 1.7372 | 5.1400 | H8 | 2.4769 | 2.0439 | 3.1534 | 2.2998 | 3.7789 | 1.0010 | 1.7372 | | 4.2816 | H9 | 1.8549 | 3.2062 | 3.6423 | 2.2954 | 0.9654 | 4.2263 | 5.1400 | 4.2816 | |
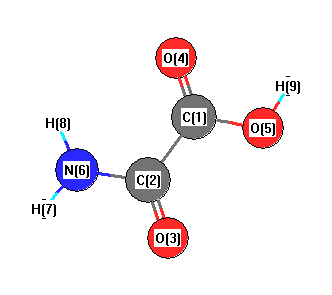 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
122.259 |
|
C1 |
C2 |
N6 |
111.346 |
| C1 |
O5 |
H9 |
107.397 |
|
C2 |
C1 |
O4 |
122.830 |
| C2 |
C1 |
O5 |
112.420 |
|
C2 |
N6 |
H7 |
118.984 |
| C2 |
N6 |
H8 |
120.644 |
|
O3 |
C2 |
N6 |
126.395 |
| O4 |
C1 |
O5 |
124.750 |
|
H7 |
N6 |
H8 |
120.373 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -357.331947 |
| Energy at 298.15K | -357.337370 |
| Nuclear repulsion energy | 235.237737 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3880 |
3584 |
109.43 |
|
|
|
| 2 |
A' |
3802 |
3512 |
160.35 |
|
|
|
| 3 |
A' |
3748 |
3462 |
100.61 |
|
|
|
| 4 |
A' |
2011 |
1857 |
217.79 |
|
|
|
| 5 |
A' |
1931 |
1784 |
444.60 |
|
|
|
| 6 |
A' |
1718 |
1587 |
81.70 |
|
|
|
| 7 |
A' |
1565 |
1446 |
197.81 |
|
|
|
| 8 |
A' |
1435 |
1326 |
474.31 |
|
|
|
| 9 |
A' |
1303 |
1204 |
6.18 |
|
|
|
| 10 |
A' |
1175 |
1085 |
3.31 |
|
|
|
| 11 |
A' |
864 |
798 |
10.78 |
|
|
|
| 12 |
A' |
662 |
611 |
19.62 |
|
|
|
| 13 |
A' |
579 |
535 |
2.93 |
|
|
|
| 14 |
A' |
437 |
404 |
7.10 |
|
|
|
| 15 |
A' |
297 |
275 |
40.27 |
|
|
|
| 16 |
A" |
850 |
785 |
0.46 |
|
|
|
| 17 |
A" |
741 |
685 |
146.48 |
|
|
|
| 18 |
A" |
684 |
631 |
9.29 |
|
|
|
| 19 |
A" |
492 |
454 |
154.95 |
|
|
|
| 20 |
A" |
393 |
363 |
193.98 |
|
|
|
| 21 |
A" |
110 |
101 |
4.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14337.6 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 13243.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.006 |
-0.789 |
0.000 |
| C2 |
0.000 |
0.744 |
0.000 |
| O3 |
-1.063 |
1.332 |
0.000 |
| O4 |
1.015 |
-1.431 |
0.000 |
| O5 |
-1.212 |
-1.293 |
0.000 |
| N6 |
1.218 |
1.284 |
0.000 |
| H7 |
1.322 |
2.280 |
0.000 |
| H8 |
2.025 |
0.689 |
0.000 |
| H9 |
-1.834 |
-0.548 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5333 | 2.3751 | 1.1957 | 1.3185 | 2.4010 | 3.3394 | 2.5014 | 1.8556 |
C2 | 1.5333 | | 1.2142 | 2.4004 | 2.3707 | 1.3321 | 2.0265 | 2.0253 | 2.2432 | O3 | 2.3751 | 1.2142 | | 3.4569 | 2.6294 | 2.2810 | 2.5662 | 3.1534 | 2.0319 | O4 | 1.1957 | 2.4004 | 3.4569 | | 2.2313 | 2.7223 | 3.7238 | 2.3478 | 2.9823 | O5 | 1.3185 | 2.3707 | 2.6294 | 2.2313 | | 3.5419 | 4.3807 | 3.7952 | 0.9704 | N6 | 2.4010 | 1.3321 | 2.2810 | 2.7223 | 3.5419 | | 1.0018 | 1.0024 | 3.5591 | H7 | 3.3394 | 2.0265 | 2.5662 | 3.7238 | 4.3807 | 1.0018 | | 1.7396 | 4.2375 | H8 | 2.5014 | 2.0253 | 3.1534 | 2.3478 | 3.7952 | 1.0024 | 1.7396 | | 4.0515 | H9 | 1.8556 | 2.2432 | 2.0319 | 2.9823 | 0.9704 | 3.5591 | 4.2375 | 4.0515 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.181 |
|
C1 |
C2 |
N6 |
113.657 |
| C1 |
O5 |
H9 |
107.354 |
|
C2 |
C1 |
O4 |
122.707 |
| C2 |
C1 |
O5 |
112.245 |
|
C2 |
N6 |
H7 |
119.859 |
| C2 |
N6 |
H8 |
119.691 |
|
O3 |
C2 |
N6 |
127.162 |
| O4 |
C1 |
O5 |
125.048 |
|
H7 |
N6 |
H8 |
120.450 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
