Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -358.319572 |
| Energy at 298.15K | -358.324581 |
| Nuclear repulsion energy | 232.268236 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3750 |
3588 |
80.04 |
|
|
|
| 2 |
A' |
3720 |
3559 |
68.76 |
|
|
|
| 3 |
A' |
3615 |
3459 |
62.18 |
|
|
|
| 4 |
A' |
1860 |
1780 |
56.98 |
|
|
|
| 5 |
A' |
1841 |
1761 |
497.23 |
|
|
|
| 6 |
A' |
1622 |
1551 |
130.11 |
|
|
|
| 7 |
A' |
1449 |
1386 |
19.10 |
|
|
|
| 8 |
A' |
1339 |
1281 |
61.85 |
|
|
|
| 9 |
A' |
1212 |
1159 |
277.85 |
|
|
|
| 10 |
A' |
1106 |
1058 |
2.58 |
|
|
|
| 11 |
A' |
785 |
751 |
9.77 |
|
|
|
| 12 |
A' |
611 |
584 |
73.34 |
|
|
|
| 13 |
A' |
529 |
507 |
0.34 |
|
|
|
| 14 |
A' |
414 |
396 |
4.34 |
|
|
|
| 15 |
A' |
267 |
256 |
15.33 |
|
|
|
| 16 |
A" |
824 |
789 |
4.81 |
|
|
|
| 17 |
A" |
694 |
664 |
151.13 |
|
|
|
| 18 |
A" |
644 |
616 |
13.97 |
|
|
|
| 19 |
A" |
437 |
418 |
19.34 |
|
|
|
| 20 |
A" |
322 |
309 |
249.66 |
|
|
|
| 21 |
A" |
69 |
66 |
2.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13553.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12966.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.752 |
0.000 |
| C2 |
-0.057 |
-0.788 |
0.000 |
| O3 |
-1.099 |
-1.412 |
0.000 |
| O4 |
1.037 |
1.380 |
0.000 |
| O5 |
-1.215 |
1.297 |
0.000 |
| N6 |
1.191 |
-1.308 |
0.000 |
| H7 |
1.311 |
-2.309 |
0.000 |
| H8 |
1.992 |
-0.693 |
0.000 |
| H9 |
-1.084 |
2.262 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5411 | 2.4271 | 1.2126 | 1.3318 | 2.3792 | 3.3297 | 2.4612 | 1.8592 |
C2 | 1.5411 | | 1.2149 | 2.4286 | 2.3854 | 1.3518 | 2.0454 | 2.0512 | 3.2190 | O3 | 2.4271 | 1.2149 | | 3.5157 | 2.7115 | 2.2929 | 2.5719 | 3.1740 | 3.6746 | O4 | 1.2126 | 2.4286 | 3.5157 | | 2.2538 | 2.6922 | 3.6991 | 2.2828 | 2.2970 | O5 | 1.3318 | 2.3854 | 2.7115 | 2.2538 | | 3.5462 | 4.4028 | 3.7747 | 0.9745 | N6 | 2.3792 | 1.3518 | 2.2929 | 2.6922 | 3.5462 | | 1.0084 | 1.0095 | 4.2334 | H7 | 3.3297 | 2.0454 | 2.5719 | 3.6991 | 4.4028 | 1.0084 | | 1.7533 | 5.1607 | H8 | 2.4612 | 2.0512 | 3.1740 | 2.2828 | 3.7747 | 1.0095 | 1.7533 | | 4.2659 | H9 | 1.8592 | 3.2190 | 3.6746 | 2.2970 | 0.9745 | 4.2334 | 5.1607 | 4.2659 | |
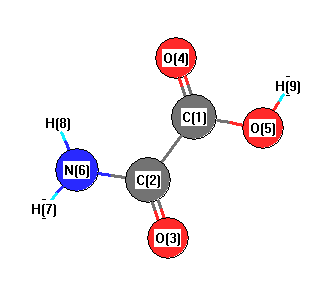 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
123.004 |
|
C1 |
C2 |
N6 |
110.487 |
| C1 |
O5 |
H9 |
106.409 |
|
C2 |
C1 |
O4 |
123.317 |
| C2 |
C1 |
O5 |
112.055 |
|
C2 |
N6 |
H7 |
119.427 |
| C2 |
N6 |
H8 |
119.912 |
|
O3 |
C2 |
N6 |
126.509 |
| O4 |
C1 |
O5 |
124.628 |
|
H7 |
N6 |
H8 |
120.661 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.535 |
|
|
0.539 |
| 2 |
C |
0.556 |
|
|
0.621 |
| 3 |
O |
-0.482 |
|
|
-0.481 |
| 4 |
O |
-0.477 |
|
|
-0.490 |
| 5 |
O |
-0.541 |
|
|
-0.558 |
| 6 |
N |
-0.781 |
|
|
-0.949 |
| 7 |
H |
0.372 |
|
|
0.445 |
| 8 |
H |
0.386 |
|
|
0.420 |
| 9 |
H |
0.432 |
|
|
0.453 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.192 |
0.694 |
0.000 |
2.299 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
2.216 |
0.686 |
0.000 |
2.320 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.756 |
-7.835 |
-0.000 |
| y |
-7.835 |
-38.133 |
0.000 |
| z |
-0.000 |
0.000 |
-32.964 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
11.793 |
-7.835 |
-0.000 |
| y |
-7.835 |
-9.773 |
0.000 |
| z |
-0.000 |
0.000 |
-2.020 |
|
| Polar |
|---|
| 3z2-r2 | -4.040 |
|---|
| x2-y2 | 14.377 |
|---|
| xy | -7.835 |
|---|
| xz | -0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
141.307 |
| (<r2>)1/2 |
11.887 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -358.326390 |
| Energy at 298.15K | -358.331668 |
| Nuclear repulsion energy | 233.879160 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3738 |
3576 |
86.80 |
|
|
|
| 2 |
A' |
3606 |
3450 |
72.71 |
|
|
|
| 3 |
A' |
3518 |
3366 |
143.16 |
|
|
|
| 4 |
A' |
1897 |
1815 |
219.63 |
|
|
|
| 5 |
A' |
1829 |
1750 |
286.30 |
|
|
|
| 6 |
A' |
1626 |
1555 |
76.00 |
|
|
|
| 7 |
A' |
1476 |
1412 |
264.65 |
|
|
|
| 8 |
A' |
1372 |
1313 |
330.16 |
|
|
|
| 9 |
A' |
1237 |
1184 |
4.96 |
|
|
|
| 10 |
A' |
1110 |
1062 |
5.68 |
|
|
|
| 11 |
A' |
813 |
778 |
9.64 |
|
|
|
| 12 |
A' |
631 |
604 |
12.12 |
|
|
|
| 13 |
A' |
547 |
524 |
1.83 |
|
|
|
| 14 |
A' |
407 |
390 |
8.98 |
|
|
|
| 15 |
A' |
273 |
261 |
41.87 |
|
|
|
| 16 |
A" |
813 |
778 |
17.39 |
|
|
|
| 17 |
A" |
776 |
743 |
97.50 |
|
|
|
| 18 |
A" |
672 |
643 |
2.98 |
|
|
|
| 19 |
A" |
470 |
450 |
156.18 |
|
|
|
| 20 |
A" |
379 |
363 |
141.34 |
|
|
|
| 21 |
A" |
122 |
116 |
4.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13657.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 13065.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.012 |
-0.795 |
0.000 |
| C2 |
0.000 |
0.746 |
0.000 |
| O3 |
-1.076 |
1.335 |
0.000 |
| O4 |
1.024 |
-1.454 |
0.000 |
| O5 |
-1.223 |
-1.279 |
0.000 |
| N6 |
1.225 |
1.287 |
0.000 |
| H7 |
1.343 |
2.289 |
0.000 |
| H8 |
2.027 |
0.672 |
0.000 |
| H9 |
-1.807 |
-0.485 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5413 | 2.3923 | 1.2072 | 1.3267 | 2.4096 | 3.3592 | 2.4933 | 1.8452 |
C2 | 1.5413 | | 1.2269 | 2.4262 | 2.3658 | 1.3387 | 2.0456 | 2.0286 | 2.1861 | O3 | 2.3923 | 1.2269 | | 3.4910 | 2.6184 | 2.3011 | 2.6003 | 3.1732 | 1.9612 | O4 | 1.2072 | 2.4262 | 3.4910 | | 2.2539 | 2.7480 | 3.7563 | 2.3514 | 2.9919 | O5 | 1.3267 | 2.3658 | 2.6184 | 2.2539 | | 3.5463 | 4.3951 | 3.7916 | 0.9856 | N6 | 2.4096 | 1.3387 | 2.3011 | 2.7480 | 3.5463 | | 1.0090 | 1.0108 | 3.5111 | H7 | 3.3592 | 2.0456 | 2.6003 | 3.7563 | 4.3951 | 1.0090 | | 1.7551 | 4.1970 | H8 | 2.4933 | 2.0286 | 3.1732 | 2.3514 | 3.7916 | 1.0108 | 1.7551 | | 4.0050 | H9 | 1.8452 | 2.1861 | 1.9612 | 2.9919 | 0.9856 | 3.5111 | 4.1970 | 4.0050 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.152 |
|
C1 |
C2 |
N6 |
113.393 |
| C1 |
O5 |
H9 |
104.914 |
|
C2 |
C1 |
O4 |
123.499 |
| C2 |
C1 |
O5 |
110.941 |
|
C2 |
N6 |
H7 |
120.584 |
| C2 |
N6 |
H8 |
118.742 |
|
O3 |
C2 |
N6 |
127.455 |
| O4 |
C1 |
O5 |
125.560 |
|
H7 |
N6 |
H8 |
120.674 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.576 |
|
|
|
| 2 |
C |
0.538 |
|
|
|
| 3 |
O |
-0.528 |
|
|
|
| 4 |
O |
-0.466 |
|
|
|
| 5 |
O |
-0.572 |
|
|
|
| 6 |
N |
-0.776 |
|
|
|
| 7 |
H |
0.379 |
|
|
|
| 8 |
H |
0.395 |
|
|
|
| 9 |
H |
0.453 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.563 |
2.689 |
0.000 |
3.110 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.724 |
7.121 |
0.000 |
| y |
7.121 |
-36.581 |
0.000 |
| z |
0.000 |
0.000 |
-32.924 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.028 |
7.121 |
0.000 |
| y |
7.121 |
-5.257 |
0.000 |
| z |
0.000 |
0.000 |
0.229 |
|
| Polar |
|---|
| 3z2-r2 | 0.458 |
|---|
| x2-y2 | 6.857 |
|---|
| xy | 7.121 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.869 |
0.021 |
0.000 |
| y |
0.021 |
5.405 |
0.000 |
| z |
0.000 |
0.000 |
2.462 |
<r2> (average value of r
2) Å
2
| <r2> |
138.865 |
| (<r2>)1/2 |
11.784 |
