Jump to
S1C2
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -357.483720 |
| Energy at 298.15K | -357.488658 |
| HF Energy | -356.541210 |
| Nuclear repulsion energy | 232.475827 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3798 |
3565 |
87.21 |
|
|
|
| 2 |
A' |
3765 |
3534 |
83.81 |
|
|
|
| 3 |
A' |
3668 |
3443 |
87.01 |
|
|
|
| 4 |
A' |
1920 |
1802 |
1.73 |
|
|
|
| 5 |
A' |
1901 |
1784 |
599.84 |
|
|
|
| 6 |
A' |
1661 |
1559 |
128.71 |
|
|
|
| 7 |
A' |
1503 |
1411 |
21.76 |
|
|
|
| 8 |
A' |
1378 |
1293 |
61.85 |
|
|
|
| 9 |
A' |
1252 |
1175 |
327.96 |
|
|
|
| 10 |
A' |
1138 |
1069 |
3.33 |
|
|
|
| 11 |
A' |
813 |
763 |
8.67 |
|
|
|
| 12 |
A' |
624 |
586 |
82.34 |
|
|
|
| 13 |
A' |
548 |
514 |
0.18 |
|
|
|
| 14 |
A' |
429 |
403 |
3.89 |
|
|
|
| 15 |
A' |
281 |
263 |
14.36 |
|
|
|
| 16 |
A" |
819 |
769 |
7.11 |
|
|
|
| 17 |
A" |
684 |
642 |
167.08 |
|
|
|
| 18 |
A" |
633 |
594 |
12.57 |
|
|
|
| 19 |
A" |
436 |
410 |
3.38 |
|
|
|
| 20 |
A" |
255 |
239 |
300.71 |
|
|
|
| 21 |
A" |
43 |
40 |
0.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13773.2 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 12927.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.751 |
0.000 |
| C2 |
-0.053 |
-0.784 |
0.000 |
| O3 |
-1.099 |
-1.399 |
0.000 |
| O4 |
1.037 |
1.374 |
0.000 |
| O5 |
-1.215 |
1.300 |
0.000 |
| N6 |
1.189 |
-1.317 |
0.000 |
| H7 |
1.291 |
-2.320 |
0.000 |
| H8 |
2.001 |
-0.719 |
0.000 |
| H9 |
-1.084 |
2.266 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5358 | 2.4145 | 1.2102 | 1.3334 | 2.3852 | 3.3306 | 2.4827 | 1.8627 |
C2 | 1.5358 | | 1.2136 | 2.4179 | 2.3862 | 1.3517 | 2.0406 | 2.0549 | 3.2194 | O3 | 2.4145 | 1.2136 | | 3.5007 | 2.7015 | 2.2902 | 2.5617 | 3.1739 | 3.6648 | O4 | 1.2102 | 2.4179 | 3.5007 | | 2.2535 | 2.6951 | 3.7021 | 2.3041 | 2.3007 | O5 | 1.3334 | 2.3862 | 2.7015 | 2.2535 | | 3.5537 | 4.4025 | 3.7971 | 0.9747 | N6 | 2.3852 | 1.3517 | 2.2902 | 2.6951 | 3.5537 | | 1.0078 | 1.0079 | 4.2428 | H7 | 3.3306 | 2.0406 | 2.5617 | 3.7021 | 4.4025 | 1.0078 | | 1.7507 | 5.1637 | H8 | 2.4827 | 2.0549 | 3.1739 | 2.3041 | 3.7971 | 1.0079 | 1.7507 | | 4.2922 | H9 | 1.8627 | 3.2194 | 3.6648 | 2.3007 | 0.9747 | 4.2428 | 5.1637 | 4.2922 | |
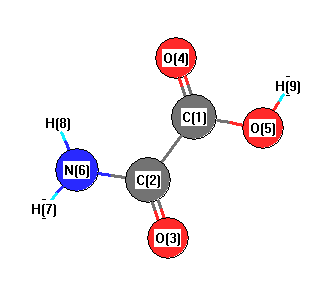 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
122.416 |
|
C1 |
C2 |
N6 |
111.230 |
| C1 |
O5 |
H9 |
106.581 |
|
C2 |
C1 |
O4 |
122.977 |
| C2 |
C1 |
O5 |
112.352 |
|
C2 |
N6 |
H7 |
119.014 |
| C2 |
N6 |
H8 |
120.416 |
|
O3 |
C2 |
N6 |
126.354 |
| O4 |
C1 |
O5 |
124.670 |
|
H7 |
N6 |
H8 |
120.570 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -357.489280 |
| Energy at 298.15K | -357.494543 |
| Nuclear repulsion energy | 233.803640 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3785 |
3553 |
95.64 |
|
|
|
| 2 |
A' |
3661 |
3437 |
161.62 |
|
|
|
| 3 |
A' |
3655 |
3431 |
71.38 |
|
|
|
| 4 |
A' |
1953 |
1833 |
198.80 |
|
|
|
| 5 |
A' |
1885 |
1769 |
363.78 |
|
|
|
| 6 |
A' |
1665 |
1563 |
77.57 |
|
|
|
| 7 |
A' |
1521 |
1428 |
223.04 |
|
|
|
| 8 |
A' |
1403 |
1316 |
413.30 |
|
|
|
| 9 |
A' |
1271 |
1193 |
9.90 |
|
|
|
| 10 |
A' |
1142 |
1072 |
4.39 |
|
|
|
| 11 |
A' |
836 |
785 |
11.32 |
|
|
|
| 12 |
A' |
641 |
602 |
16.79 |
|
|
|
| 13 |
A' |
563 |
529 |
2.65 |
|
|
|
| 14 |
A' |
425 |
399 |
7.07 |
|
|
|
| 15 |
A' |
290 |
273 |
39.12 |
|
|
|
| 16 |
A" |
802 |
752 |
2.58 |
|
|
|
| 17 |
A" |
731 |
686 |
134.77 |
|
|
|
| 18 |
A" |
665 |
624 |
7.27 |
|
|
|
| 19 |
A" |
462 |
433 |
97.86 |
|
|
|
| 20 |
A" |
349 |
328 |
231.82 |
|
|
|
| 21 |
A" |
108 |
101 |
2.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13905.9 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 13052.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.007 |
-0.791 |
0.000 |
| C2 |
0.000 |
0.747 |
0.000 |
| O3 |
-1.070 |
1.340 |
0.000 |
| O4 |
1.023 |
-1.440 |
0.000 |
| O5 |
-1.222 |
-1.299 |
0.000 |
| N6 |
1.226 |
1.288 |
0.000 |
| H7 |
1.334 |
2.290 |
0.000 |
| H8 |
2.034 |
0.683 |
0.000 |
| H9 |
-1.835 |
-0.534 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5374 | 2.3880 | 1.2053 | 1.3299 | 2.4092 | 3.3545 | 2.5057 | 1.8600 |
C2 | 1.5374 | | 1.2240 | 2.4140 | 2.3827 | 1.3398 | 2.0401 | 2.0349 | 2.2374 | O3 | 2.3880 | 1.2240 | | 3.4801 | 2.6435 | 2.2968 | 2.5853 | 3.1732 | 2.0238 | O4 | 1.2053 | 2.4140 | 3.4801 | | 2.2489 | 2.7351 | 3.7432 | 2.3513 | 2.9979 | O5 | 1.3299 | 2.3827 | 2.6435 | 2.2489 | | 3.5609 | 4.4061 | 3.8113 | 0.9806 | N6 | 2.4092 | 1.3398 | 2.2968 | 2.7351 | 3.5609 | | 1.0084 | 1.0094 | 3.5615 | H7 | 3.3545 | 2.0401 | 2.5853 | 3.7432 | 4.4061 | 1.0084 | | 1.7532 | 4.2446 | H8 | 2.5057 | 2.0349 | 3.1732 | 2.3513 | 3.8113 | 1.0094 | 1.7532 | | 4.0555 | H9 | 1.8600 | 2.2374 | 2.0238 | 2.9979 | 0.9806 | 3.5615 | 4.2446 | 4.0555 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.281 |
|
C1 |
C2 |
N6 |
113.539 |
| C1 |
O5 |
H9 |
106.236 |
|
C2 |
C1 |
O4 |
122.866 |
| C2 |
C1 |
O5 |
112.195 |
|
C2 |
N6 |
H7 |
119.974 |
| C2 |
N6 |
H8 |
119.378 |
|
O3 |
C2 |
N6 |
127.180 |
| O4 |
C1 |
O5 |
124.939 |
|
H7 |
N6 |
H8 |
120.648 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
