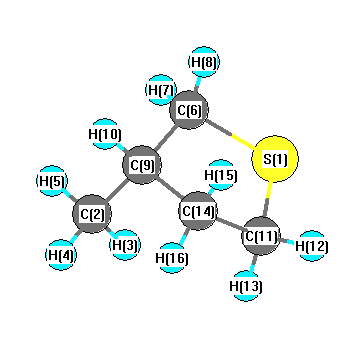
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3144 |
3008 |
15.34 |
|
|
|
| 2 |
A |
3138 |
3002 |
27.94 |
|
|
|
| 3 |
A |
3137 |
3001 |
18.59 |
|
|
|
| 4 |
A |
3130 |
2995 |
24.11 |
|
|
|
| 5 |
A |
3110 |
2976 |
26.05 |
|
|
|
| 6 |
A |
3088 |
2954 |
10.58 |
|
|
|
| 7 |
A |
3084 |
2951 |
52.11 |
|
|
|
| 8 |
A |
3065 |
2933 |
34.28 |
|
|
|
| 9 |
A |
3057 |
2924 |
24.89 |
|
|
|
| 10 |
A |
3051 |
2919 |
8.84 |
|
|
|
| 11 |
A |
1536 |
1470 |
6.23 |
|
|
|
| 12 |
A |
1531 |
1465 |
1.52 |
|
|
|
| 13 |
A |
1518 |
1453 |
6.00 |
|
|
|
| 14 |
A |
1511 |
1446 |
3.57 |
|
|
|
| 15 |
A |
1502 |
1437 |
4.13 |
|
|
|
| 16 |
A |
1435 |
1373 |
10.66 |
|
|
|
| 17 |
A |
1386 |
1326 |
5.64 |
|
|
|
| 18 |
A |
1358 |
1299 |
1.00 |
|
|
|
| 19 |
A |
1332 |
1275 |
3.76 |
|
|
|
| 20 |
A |
1312 |
1255 |
6.13 |
|
|
|
| 21 |
A |
1274 |
1219 |
13.27 |
|
|
|
| 22 |
A |
1249 |
1195 |
4.93 |
|
|
|
| 23 |
A |
1196 |
1144 |
2.20 |
|
|
|
| 24 |
A |
1151 |
1101 |
0.15 |
|
|
|
| 25 |
A |
1093 |
1046 |
1.08 |
|
|
|
| 26 |
A |
1082 |
1035 |
1.12 |
|
|
|
| 27 |
A |
1053 |
1007 |
0.44 |
|
|
|
| 28 |
A |
1008 |
965 |
1.82 |
|
|
|
| 29 |
A |
973 |
931 |
0.22 |
|
|
|
| 30 |
A |
943 |
902 |
0.43 |
|
|
|
| 31 |
A |
912 |
873 |
3.22 |
|
|
|
| 32 |
A |
866 |
829 |
1.12 |
|
|
|
| 33 |
A |
811 |
776 |
1.54 |
|
|
|
| 34 |
A |
730 |
698 |
5.26 |
|
|
|
| 35 |
A |
700 |
670 |
2.51 |
|
|
|
| 36 |
A |
615 |
588 |
1.05 |
|
|
|
| 37 |
A |
497 |
475 |
0.10 |
|
|
|
| 38 |
A |
378 |
361 |
0.14 |
|
|
|
| 39 |
A |
352 |
337 |
0.14 |
|
|
|
| 40 |
A |
233 |
223 |
0.09 |
|
|
|
| 41 |
A |
196 |
187 |
0.31 |
|
|
|
| 42 |
A |
111 |
106 |
2.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31423.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30062.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.078 |
|
|
-0.277 |
| 2 |
C |
-0.511 |
|
|
-0.325 |
| 3 |
H |
0.187 |
|
|
0.091 |
| 4 |
H |
0.167 |
|
|
0.102 |
| 5 |
H |
0.170 |
|
|
0.076 |
| 6 |
C |
-0.467 |
|
|
-0.147 |
| 7 |
H |
0.198 |
|
|
0.124 |
| 8 |
H |
0.196 |
|
|
0.091 |
| 9 |
C |
-0.140 |
|
|
0.171 |
| 10 |
H |
0.167 |
|
|
0.022 |
| 11 |
C |
-0.475 |
|
|
0.003 |
| 12 |
H |
0.201 |
|
|
0.087 |
| 13 |
H |
0.197 |
|
|
0.081 |
| 14 |
C |
-0.317 |
|
|
-0.318 |
| 15 |
H |
0.177 |
|
|
0.117 |
| 16 |
H |
0.172 |
|
|
0.103 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.966 |
0.839 |
0.715 |
2.254 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.965 |
0.874 |
0.717 |
2.267 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.426 |
-2.001 |
-0.150 |
| y |
-2.001 |
-42.758 |
-1.257 |
| z |
-0.150 |
-1.257 |
-45.389 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.353 |
-2.001 |
-0.150 |
| y |
-2.001 |
4.150 |
-1.257 |
| z |
-0.150 |
-1.257 |
0.203 |
|
| Polar |
|---|
| 3z2-r2 | 0.406 |
|---|
| x2-y2 | -5.668 |
|---|
| xy | -2.001 |
|---|
| xz | -0.150 |
|---|
| yz | -1.257 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
194.625 |
| (<r2>)1/2 |
13.951 |