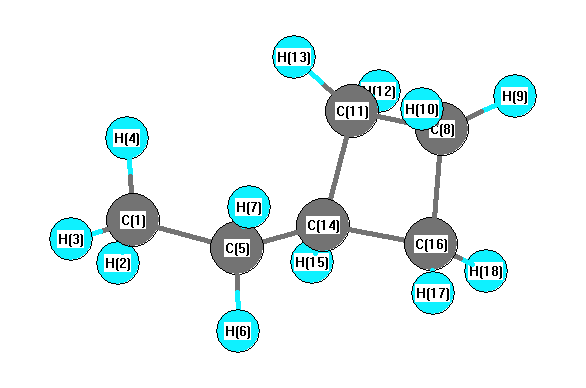
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3140 |
3004 |
80.10 |
|
|
|
| 2 |
A |
3129 |
2994 |
39.56 |
|
|
|
| 3 |
A |
3125 |
2990 |
43.85 |
|
|
|
| 4 |
A |
3122 |
2987 |
4.51 |
|
|
|
| 5 |
A |
3114 |
2979 |
13.61 |
|
|
|
| 6 |
A |
3084 |
2951 |
18.72 |
|
|
|
| 7 |
A |
3075 |
2942 |
63.82 |
|
|
|
| 8 |
A |
3070 |
2937 |
61.46 |
|
|
|
| 9 |
A |
3068 |
2936 |
30.15 |
|
|
|
| 10 |
A |
3053 |
2921 |
36.71 |
|
|
|
| 11 |
A |
3051 |
2919 |
9.31 |
|
|
|
| 12 |
A |
3032 |
2900 |
23.25 |
|
|
|
| 13 |
A |
1536 |
1469 |
2.42 |
|
|
|
| 14 |
A |
1530 |
1464 |
4.15 |
|
|
|
| 15 |
A |
1525 |
1459 |
6.45 |
|
|
|
| 16 |
A |
1511 |
1446 |
0.73 |
|
|
|
| 17 |
A |
1506 |
1441 |
1.55 |
|
|
|
| 18 |
A |
1499 |
1434 |
1.71 |
|
|
|
| 19 |
A |
1434 |
1372 |
4.81 |
|
|
|
| 20 |
A |
1409 |
1348 |
0.18 |
|
|
|
| 21 |
A |
1355 |
1297 |
5.97 |
|
|
|
| 22 |
A |
1318 |
1261 |
0.96 |
|
|
|
| 23 |
A |
1301 |
1245 |
1.57 |
|
|
|
| 24 |
A |
1294 |
1238 |
0.45 |
|
|
|
| 25 |
A |
1272 |
1216 |
0.07 |
|
|
|
| 26 |
A |
1268 |
1213 |
0.00 |
|
|
|
| 27 |
A |
1256 |
1202 |
0.10 |
|
|
|
| 28 |
A |
1205 |
1153 |
0.94 |
|
|
|
| 29 |
A |
1198 |
1146 |
0.11 |
|
|
|
| 30 |
A |
1146 |
1096 |
1.25 |
|
|
|
| 31 |
A |
1095 |
1047 |
0.07 |
|
|
|
| 32 |
A |
1041 |
996 |
0.08 |
|
|
|
| 33 |
A |
1015 |
971 |
0.35 |
|
|
|
| 34 |
A |
1004 |
960 |
3.65 |
|
|
|
| 35 |
A |
975 |
933 |
0.56 |
|
|
|
| 36 |
A |
952 |
911 |
1.79 |
|
|
|
| 37 |
A |
938 |
898 |
0.95 |
|
|
|
| 38 |
A |
867 |
830 |
1.01 |
|
|
|
| 39 |
A |
822 |
786 |
0.57 |
|
|
|
| 40 |
A |
777 |
743 |
0.37 |
|
|
|
| 41 |
A |
761 |
728 |
1.82 |
|
|
|
| 42 |
A |
677 |
647 |
3.54 |
|
|
|
| 43 |
A |
419 |
400 |
0.14 |
|
|
|
| 44 |
A |
361 |
345 |
0.17 |
|
|
|
| 45 |
A |
247 |
236 |
0.06 |
|
|
|
| 46 |
A |
211 |
202 |
0.03 |
|
|
|
| 47 |
A |
131 |
125 |
0.01 |
|
|
|
| 48 |
A |
106 |
102 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37010.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 35408.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.510 |
|
|
-0.327 |
| 2 |
H |
0.164 |
|
|
0.083 |
| 3 |
H |
0.166 |
|
|
0.085 |
| 4 |
H |
0.167 |
|
|
0.077 |
| 5 |
C |
-0.310 |
|
|
0.019 |
| 6 |
H |
0.159 |
|
|
0.005 |
| 7 |
H |
0.154 |
|
|
0.049 |
| 8 |
C |
-0.328 |
|
|
0.032 |
| 9 |
H |
0.161 |
|
|
0.026 |
| 10 |
H |
0.164 |
|
|
0.028 |
| 11 |
C |
-0.324 |
|
|
-0.312 |
| 12 |
H |
0.162 |
|
|
0.098 |
| 13 |
H |
0.158 |
|
|
0.087 |
| 14 |
C |
-0.124 |
|
|
0.190 |
| 15 |
H |
0.147 |
|
|
-0.007 |
| 16 |
C |
-0.326 |
|
|
-0.289 |
| 17 |
H |
0.157 |
|
|
0.059 |
| 18 |
H |
0.161 |
|
|
0.097 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.001 |
-0.029 |
0.036 |
0.047 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.005 |
0.004 |
0.025 |
0.026 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.802 |
-0.285 |
-0.505 |
| y |
-0.285 |
-39.788 |
0.004 |
| z |
-0.505 |
0.004 |
-38.790 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.513 |
-0.285 |
-0.505 |
| y |
-0.285 |
-0.492 |
0.004 |
| z |
-0.505 |
0.004 |
1.005 |
|
| Polar |
|---|
| 3z2-r2 | 2.010 |
|---|
| x2-y2 | -0.014 |
|---|
| xy | -0.285 |
|---|
| xz | -0.505 |
|---|
| yz | 0.004 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
193.753 |
| (<r2>)1/2 |
13.920 |