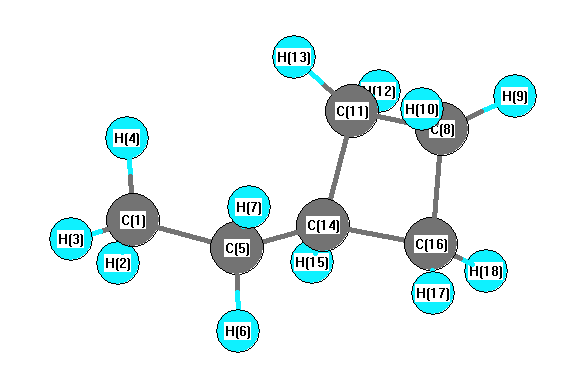
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3064 |
3013 |
77.38 |
|
|
|
| 2 |
A |
3056 |
3006 |
33.24 |
|
|
|
| 3 |
A |
3050 |
2999 |
44.10 |
|
|
|
| 4 |
A |
3045 |
2995 |
2.36 |
|
|
|
| 5 |
A |
3037 |
2987 |
14.58 |
|
|
|
| 6 |
A |
3007 |
2958 |
25.65 |
|
|
|
| 7 |
A |
2997 |
2948 |
46.53 |
|
|
|
| 8 |
A |
2992 |
2943 |
77.33 |
|
|
|
| 9 |
A |
2991 |
2942 |
22.10 |
|
|
|
| 10 |
A |
2979 |
2929 |
29.67 |
|
|
|
| 11 |
A |
2972 |
2923 |
18.25 |
|
|
|
| 12 |
A |
2953 |
2904 |
20.97 |
|
|
|
| 13 |
A |
1490 |
1465 |
3.92 |
|
|
|
| 14 |
A |
1486 |
1462 |
2.62 |
|
|
|
| 15 |
A |
1481 |
1457 |
5.56 |
|
|
|
| 16 |
A |
1467 |
1443 |
1.02 |
|
|
|
| 17 |
A |
1462 |
1438 |
1.26 |
|
|
|
| 18 |
A |
1453 |
1429 |
1.39 |
|
|
|
| 19 |
A |
1388 |
1365 |
4.32 |
|
|
|
| 20 |
A |
1360 |
1337 |
0.26 |
|
|
|
| 21 |
A |
1312 |
1290 |
7.18 |
|
|
|
| 22 |
A |
1277 |
1256 |
1.42 |
|
|
|
| 23 |
A |
1254 |
1234 |
1.31 |
|
|
|
| 24 |
A |
1248 |
1228 |
0.39 |
|
|
|
| 25 |
A |
1230 |
1210 |
0.09 |
|
|
|
| 26 |
A |
1223 |
1203 |
0.01 |
|
|
|
| 27 |
A |
1218 |
1198 |
0.14 |
|
|
|
| 28 |
A |
1161 |
1142 |
0.40 |
|
|
|
| 29 |
A |
1161 |
1142 |
0.64 |
|
|
|
| 30 |
A |
1111 |
1092 |
1.33 |
|
|
|
| 31 |
A |
1065 |
1048 |
0.07 |
|
|
|
| 32 |
A |
1012 |
995 |
0.02 |
|
|
|
| 33 |
A |
986 |
970 |
0.45 |
|
|
|
| 34 |
A |
976 |
960 |
3.75 |
|
|
|
| 35 |
A |
948 |
932 |
0.52 |
|
|
|
| 36 |
A |
925 |
909 |
1.48 |
|
|
|
| 37 |
A |
910 |
895 |
0.86 |
|
|
|
| 38 |
A |
844 |
830 |
1.03 |
|
|
|
| 39 |
A |
801 |
788 |
0.63 |
|
|
|
| 40 |
A |
755 |
742 |
0.52 |
|
|
|
| 41 |
A |
742 |
730 |
1.79 |
|
|
|
| 42 |
A |
654 |
643 |
3.76 |
|
|
|
| 43 |
A |
410 |
403 |
0.16 |
|
|
|
| 44 |
A |
353 |
347 |
0.20 |
|
|
|
| 45 |
A |
240 |
236 |
0.07 |
|
|
|
| 46 |
A |
209 |
205 |
0.04 |
|
|
|
| 47 |
A |
132 |
130 |
0.01 |
|
|
|
| 48 |
A |
106 |
104 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35995.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 35401.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.485 |
|
|
|
| 2 |
H |
0.155 |
|
|
|
| 3 |
H |
0.156 |
|
|
|
| 4 |
H |
0.158 |
|
|
|
| 5 |
C |
-0.285 |
|
|
|
| 6 |
H |
0.146 |
|
|
|
| 7 |
H |
0.142 |
|
|
|
| 8 |
C |
-0.303 |
|
|
|
| 9 |
H |
0.148 |
|
|
|
| 10 |
H |
0.152 |
|
|
|
| 11 |
C |
-0.299 |
|
|
|
| 12 |
H |
0.148 |
|
|
|
| 13 |
H |
0.145 |
|
|
|
| 14 |
C |
-0.101 |
|
|
|
| 15 |
H |
0.130 |
|
|
|
| 16 |
C |
-0.300 |
|
|
|
| 17 |
H |
0.144 |
|
|
|
| 18 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.001 |
-0.025 |
0.033 |
0.042 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.966 |
-0.282 |
-0.502 |
| y |
-0.282 |
-40.022 |
0.010 |
| z |
-0.502 |
0.010 |
-39.051 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.430 |
-0.282 |
-0.502 |
| y |
-0.282 |
-0.513 |
0.010 |
| z |
-0.502 |
0.010 |
0.943 |
|
| Polar |
|---|
| 3z2-r2 | 1.886 |
|---|
| x2-y2 | 0.055 |
|---|
| xy | -0.282 |
|---|
| xz | -0.502 |
|---|
| yz | 0.010 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.506 |
-0.015 |
-0.053 |
| y |
-0.015 |
8.950 |
0.036 |
| z |
-0.053 |
0.036 |
8.526 |
<r2> (average value of r
2) Å
2
| <r2> |
195.060 |
| (<r2>)1/2 |
13.966 |