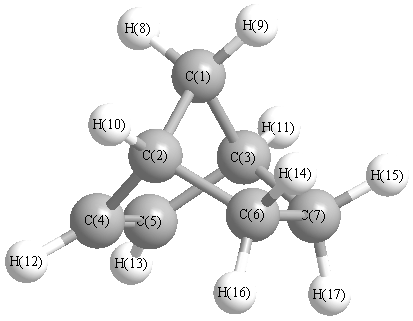
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3254 |
3088 |
20.96 |
|
|
|
| 2 |
A' |
3155 |
2995 |
40.73 |
|
|
|
| 3 |
A' |
3152 |
2992 |
44.70 |
|
|
|
| 4 |
A' |
3147 |
2987 |
8.41 |
|
|
|
| 5 |
A' |
3101 |
2943 |
50.34 |
|
|
|
| 6 |
A' |
3086 |
2930 |
35.09 |
|
|
|
| 7 |
A' |
1652 |
1568 |
0.88 |
|
|
|
| 8 |
A' |
1562 |
1483 |
0.34 |
|
|
|
| 9 |
A' |
1537 |
1459 |
1.90 |
|
|
|
| 10 |
A' |
1363 |
1294 |
0.68 |
|
|
|
| 11 |
A' |
1339 |
1271 |
0.48 |
|
|
|
| 12 |
A' |
1220 |
1158 |
0.30 |
|
|
|
| 13 |
A' |
1172 |
1112 |
4.61 |
|
|
|
| 14 |
A' |
1135 |
1077 |
0.77 |
|
|
|
| 15 |
A' |
1061 |
1007 |
1.24 |
|
|
|
| 16 |
A' |
994 |
943 |
0.35 |
|
|
|
| 17 |
A' |
969 |
920 |
0.53 |
|
|
|
| 18 |
A' |
930 |
883 |
0.71 |
|
|
|
| 19 |
A' |
901 |
855 |
5.01 |
|
|
|
| 20 |
A' |
838 |
796 |
0.34 |
|
|
|
| 21 |
A' |
789 |
749 |
2.80 |
|
|
|
| 22 |
A' |
734 |
697 |
36.64 |
|
|
|
| 23 |
A' |
484 |
459 |
0.82 |
|
|
|
| 24 |
A' |
385 |
365 |
1.99 |
|
|
|
| 25 |
A" |
3230 |
3066 |
9.59 |
|
|
|
| 26 |
A" |
3145 |
2985 |
71.59 |
|
|
|
| 27 |
A" |
3137 |
2978 |
4.64 |
|
|
|
| 28 |
A" |
3089 |
2932 |
25.58 |
|
|
|
| 29 |
A" |
1539 |
1460 |
0.37 |
|
|
|
| 30 |
A" |
1396 |
1325 |
7.02 |
|
|
|
| 31 |
A" |
1334 |
1266 |
1.33 |
|
|
|
| 32 |
A" |
1317 |
1250 |
0.00 |
|
|
|
| 33 |
A" |
1309 |
1243 |
0.48 |
|
|
|
| 34 |
A" |
1265 |
1201 |
0.41 |
|
|
|
| 35 |
A" |
1216 |
1154 |
0.13 |
|
|
|
| 36 |
A" |
1162 |
1103 |
0.30 |
|
|
|
| 37 |
A" |
1072 |
1018 |
0.86 |
|
|
|
| 38 |
A" |
981 |
931 |
0.01 |
|
|
|
| 39 |
A" |
958 |
909 |
0.02 |
|
|
|
| 40 |
A" |
928 |
881 |
4.83 |
|
|
|
| 41 |
A" |
855 |
812 |
2.23 |
|
|
|
| 42 |
A" |
812 |
771 |
1.56 |
|
|
|
| 43 |
A" |
681 |
646 |
0.53 |
|
|
|
| 44 |
A" |
487 |
463 |
0.18 |
|
|
|
| 45 |
A" |
256 |
243 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34063.4 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 32333.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.275 |
|
|
|
| 2 |
C |
-0.170 |
|
|
|
| 3 |
C |
-0.170 |
|
|
|
| 4 |
C |
-0.123 |
|
|
|
| 5 |
C |
-0.123 |
|
|
|
| 6 |
C |
-0.282 |
|
|
|
| 7 |
C |
-0.282 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
| 10 |
H |
0.136 |
|
|
|
| 11 |
H |
0.136 |
|
|
|
| 12 |
H |
0.142 |
|
|
|
| 13 |
H |
0.142 |
|
|
|
| 14 |
H |
0.145 |
|
|
|
| 15 |
H |
0.145 |
|
|
|
| 16 |
H |
0.148 |
|
|
|
| 17 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.051 |
-0.226 |
0.000 |
0.232 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.303 |
-0.364 |
0.000 |
| y |
-0.364 |
-44.781 |
0.000 |
| z |
0.000 |
0.000 |
-42.110 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.143 |
-0.364 |
0.000 |
| y |
-0.364 |
-2.574 |
0.000 |
| z |
0.000 |
0.000 |
1.432 |
|
| Polar |
|---|
| 3z2-r2 | 2.864 |
|---|
| x2-y2 | 2.478 |
|---|
| xy | -0.364 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.468 |
-0.657 |
0.000 |
| y |
-0.657 |
8.790 |
0.000 |
| z |
0.000 |
0.000 |
10.252 |
<r2> (average value of r
2) Å
2
| <r2> |
164.928 |
| (<r2>)1/2 |
12.842 |