Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -193.035201 |
| Energy at 298.15K | -193.042416 |
| HF Energy | -193.035201 |
| Nuclear repulsion energy | 128.316114 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3107 |
2972 |
25.84 |
|
|
|
| 2 |
A1 |
3047 |
2915 |
2.29 |
|
|
|
| 3 |
A1 |
1574 |
1506 |
0.01 |
|
|
|
| 4 |
A1 |
1515 |
1449 |
2.22 |
|
|
|
| 5 |
A1 |
1400 |
1340 |
4.14 |
|
|
|
| 6 |
A1 |
1073 |
1027 |
7.31 |
|
|
|
| 7 |
A1 |
956 |
914 |
27.79 |
|
|
|
| 8 |
A1 |
814 |
779 |
3.44 |
|
|
|
| 9 |
A2 |
3082 |
2949 |
0.00 |
|
|
|
| 10 |
A2 |
1246 |
1192 |
0.00 |
|
|
|
| 11 |
A2 |
1176 |
1125 |
0.00 |
|
|
|
| 12 |
A2 |
848 |
811 |
0.00 |
|
|
|
| 13 |
B1 |
3163 |
3026 |
32.44 |
|
|
|
| 14 |
B1 |
3082 |
2948 |
95.97 |
|
|
|
| 15 |
B1 |
1206 |
1154 |
1.13 |
|
|
|
| 16 |
B1 |
1159 |
1109 |
1.60 |
|
|
|
| 17 |
B1 |
775 |
741 |
0.37 |
|
|
|
| 18 |
B1 |
54 |
52 |
2.62 |
|
|
|
| 19 |
B2 |
3035 |
2904 |
165.40 |
|
|
|
| 20 |
B2 |
1542 |
1475 |
4.85 |
|
|
|
| 21 |
B2 |
1323 |
1266 |
0.36 |
|
|
|
| 22 |
B2 |
1274 |
1219 |
10.19 |
|
|
|
| 23 |
B2 |
1076 |
1029 |
105.61 |
|
|
|
| 24 |
B2 |
967 |
925 |
4.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19246.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 18413.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.069 |
| C2 |
0.000 |
0.000 |
-1.075 |
| C3 |
0.000 |
1.031 |
0.065 |
| C4 |
0.000 |
-1.031 |
0.065 |
| H5 |
0.891 |
0.000 |
-1.708 |
| H6 |
-0.891 |
0.000 |
-1.708 |
| H7 |
0.893 |
1.667 |
0.133 |
| H8 |
-0.893 |
1.667 |
0.133 |
| H9 |
-0.893 |
-1.667 |
0.133 |
| H10 |
0.893 |
-1.667 |
0.133 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1439 | 1.4388 | 1.4388 | 2.9162 | 2.9162 | 2.1103 | 2.1103 | 2.1103 | 2.1103 |
C2 | 2.1439 | | 1.5374 | 1.5374 | 1.0929 | 1.0929 | 2.2440 | 2.2440 | 2.2440 | 2.2440 | C3 | 1.4388 | 1.5374 | | 2.0621 | 2.2364 | 2.2364 | 1.0983 | 1.0983 | 2.8431 | 2.8431 | C4 | 1.4388 | 1.5374 | 2.0621 | | 2.2364 | 2.2364 | 2.8431 | 2.8431 | 1.0983 | 1.0983 | H5 | 2.9162 | 1.0929 | 2.2364 | 2.2364 | | 1.7820 | 2.4836 | 3.0577 | 3.0577 | 2.4836 | H6 | 2.9162 | 1.0929 | 2.2364 | 2.2364 | 1.7820 | | 3.0577 | 2.4836 | 2.4836 | 3.0577 | H7 | 2.1103 | 2.2440 | 1.0983 | 2.8431 | 2.4836 | 3.0577 | | 1.7852 | 3.7826 | 3.3348 | H8 | 2.1103 | 2.2440 | 1.0983 | 2.8431 | 3.0577 | 2.4836 | 1.7852 | | 3.3348 | 3.7826 | H9 | 2.1103 | 2.2440 | 2.8431 | 1.0983 | 3.0577 | 2.4836 | 3.7826 | 3.3348 | | 1.7852 | H10 | 2.1103 | 2.2440 | 2.8431 | 1.0983 | 2.4836 | 3.0577 | 3.3348 | 3.7826 | 1.7852 | |
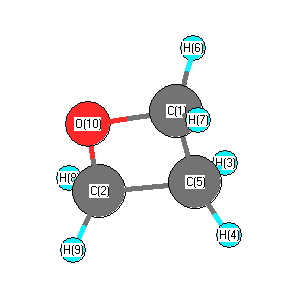 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.106 |
|
O1 |
C3 |
H7 |
111.866 |
| O1 |
C3 |
H8 |
111.866 |
|
O1 |
C4 |
C2 |
92.106 |
| O1 |
C4 |
H9 |
111.866 |
|
O1 |
C4 |
H10 |
111.866 |
| C2 |
C3 |
H7 |
115.725 |
|
C2 |
C3 |
H8 |
115.725 |
| C2 |
C4 |
H9 |
115.725 |
|
C2 |
C4 |
H10 |
115.725 |
| C3 |
O1 |
C4 |
91.553 |
|
C3 |
C2 |
C4 |
84.234 |
| C3 |
C2 |
H5 |
115.439 |
|
C3 |
C2 |
H6 |
115.439 |
| C4 |
C2 |
H5 |
115.439 |
|
C4 |
C2 |
H6 |
115.439 |
| H5 |
C2 |
H6 |
109.228 |
|
H7 |
C3 |
H8 |
108.728 |
| H9 |
C4 |
H10 |
108.728 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.497 |
|
|
|
| 2 |
C |
-0.420 |
|
|
|
| 3 |
C |
-0.040 |
|
|
|
| 4 |
C |
-0.040 |
|
|
|
| 5 |
H |
0.180 |
|
|
|
| 6 |
H |
0.180 |
|
|
|
| 7 |
H |
0.159 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
H |
0.159 |
|
|
|
| 10 |
H |
0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.984 |
1.984 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.879 |
0.000 |
0.000 |
| y |
0.000 |
-22.620 |
0.000 |
| z |
0.000 |
0.000 |
-27.316 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.089 |
0.000 |
0.000 |
| y |
0.000 |
2.977 |
0.000 |
| z |
0.000 |
0.000 |
-4.067 |
|
| Polar |
|---|
| 3z2-r2 | -8.133 |
|---|
| x2-y2 | -1.258 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.652 |
0.000 |
0.000 |
| y |
0.000 |
5.573 |
0.000 |
| z |
0.000 |
0.000 |
4.453 |
<r2> (average value of r
2) Å
2
| <r2> |
66.050 |
| (<r2>)1/2 |
8.127 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -193.035200 |
| Energy at 298.15K | -193.042417 |
| HF Energy | -193.035200 |
| Nuclear repulsion energy | 128.315879 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3163 |
3026 |
32.50 |
|
|
|
| 2 |
A' |
3107 |
2972 |
25.89 |
|
|
|
| 3 |
A' |
3082 |
2948 |
95.96 |
|
|
|
| 4 |
A' |
3047 |
2915 |
2.29 |
|
|
|
| 5 |
A' |
1574 |
1506 |
0.01 |
|
|
|
| 6 |
A' |
1515 |
1449 |
2.22 |
|
|
|
| 7 |
A' |
1400 |
1340 |
4.14 |
|
|
|
| 8 |
A' |
1206 |
1154 |
1.14 |
|
|
|
| 9 |
A' |
1158 |
1108 |
1.60 |
|
|
|
| 10 |
A' |
1073 |
1027 |
7.32 |
|
|
|
| 11 |
A' |
956 |
914 |
27.78 |
|
|
|
| 12 |
A' |
814 |
779 |
3.42 |
|
|
|
| 13 |
A' |
775 |
741 |
0.37 |
|
|
|
| 14 |
A' |
55 |
53 |
2.62 |
|
|
|
| 15 |
A" |
3082 |
2949 |
0.00 |
|
|
|
| 16 |
A" |
3035 |
2904 |
165.39 |
|
|
|
| 17 |
A" |
1542 |
1475 |
4.87 |
|
|
|
| 18 |
A" |
1323 |
1266 |
0.37 |
|
|
|
| 19 |
A" |
1275 |
1219 |
10.17 |
|
|
|
| 20 |
A" |
1246 |
1192 |
0.00 |
|
|
|
| 21 |
A" |
1176 |
1125 |
0.00 |
|
|
|
| 22 |
A" |
1076 |
1029 |
105.62 |
|
|
|
| 23 |
A" |
967 |
925 |
4.29 |
|
|
|
| 24 |
A" |
848 |
811 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19246.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 18413.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.001 |
-1.069 |
0.000 |
| C2 |
0.000 |
1.075 |
0.000 |
| C3 |
0.000 |
-0.065 |
1.031 |
| C4 |
0.000 |
-0.065 |
-1.031 |
| H5 |
0.891 |
1.709 |
0.000 |
| H6 |
-0.891 |
1.707 |
0.000 |
| H7 |
0.894 |
-0.133 |
1.667 |
| H8 |
-0.892 |
-0.132 |
1.669 |
| H9 |
0.894 |
-0.133 |
-1.667 |
| H10 |
-0.892 |
-0.132 |
-1.669 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1435 | 1.4387 | 1.4387 | 2.9169 | 2.9148 | 2.1107 | 2.1105 | 2.1107 | 2.1105 |
C2 | 2.1435 | | 1.5375 | 1.5375 | 1.0929 | 1.0928 | 2.2436 | 2.2442 | 2.2436 | 2.2442 | C3 | 1.4387 | 1.5375 | | 2.0625 | 2.2368 | 2.2360 | 1.0983 | 1.0983 | 2.8428 | 2.8442 | C4 | 1.4387 | 1.5375 | 2.0625 | | 2.2368 | 2.2360 | 2.8428 | 2.8442 | 1.0983 | 1.0983 | H5 | 2.9169 | 1.0929 | 2.2368 | 2.2368 | | 1.7818 | 2.4835 | 3.0576 | 2.4835 | 3.0576 | H6 | 2.9148 | 1.0928 | 2.2360 | 2.2360 | 1.7818 | | 3.0573 | 2.4832 | 3.0573 | 2.4832 | H7 | 2.1107 | 2.2436 | 1.0983 | 2.8428 | 2.4835 | 3.0573 | | 1.7852 | 3.3335 | 3.7831 | H8 | 2.1105 | 2.2442 | 1.0983 | 2.8442 | 3.0576 | 2.4832 | 1.7852 | | 3.7831 | 3.3372 | H9 | 2.1107 | 2.2436 | 2.8428 | 1.0983 | 2.4835 | 3.0573 | 3.3335 | 3.7831 | | 1.7852 | H10 | 2.1105 | 2.2442 | 2.8442 | 1.0983 | 3.0576 | 2.4832 | 3.7831 | 3.3372 | 1.7852 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.083 |
|
O1 |
C3 |
H7 |
111.904 |
| O1 |
C3 |
H8 |
111.886 |
|
O1 |
C4 |
C2 |
92.083 |
| O1 |
C4 |
H9 |
111.904 |
|
O1 |
C4 |
H10 |
111.886 |
| C2 |
C3 |
H7 |
115.686 |
|
C2 |
C3 |
H8 |
115.735 |
| C2 |
C4 |
H9 |
115.686 |
|
C2 |
C4 |
H10 |
115.735 |
| C3 |
O1 |
C4 |
91.581 |
|
C3 |
C2 |
C4 |
84.252 |
| C3 |
C2 |
H5 |
115.472 |
|
C3 |
C2 |
H6 |
115.409 |
| C4 |
C2 |
H5 |
115.472 |
|
C4 |
C2 |
H6 |
115.409 |
| H5 |
C2 |
H6 |
109.212 |
|
H7 |
C3 |
H8 |
108.726 |
| H9 |
C4 |
H10 |
108.726 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.496 |
|
|
|
| 2 |
C |
-0.420 |
|
|
|
| 3 |
C |
-0.040 |
|
|
|
| 4 |
C |
-0.040 |
|
|
|
| 5 |
H |
0.180 |
|
|
|
| 6 |
H |
0.180 |
|
|
|
| 7 |
H |
0.159 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
H |
0.159 |
|
|
|
| 10 |
H |
0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.002 |
1.984 |
0.000 |
1.984 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.880 |
-0.002 |
0.000 |
| y |
-0.002 |
-27.316 |
0.000 |
| z |
0.000 |
0.000 |
-22.618 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.087 |
-0.002 |
0.000 |
| y |
-0.002 |
-4.068 |
0.000 |
| z |
0.000 |
0.000 |
2.980 |
|
| Polar |
|---|
| 3z2-r2 | 5.960 |
|---|
| x2-y2 | 3.437 |
|---|
| xy | -0.002 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.652 |
0.001 |
0.000 |
| y |
0.001 |
4.452 |
0.000 |
| z |
0.000 |
0.000 |
5.574 |
<r2> (average value of r
2) Å
2
| <r2> |
66.051 |
| (<r2>)1/2 |
8.127 |
