Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -192.884180 |
| Energy at 298.15K | |
| HF Energy | -192.698463 |
| Nuclear repulsion energy | 128.021856 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3128 |
2969 |
24.74 |
|
|
|
| 2 |
A1 |
3078 |
2921 |
2.21 |
|
|
|
| 3 |
A1 |
1598 |
1516 |
0.00 |
|
|
|
| 4 |
A1 |
1541 |
1463 |
1.64 |
|
|
|
| 5 |
A1 |
1413 |
1341 |
5.03 |
|
|
|
| 6 |
A1 |
1066 |
1012 |
5.04 |
|
|
|
| 7 |
A1 |
943 |
895 |
28.21 |
|
|
|
| 8 |
A1 |
818 |
776 |
4.11 |
|
|
|
| 9 |
A2 |
3117 |
2958 |
0.00 |
|
|
|
| 10 |
A2 |
1263 |
1199 |
0.00 |
|
|
|
| 11 |
A2 |
1183 |
1123 |
0.00 |
|
|
|
| 12 |
A2 |
859 |
815 |
0.00 |
|
|
|
| 13 |
B1 |
3184 |
3022 |
36.45 |
|
|
|
| 14 |
B1 |
3116 |
2957 |
86.97 |
|
|
|
| 15 |
B1 |
1222 |
1159 |
1.41 |
|
|
|
| 16 |
B1 |
1167 |
1108 |
1.66 |
|
|
|
| 17 |
B1 |
784 |
744 |
0.15 |
|
|
|
| 18 |
B1 |
18i |
17i |
2.92 |
|
|
|
| 19 |
B2 |
3068 |
2912 |
150.44 |
|
|
|
| 20 |
B2 |
1567 |
1488 |
5.80 |
|
|
|
| 21 |
B2 |
1336 |
1268 |
0.70 |
|
|
|
| 22 |
B2 |
1293 |
1227 |
7.87 |
|
|
|
| 23 |
B2 |
1056 |
1002 |
97.86 |
|
|
|
| 24 |
B2 |
967 |
918 |
6.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19372.1 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 18388.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.074 |
| C2 |
0.000 |
0.000 |
-1.076 |
| C3 |
0.000 |
1.036 |
0.063 |
| C4 |
0.000 |
-1.036 |
0.063 |
| H5 |
0.889 |
0.000 |
-1.707 |
| H6 |
-0.889 |
0.000 |
-1.707 |
| H7 |
0.891 |
1.669 |
0.131 |
| H8 |
-0.891 |
1.669 |
0.131 |
| H9 |
-0.891 |
-1.669 |
0.131 |
| H10 |
0.891 |
-1.669 |
0.131 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1491 | 1.4473 | 1.4473 | 2.9195 | 2.9195 | 2.1139 | 2.1139 | 2.1139 | 2.1139 |
C2 | 2.1491 | | 1.5394 | 1.5394 | 1.0909 | 1.0909 | 2.2441 | 2.2441 | 2.2441 | 2.2441 | C3 | 1.4473 | 1.5394 | | 2.0721 | 2.2356 | 2.2356 | 1.0954 | 1.0954 | 2.8489 | 2.8489 | C4 | 1.4473 | 1.5394 | 2.0721 | | 2.2356 | 2.2356 | 2.8489 | 2.8489 | 1.0954 | 1.0954 | H5 | 2.9195 | 1.0909 | 2.2356 | 2.2356 | | 1.7789 | 2.4828 | 3.0554 | 3.0554 | 2.4828 | H6 | 2.9195 | 1.0909 | 2.2356 | 2.2356 | 1.7789 | | 3.0554 | 2.4828 | 2.4828 | 3.0554 | H7 | 2.1139 | 2.2441 | 1.0954 | 2.8489 | 2.4828 | 3.0554 | | 1.7829 | 3.7842 | 3.3379 | H8 | 2.1139 | 2.2441 | 1.0954 | 2.8489 | 3.0554 | 2.4828 | 1.7829 | | 3.3379 | 3.7842 | H9 | 2.1139 | 2.2441 | 2.8489 | 1.0954 | 3.0554 | 2.4828 | 3.7842 | 3.3379 | | 1.7829 | H10 | 2.1139 | 2.2441 | 2.8489 | 1.0954 | 2.4828 | 3.0554 | 3.3379 | 3.7842 | 1.7829 | |
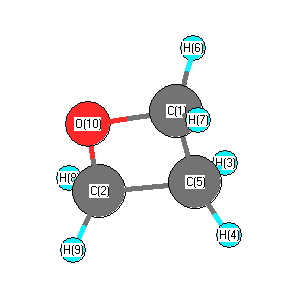 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.987 |
|
O1 |
C3 |
H7 |
111.734 |
| O1 |
C3 |
H8 |
111.734 |
|
O1 |
C4 |
C2 |
91.987 |
| O1 |
C4 |
H9 |
111.734 |
|
O1 |
C4 |
H10 |
111.734 |
| C2 |
C3 |
H7 |
115.769 |
|
C2 |
C3 |
H8 |
115.769 |
| C2 |
C4 |
H9 |
115.769 |
|
C2 |
C4 |
H10 |
115.769 |
| C3 |
O1 |
C4 |
91.426 |
|
C3 |
C2 |
C4 |
84.600 |
| C3 |
C2 |
H5 |
115.356 |
|
C3 |
C2 |
H6 |
115.356 |
| C4 |
C2 |
H5 |
115.356 |
|
C4 |
C2 |
H6 |
115.356 |
| H5 |
C2 |
H6 |
109.241 |
|
H7 |
C3 |
H8 |
108.943 |
| H9 |
C4 |
H10 |
108.943 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.517 |
|
|
|
| 2 |
C |
-0.380 |
|
|
|
| 3 |
C |
-0.005 |
|
|
|
| 4 |
C |
-0.005 |
|
|
|
| 5 |
H |
0.164 |
|
|
|
| 6 |
H |
0.164 |
|
|
|
| 7 |
H |
0.145 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
H |
0.145 |
|
|
|
| 10 |
H |
0.145 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.044 |
2.044 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.180 |
0.000 |
0.000 |
| y |
0.000 |
-22.814 |
0.000 |
| z |
0.000 |
0.000 |
-27.671 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.063 |
0.000 |
0.000 |
| y |
0.000 |
3.112 |
0.000 |
| z |
0.000 |
0.000 |
-4.175 |
|
| Polar |
|---|
| 3z2-r2 | -8.349 |
|---|
| x2-y2 | -1.366 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.594 |
0.000 |
0.000 |
| y |
0.000 |
5.514 |
0.000 |
| z |
0.000 |
0.000 |
4.408 |
<r2> (average value of r
2) Å
2
| <r2> |
66.434 |
| (<r2>)1/2 |
8.151 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -192.884180 |
| Energy at 298.15K | -192.891339 |
| HF Energy | -192.698453 |
| Nuclear repulsion energy | 128.031037 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3184 |
3022 |
36.61 |
|
|
|
| 2 |
A' |
3128 |
2969 |
25.29 |
|
|
|
| 3 |
A' |
3116 |
2958 |
84.34 |
|
|
|
| 4 |
A' |
3077 |
2921 |
4.35 |
|
|
|
| 5 |
A' |
1597 |
1516 |
0.01 |
|
|
|
| 6 |
A' |
1541 |
1463 |
1.64 |
|
|
|
| 7 |
A' |
1413 |
1341 |
5.02 |
|
|
|
| 8 |
A' |
1222 |
1160 |
1.39 |
|
|
|
| 9 |
A' |
1168 |
1108 |
1.71 |
|
|
|
| 10 |
A' |
1066 |
1012 |
5.09 |
|
|
|
| 11 |
A' |
942 |
895 |
28.09 |
|
|
|
| 12 |
A' |
823 |
781 |
4.06 |
|
|
|
| 13 |
A' |
779 |
739 |
0.40 |
|
|
|
| 14 |
A' |
20 |
19 |
2.95 |
|
|
|
| 15 |
A" |
3117 |
2959 |
1.62 |
|
|
|
| 16 |
A" |
3068 |
2912 |
148.46 |
|
|
|
| 17 |
A" |
1567 |
1488 |
5.78 |
|
|
|
| 18 |
A" |
1336 |
1268 |
0.68 |
|
|
|
| 19 |
A" |
1293 |
1228 |
7.73 |
|
|
|
| 20 |
A" |
1262 |
1198 |
0.16 |
|
|
|
| 21 |
A" |
1182 |
1122 |
0.03 |
|
|
|
| 22 |
A" |
1055 |
1002 |
97.61 |
|
|
|
| 23 |
A" |
968 |
918 |
6.49 |
|
|
|
| 24 |
A" |
858 |
815 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19391.2 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 18406.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.045 |
-1.073 |
0.000 |
| C2 |
-0.012 |
1.075 |
0.000 |
| C3 |
-0.012 |
-0.063 |
1.036 |
| C4 |
-0.012 |
-0.063 |
-1.036 |
| H5 |
0.884 |
1.698 |
0.000 |
| H6 |
-0.895 |
1.716 |
0.000 |
| H7 |
0.858 |
-0.112 |
1.698 |
| H8 |
-0.923 |
-0.151 |
1.638 |
| H9 |
0.858 |
-0.112 |
-1.698 |
| H10 |
-0.923 |
-0.151 |
-1.638 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1487 | 1.4473 | 1.4473 | 2.8946 | 2.9427 | 2.1140 | 2.1139 | 2.1140 | 2.1139 |
C2 | 2.1487 | | 1.5393 | 1.5393 | 1.0909 | 1.0909 | 2.2479 | 2.2401 | 2.2479 | 2.2401 | C3 | 1.4473 | 1.5393 | | 2.0716 | 2.2309 | 2.2403 | 1.0951 | 1.0957 | 2.8699 | 2.8262 | C4 | 1.4473 | 1.5393 | 2.0716 | | 2.2309 | 2.2403 | 2.8699 | 2.8262 | 1.0951 | 1.0957 | H5 | 2.8946 | 1.0909 | 2.2309 | 2.2309 | | 1.7789 | 2.4823 | 3.0607 | 2.4823 | 3.0607 | H6 | 2.9427 | 1.0909 | 2.2403 | 2.2403 | 1.7789 | | 3.0499 | 2.4842 | 3.0499 | 2.4842 | H7 | 2.1140 | 2.2479 | 1.0951 | 2.8699 | 2.4823 | 3.0499 | | 1.7830 | 3.3969 | 3.7826 | H8 | 2.1139 | 2.2401 | 1.0957 | 2.8262 | 3.0607 | 2.4842 | 1.7830 | | 3.7826 | 3.2761 | H9 | 2.1140 | 2.2479 | 2.8699 | 1.0951 | 2.4823 | 3.0499 | 3.3969 | 3.7826 | | 1.7830 | H10 | 2.1139 | 2.2401 | 2.8262 | 1.0957 | 3.0607 | 2.4842 | 3.7826 | 3.2761 | 1.7830 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.964 |
|
O1 |
C3 |
H7 |
111.762 |
| O1 |
C3 |
H8 |
111.715 |
|
O1 |
C4 |
C2 |
91.964 |
| O1 |
C4 |
H9 |
111.762 |
|
O1 |
C4 |
H10 |
111.715 |
| C2 |
C3 |
H7 |
116.121 |
|
C2 |
C3 |
H8 |
115.420 |
| C2 |
C4 |
H9 |
116.121 |
|
C2 |
C4 |
H10 |
115.420 |
| C3 |
O1 |
C4 |
91.397 |
|
C3 |
C2 |
C4 |
84.582 |
| C3 |
C2 |
H5 |
114.964 |
|
C3 |
C2 |
H6 |
115.756 |
| C4 |
C2 |
H5 |
114.964 |
|
C4 |
C2 |
H6 |
115.756 |
| H5 |
C2 |
H6 |
109.238 |
|
H7 |
C3 |
H8 |
108.950 |
| H9 |
C4 |
H10 |
108.950 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.517 |
|
|
|
| 2 |
C |
-0.380 |
|
|
|
| 3 |
C |
-0.005 |
|
|
|
| 4 |
C |
-0.005 |
|
|
|
| 5 |
H |
0.164 |
|
|
|
| 6 |
H |
0.163 |
|
|
|
| 7 |
H |
0.146 |
|
|
|
| 8 |
H |
0.144 |
|
|
|
| 9 |
H |
0.146 |
|
|
|
| 10 |
H |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.087 |
2.041 |
0.000 |
2.043 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.188 |
0.138 |
0.000 |
| y |
0.138 |
-27.666 |
0.000 |
| z |
0.000 |
0.000 |
-22.811 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.050 |
0.138 |
0.000 |
| y |
0.138 |
-4.166 |
0.000 |
| z |
0.000 |
0.000 |
3.117 |
|
| Polar |
|---|
| 3z2-r2 | 6.233 |
|---|
| x2-y2 | 3.478 |
|---|
| xy | 0.138 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.596 |
0.001 |
0.000 |
| y |
0.001 |
4.409 |
0.000 |
| z |
0.000 |
0.000 |
5.514 |
<r2> (average value of r
2) Å
2
| <r2> |
66.422 |
| (<r2>)1/2 |
8.150 |
