Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -192.860098 |
| Energy at 298.15K | -192.867174 |
| HF Energy | -192.860098 |
| Nuclear repulsion energy | 127.376073 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3030 |
2980 |
26.58 |
|
|
|
| 2 |
A1 |
2958 |
2909 |
1.50 |
|
|
|
| 3 |
A1 |
1528 |
1503 |
0.02 |
|
|
|
| 4 |
A1 |
1471 |
1447 |
1.81 |
|
|
|
| 5 |
A1 |
1353 |
1330 |
3.52 |
|
|
|
| 6 |
A1 |
1039 |
1022 |
4.24 |
|
|
|
| 7 |
A1 |
917 |
902 |
24.96 |
|
|
|
| 8 |
A1 |
783 |
770 |
3.06 |
|
|
|
| 9 |
A2 |
2988 |
2938 |
0.00 |
|
|
|
| 10 |
A2 |
1208 |
1188 |
0.00 |
|
|
|
| 11 |
A2 |
1138 |
1119 |
0.00 |
|
|
|
| 12 |
A2 |
825 |
812 |
0.00 |
|
|
|
| 13 |
B1 |
3086 |
3035 |
28.54 |
|
|
|
| 14 |
B1 |
2987 |
2937 |
104.75 |
|
|
|
| 15 |
B1 |
1164 |
1144 |
1.53 |
|
|
|
| 16 |
B1 |
1115 |
1097 |
0.53 |
|
|
|
| 17 |
B1 |
753 |
741 |
0.37 |
|
|
|
| 18 |
B1 |
25 |
25 |
1.76 |
|
|
|
| 19 |
B2 |
2945 |
2896 |
175.81 |
|
|
|
| 20 |
B2 |
1496 |
1472 |
5.88 |
|
|
|
| 21 |
B2 |
1276 |
1255 |
0.50 |
|
|
|
| 22 |
B2 |
1228 |
1208 |
9.08 |
|
|
|
| 23 |
B2 |
1027 |
1010 |
97.50 |
|
|
|
| 24 |
B2 |
943 |
928 |
8.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18640.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 18332.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.080 |
| C2 |
0.000 |
0.000 |
-1.080 |
| C3 |
0.000 |
1.037 |
0.063 |
| C4 |
0.000 |
-1.037 |
0.063 |
| H5 |
0.897 |
0.000 |
-1.718 |
| H6 |
-0.897 |
0.000 |
-1.718 |
| H7 |
0.899 |
1.680 |
0.129 |
| H8 |
-0.899 |
1.680 |
0.129 |
| H9 |
-0.899 |
-1.680 |
0.129 |
| H10 |
0.899 |
-1.680 |
0.129 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1608 | 1.4528 | 1.4528 | 2.9388 | 2.9388 | 2.1299 | 2.1299 | 2.1299 | 2.1299 |
C2 | 2.1608 | | 1.5438 | 1.5438 | 1.1007 | 1.1007 | 2.2573 | 2.2573 | 2.2573 | 2.2573 | C3 | 1.4528 | 1.5438 | | 2.0744 | 2.2479 | 2.2479 | 1.1076 | 1.1076 | 2.8631 | 2.8631 | C4 | 1.4528 | 1.5438 | 2.0744 | | 2.2479 | 2.2479 | 2.8631 | 2.8631 | 1.1076 | 1.1076 | H5 | 2.9388 | 1.1007 | 2.2479 | 2.2479 | | 1.7944 | 2.4971 | 3.0763 | 3.0763 | 2.4971 | H6 | 2.9388 | 1.1007 | 2.2479 | 2.2479 | 1.7944 | | 3.0763 | 2.4971 | 2.4971 | 3.0763 | H7 | 2.1299 | 2.2573 | 1.1076 | 2.8631 | 2.4971 | 3.0763 | | 1.7988 | 3.8115 | 3.3603 | H8 | 2.1299 | 2.2573 | 1.1076 | 2.8631 | 3.0763 | 2.4971 | 1.7988 | | 3.3603 | 3.8115 | H9 | 2.1299 | 2.2573 | 2.8631 | 1.1076 | 3.0763 | 2.4971 | 3.8115 | 3.3603 | | 1.7988 | H10 | 2.1299 | 2.2573 | 2.8631 | 1.1076 | 2.4971 | 3.0763 | 3.3603 | 3.8115 | 1.7988 | |
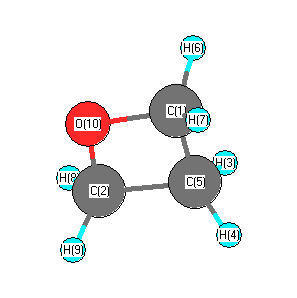 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.235 |
|
O1 |
C3 |
H7 |
111.874 |
| O1 |
C3 |
H8 |
111.874 |
|
O1 |
C4 |
C2 |
92.235 |
| O1 |
C4 |
H9 |
111.874 |
|
O1 |
C4 |
H10 |
111.874 |
| C2 |
C3 |
H7 |
115.741 |
|
C2 |
C3 |
H8 |
115.741 |
| C2 |
C4 |
H9 |
115.741 |
|
C2 |
C4 |
H10 |
115.741 |
| C3 |
O1 |
C4 |
91.109 |
|
C3 |
C2 |
C4 |
84.421 |
| C3 |
C2 |
H5 |
115.411 |
|
C3 |
C2 |
H6 |
115.411 |
| C4 |
C2 |
H5 |
115.411 |
|
C4 |
C2 |
H6 |
115.411 |
| H5 |
C2 |
H6 |
109.193 |
|
H7 |
C3 |
H8 |
108.594 |
| H9 |
C4 |
H10 |
108.594 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.451 |
|
|
|
| 2 |
C |
-0.384 |
|
|
|
| 3 |
C |
-0.044 |
|
|
|
| 4 |
C |
-0.044 |
|
|
|
| 5 |
H |
0.167 |
|
|
|
| 6 |
H |
0.167 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.879 |
1.879 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.074 |
0.000 |
0.000 |
| y |
0.000 |
-22.944 |
0.000 |
| z |
0.000 |
0.000 |
-27.289 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.042 |
0.000 |
0.000 |
| y |
0.000 |
2.738 |
0.000 |
| z |
0.000 |
0.000 |
-3.780 |
|
| Polar |
|---|
| 3z2-r2 | -7.559 |
|---|
| x2-y2 | -1.131 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.774 |
0.000 |
0.000 |
| y |
0.000 |
5.778 |
0.000 |
| z |
0.000 |
0.000 |
4.570 |
<r2> (average value of r
2) Å
2
| <r2> |
66.877 |
| (<r2>)1/2 |
8.178 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -192.860099 |
| Energy at 298.15K | -192.867173 |
| HF Energy | -192.860099 |
| Nuclear repulsion energy | 127.377609 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3085 |
3034 |
28.66 |
|
|
|
| 2 |
A' |
3030 |
2980 |
26.61 |
|
|
|
| 3 |
A' |
2987 |
2938 |
104.62 |
|
|
|
| 4 |
A' |
2958 |
2909 |
1.50 |
|
|
|
| 5 |
A' |
1528 |
1503 |
0.02 |
|
|
|
| 6 |
A' |
1471 |
1447 |
1.81 |
|
|
|
| 7 |
A' |
1353 |
1330 |
3.52 |
|
|
|
| 8 |
A' |
1164 |
1144 |
1.53 |
|
|
|
| 9 |
A' |
1115 |
1097 |
0.53 |
|
|
|
| 10 |
A' |
1039 |
1022 |
4.23 |
|
|
|
| 11 |
A' |
917 |
902 |
24.97 |
|
|
|
| 12 |
A' |
783 |
770 |
3.06 |
|
|
|
| 13 |
A' |
753 |
740 |
0.37 |
|
|
|
| 14 |
A' |
25 |
24 |
1.76 |
|
|
|
| 15 |
A" |
2988 |
2939 |
0.00 |
|
|
|
| 16 |
A" |
2945 |
2897 |
175.77 |
|
|
|
| 17 |
A" |
1496 |
1472 |
5.88 |
|
|
|
| 18 |
A" |
1276 |
1255 |
0.50 |
|
|
|
| 19 |
A" |
1228 |
1208 |
9.07 |
|
|
|
| 20 |
A" |
1208 |
1188 |
0.00 |
|
|
|
| 21 |
A" |
1138 |
1119 |
0.00 |
|
|
|
| 22 |
A" |
1027 |
1010 |
97.49 |
|
|
|
| 23 |
A" |
943 |
928 |
8.17 |
|
|
|
| 24 |
A" |
825 |
812 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18640.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 18332.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
-1.080 |
0.000 |
| C2 |
-0.000 |
1.080 |
0.000 |
| C3 |
-0.000 |
-0.063 |
1.037 |
| C4 |
-0.000 |
-0.063 |
-1.037 |
| H5 |
0.897 |
1.718 |
0.000 |
| H6 |
-0.897 |
1.718 |
0.000 |
| H7 |
0.899 |
-0.129 |
1.680 |
| H8 |
-0.900 |
-0.129 |
1.680 |
| H9 |
0.899 |
-0.129 |
-1.680 |
| H10 |
-0.900 |
-0.129 |
-1.680 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1607 | 1.4528 | 1.4528 | 2.9387 | 2.9390 | 2.1299 | 2.1299 | 2.1299 | 2.1299 |
C2 | 2.1607 | | 1.5438 | 1.5438 | 1.1007 | 1.1007 | 2.2572 | 2.2571 | 2.2572 | 2.2571 | C3 | 1.4528 | 1.5438 | | 2.0745 | 2.2480 | 2.2480 | 1.1075 | 1.1076 | 2.8633 | 2.8629 | C4 | 1.4528 | 1.5438 | 2.0745 | | 2.2480 | 2.2480 | 2.8633 | 2.8629 | 1.1075 | 1.1076 | H5 | 2.9387 | 1.1007 | 2.2480 | 2.2480 | | 1.7942 | 2.4972 | 3.0763 | 2.4972 | 3.0763 | H6 | 2.9390 | 1.1007 | 2.2480 | 2.2480 | 1.7942 | | 3.0762 | 2.4971 | 3.0762 | 2.4971 | H7 | 2.1299 | 2.2572 | 1.1075 | 2.8633 | 2.4972 | 3.0762 | | 1.7989 | 3.3607 | 3.8115 | H8 | 2.1299 | 2.2571 | 1.1076 | 2.8629 | 3.0763 | 2.4971 | 1.7989 | | 3.8115 | 3.3598 | H9 | 2.1299 | 2.2572 | 2.8633 | 1.1075 | 2.4972 | 3.0762 | 3.3607 | 3.8115 | | 1.7989 | H10 | 2.1299 | 2.2571 | 2.8629 | 1.1076 | 3.0763 | 2.4971 | 3.8115 | 3.3598 | 1.7989 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.231 |
|
O1 |
C3 |
H7 |
111.876 |
| O1 |
C3 |
H8 |
111.877 |
|
O1 |
C4 |
C2 |
92.231 |
| O1 |
C4 |
H9 |
111.876 |
|
O1 |
C4 |
H10 |
111.877 |
| C2 |
C3 |
H7 |
115.740 |
|
C2 |
C3 |
H8 |
115.733 |
| C2 |
C4 |
H9 |
115.740 |
|
C2 |
C4 |
H10 |
115.733 |
| C3 |
O1 |
C4 |
91.112 |
|
C3 |
C2 |
C4 |
84.425 |
| C3 |
C2 |
H5 |
115.416 |
|
C3 |
C2 |
H6 |
115.419 |
| C4 |
C2 |
H5 |
115.416 |
|
C4 |
C2 |
H6 |
115.419 |
| H5 |
C2 |
H6 |
109.170 |
|
H7 |
C3 |
H8 |
108.601 |
| H9 |
C4 |
H10 |
108.601 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.451 |
|
|
|
| 2 |
C |
-0.384 |
|
|
|
| 3 |
C |
-0.044 |
|
|
|
| 4 |
C |
-0.044 |
|
|
|
| 5 |
H |
0.167 |
|
|
|
| 6 |
H |
0.167 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.001 |
1.879 |
0.000 |
1.879 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.075 |
0.001 |
0.000 |
| y |
0.001 |
-27.288 |
0.000 |
| z |
0.000 |
0.000 |
-22.943 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.041 |
0.001 |
0.000 |
| y |
0.001 |
-3.779 |
0.000 |
| z |
0.000 |
0.000 |
2.738 |
|
| Polar |
|---|
| 3z2-r2 | 5.476 |
|---|
| x2-y2 | 3.214 |
|---|
| xy | 0.001 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.773 |
0.000 |
0.000 |
| y |
0.000 |
4.570 |
0.000 |
| z |
0.000 |
0.000 |
5.778 |
<r2> (average value of r
2) Å
2
| <r2> |
66.876 |
| (<r2>)1/2 |
8.178 |
