return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8O2 (1,3-Propanediol)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-268.758025
Energy at 298.15K-268.768100
Nuclear repulsion energy194.390687
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3778 3598 18.99      
2 A 3756 3576 102.84      
3 A 3122 2973 31.61      
4 A 3112 2964 48.04      
5 A 3070 2924 61.34      
6 A 3061 2915 17.07      
7 A 3026 2881 53.27      
8 A 2998 2855 61.92      
9 A 1579 1503 1.93      
10 A 1565 1490 0.16      
11 A 1523 1450 15.88      
12 A 1506 1434 82.99      
13 A 1490 1419 16.08      
14 A 1436 1367 1.28      
15 A 1417 1350 0.26      
16 A 1342 1278 34.65      
17 A 1309 1247 2.24      
18 A 1251 1191 21.28      
19 A 1230 1172 54.07      
20 A 1167 1111 27.12      
21 A 1135 1081 13.33      
22 A 1126 1072 106.51      
23 A 991 943 2.17      
24 A 941 897 5.07      
25 A 933 889 8.84      
26 A 843 803 8.95      
27 A 549 523 178.56      
28 A 519 494 3.25      
29 A 397 378 4.61      
30 A 337 321 25.51      
31 A 289 276 76.83      
32 A 221 210 2.83      
33 A 108 103 1.36      

Unscaled Zero Point Vibrational Energy (zpe) 25562.5 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 24343.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.25603 0.13100 0.09583

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.019 1.039 -0.348
C2 1.295 0.468 0.163
C3 -1.247 0.411 0.316
H4 -0.090 0.885 -1.433
H5 -0.012 2.124 -0.165
O6 1.291 -0.948 -0.080
O7 -1.512 -0.906 -0.148
H8 2.085 -1.331 0.326
H9 -0.668 -1.387 -0.082
H10 2.147 0.942 -0.349
H11 1.394 0.668 1.243
H12 -1.122 0.432 1.414
H13 -2.142 0.999 0.079

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 O6 O7 H8 H9 H10 H11 H12 H13
C11.52131.53031.09801.10062.39462.46023.24012.52542.16842.15962.16542.1655
C21.52132.54732.15382.13521.43683.14141.97192.71291.10121.10252.72193.4787
C31.53032.54732.14952.16582.90561.42113.75981.93003.49892.81031.10581.0967
H41.09802.15382.14951.77432.66352.62363.56842.70592.48603.06703.06192.5516
H51.10062.13522.16581.77433.33793.38134.07173.57302.46882.46552.56682.4207
O62.39461.43682.90562.66353.33792.80450.97082.00832.09172.09123.15573.9496
O72.46023.14141.42112.62363.38132.80453.65330.97344.10453.58572.09332.0194
H83.24011.97193.75983.56844.07170.97083.65332.78432.37182.30593.81814.8329
H92.52542.71291.93002.70593.57302.00830.97342.78433.66383.19902.39862.8090
H102.16841.10123.49892.48602.46882.09174.10452.37183.66381.78283.74924.3106
H112.15961.10252.81033.06702.46552.09123.58572.30593.19901.78282.53233.7365
H122.16542.72191.10583.06192.56683.15572.09333.81812.39863.74922.53231.7728
H132.16553.47871.09672.55162.42073.94962.01944.83292.80904.31063.73651.7728

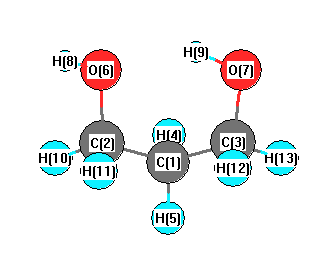
picture of 1,3-Propanediol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O6 108.065 C1 C2 H10 110.531
C1 C2 H11 109.757 C1 C3 O7 112.887
C1 C3 H12 109.404 C1 C3 H13 109.947
C2 C1 C3 113.177 C2 C1 H4 109.569
C2 C1 H5 107.979 C2 O6 H8 108.418
C3 C1 H4 108.621 C3 C1 H5 109.739
C3 O7 H9 105.901 H4 C1 H5 107.611
O6 C2 H10 110.308 O6 C2 H11 110.185
O7 C3 H12 111.264 O7 C3 H13 105.935
H10 C2 H11 107.999 H12 C3 H13 107.200
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability