Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G*
Geometric Data calculated at PBE1PBE/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -269.250726 |
| Energy at 298.15K | -269.260800 |
| HF Energy | -269.250726 |
| Nuclear repulsion energy | 195.619851 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3831 |
3641 |
28.09 |
|
|
|
| 2 |
A |
3733 |
3547 |
141.36 |
|
|
|
| 3 |
A |
3130 |
2974 |
30.07 |
|
|
|
| 4 |
A |
3111 |
2956 |
53.52 |
|
|
|
| 5 |
A |
3078 |
2925 |
36.37 |
|
|
|
| 6 |
A |
3059 |
2907 |
41.71 |
|
|
|
| 7 |
A |
3019 |
2869 |
60.55 |
|
|
|
| 8 |
A |
2976 |
2828 |
79.72 |
|
|
|
| 9 |
A |
1552 |
1475 |
2.75 |
|
|
|
| 10 |
A |
1539 |
1462 |
1.30 |
|
|
|
| 11 |
A |
1499 |
1424 |
96.85 |
|
|
|
| 12 |
A |
1487 |
1413 |
18.25 |
|
|
|
| 13 |
A |
1472 |
1399 |
5.14 |
|
|
|
| 14 |
A |
1419 |
1348 |
1.99 |
|
|
|
| 15 |
A |
1394 |
1325 |
0.16 |
|
|
|
| 16 |
A |
1328 |
1262 |
33.68 |
|
|
|
| 17 |
A |
1301 |
1237 |
3.05 |
|
|
|
| 18 |
A |
1245 |
1183 |
11.84 |
|
|
|
| 19 |
A |
1220 |
1159 |
64.48 |
|
|
|
| 20 |
A |
1150 |
1093 |
36.15 |
|
|
|
| 21 |
A |
1140 |
1083 |
73.85 |
|
|
|
| 22 |
A |
1123 |
1067 |
52.04 |
|
|
|
| 23 |
A |
990 |
941 |
1.36 |
|
|
|
| 24 |
A |
925 |
879 |
12.13 |
|
|
|
| 25 |
A |
924 |
878 |
1.30 |
|
|
|
| 26 |
A |
842 |
800 |
6.45 |
|
|
|
| 27 |
A |
572 |
544 |
167.95 |
|
|
|
| 28 |
A |
509 |
484 |
2.81 |
|
|
|
| 29 |
A |
394 |
374 |
4.32 |
|
|
|
| 30 |
A |
340 |
323 |
20.88 |
|
|
|
| 31 |
A |
290 |
276 |
71.74 |
|
|
|
| 32 |
A |
223 |
212 |
2.24 |
|
|
|
| 33 |
A |
110 |
105 |
0.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25461.6 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 24196.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.033 |
1.041 |
-0.343 |
| C2 |
1.271 |
0.472 |
0.182 |
| C3 |
-1.263 |
0.395 |
0.290 |
| H4 |
-0.081 |
0.904 |
-1.430 |
| H5 |
-0.037 |
2.122 |
-0.146 |
| O6 |
1.298 |
-0.923 |
-0.102 |
| O7 |
-1.464 |
-0.932 |
-0.129 |
| H8 |
2.062 |
-1.313 |
0.342 |
| H9 |
-0.594 |
-1.360 |
-0.069 |
| H10 |
2.128 |
0.977 |
-0.290 |
| H11 |
1.336 |
0.645 |
1.268 |
| H12 |
-1.180 |
0.466 |
1.391 |
| H13 |
-2.162 |
0.953 |
0.005 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
H4 |
H5 |
O6 |
O7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5173 | 1.5267 | 1.0969 | 1.0981 | 2.3856 | 2.4465 | 3.2251 | 2.4811 | 2.1629 | 2.1514 | 2.1565 | 2.1590 |
C2 | 1.5173 | | 2.5380 | 2.1480 | 2.1308 | 1.4237 | 3.0897 | 1.9584 | 2.6263 | 1.1005 | 1.1020 | 2.7330 | 3.4718 | C3 | 1.5267 | 2.5380 | | 2.1486 | 2.1623 | 2.9073 | 1.4055 | 3.7382 | 1.9122 | 3.4891 | 2.7884 | 1.1066 | 1.0960 | H4 | 1.0969 | 2.1480 | 2.1486 | | 1.7702 | 2.6469 | 2.6417 | 3.5562 | 2.6913 | 2.4868 | 3.0588 | 3.0592 | 2.5288 | H5 | 1.0981 | 2.1308 | 2.1623 | 1.7702 | | 3.3249 | 3.3705 | 4.0543 | 3.5267 | 2.4529 | 2.4627 | 2.5312 | 2.4305 | O6 | 2.3856 | 1.4237 | 2.9073 | 2.6469 | 3.3249 | | 2.7622 | 0.9652 | 1.9424 | 2.0817 | 2.0825 | 3.2094 | 3.9380 | O7 | 2.4465 | 3.0897 | 1.4055 | 2.6417 | 3.3705 | 2.7622 | | 3.5770 | 0.9712 | 4.0705 | 3.5035 | 2.0845 | 2.0144 | H8 | 3.2251 | 1.9584 | 3.7382 | 3.5562 | 4.0543 | 0.9652 | 3.5770 | | 2.6877 | 2.3761 | 2.2841 | 3.8436 | 4.8052 | H9 | 2.4811 | 2.6263 | 1.9122 | 2.6913 | 3.5267 | 1.9424 | 0.9712 | 2.6877 | | 3.5941 | 3.0874 | 2.4106 | 2.7954 | H10 | 2.1629 | 1.1005 | 3.4891 | 2.4868 | 2.4529 | 2.0817 | 4.0705 | 2.3761 | 3.5941 | | 1.7789 | 3.7450 | 4.3005 | H11 | 2.1514 | 1.1020 | 2.7884 | 3.0588 | 2.4627 | 2.0825 | 3.5035 | 2.2841 | 3.0874 | 1.7789 | | 2.5249 | 3.7323 | H12 | 2.1565 | 2.7330 | 1.1066 | 3.0592 | 2.5312 | 3.2094 | 2.0845 | 3.8436 | 2.4106 | 3.7450 | 2.5249 | | 1.7675 | H13 | 2.1590 | 3.4718 | 1.0960 | 2.5288 | 2.4305 | 3.9380 | 2.0144 | 4.8052 | 2.7954 | 4.3005 | 3.7323 | 1.7675 | |
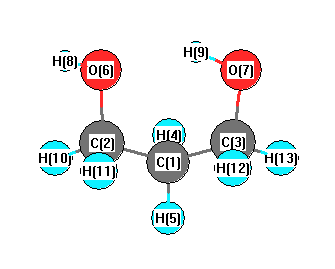 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
108.378 |
|
C1 |
C2 |
H10 |
110.416 |
| C1 |
C2 |
H11 |
109.419 |
|
C1 |
C3 |
O7 |
113.033 |
| C1 |
C3 |
H12 |
108.918 |
|
C1 |
C3 |
H13 |
109.725 |
| C2 |
C1 |
C3 |
112.974 |
|
C2 |
C1 |
H4 |
109.448 |
| C2 |
C1 |
H5 |
108.049 |
|
C2 |
O6 |
H8 |
108.603 |
| C3 |
C1 |
H4 |
108.858 |
|
C3 |
C1 |
H5 |
109.858 |
| C3 |
O7 |
H9 |
105.675 |
|
H4 |
C1 |
H5 |
107.509 |
| O6 |
C2 |
H10 |
110.456 |
|
O6 |
C2 |
H11 |
110.434 |
| O7 |
C3 |
H12 |
111.598 |
|
O7 |
C3 |
H13 |
106.616 |
| H10 |
C2 |
H11 |
107.736 |
|
H12 |
C3 |
H13 |
106.733 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.375 |
|
|
|
| 2 |
C |
-0.092 |
|
|
|
| 3 |
C |
-0.066 |
|
|
|
| 4 |
H |
0.193 |
|
|
|
| 5 |
H |
0.168 |
|
|
|
| 6 |
O |
-0.643 |
|
|
|
| 7 |
O |
-0.651 |
|
|
|
| 8 |
H |
0.420 |
|
|
|
| 9 |
H |
0.432 |
|
|
|
| 10 |
H |
0.161 |
|
|
|
| 11 |
H |
0.153 |
|
|
|
| 12 |
H |
0.131 |
|
|
|
| 13 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.127 |
1.286 |
1.265 |
3.610 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.306 |
-4.533 |
1.380 |
| y |
-4.533 |
-29.952 |
-1.459 |
| z |
1.380 |
-1.459 |
-31.499 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.420 |
-4.533 |
1.380 |
| y |
-4.533 |
0.950 |
-1.459 |
| z |
1.380 |
-1.459 |
-1.370 |
|
| Polar |
|---|
| 3z2-r2 | -2.741 |
|---|
| x2-y2 | -0.353 |
|---|
| xy | -4.533 |
|---|
| xz | 1.380 |
|---|
| yz | -1.459 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.400 |
-0.187 |
0.129 |
| y |
-0.187 |
6.014 |
0.036 |
| z |
0.129 |
0.036 |
5.236 |
<r2> (average value of r
2) Å
2
| <r2> |
127.367 |
| (<r2>)1/2 |
11.286 |
