Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/6-31G*
Geometric Data calculated at B2PLYP/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -269.262654 |
| Energy at 298.15K | -269.272726 |
| HF Energy | -269.016140 |
| Nuclear repulsion energy | 194.819391 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3784 |
3592 |
22.34 |
|
|
|
| 2 |
A |
3720 |
3531 |
122.26 |
|
|
|
| 3 |
A |
3135 |
2975 |
34.10 |
|
|
|
| 4 |
A |
3125 |
2966 |
52.12 |
|
|
|
| 5 |
A |
3083 |
2927 |
57.48 |
|
|
|
| 6 |
A |
3073 |
2917 |
24.16 |
|
|
|
| 7 |
A |
3036 |
2882 |
57.49 |
|
|
|
| 8 |
A |
2997 |
2845 |
73.77 |
|
|
|
| 9 |
A |
1575 |
1495 |
2.19 |
|
|
|
| 10 |
A |
1561 |
1482 |
0.20 |
|
|
|
| 11 |
A |
1516 |
1439 |
20.43 |
|
|
|
| 12 |
A |
1499 |
1423 |
87.26 |
|
|
|
| 13 |
A |
1485 |
1409 |
9.63 |
|
|
|
| 14 |
A |
1429 |
1357 |
1.65 |
|
|
|
| 15 |
A |
1412 |
1340 |
0.11 |
|
|
|
| 16 |
A |
1335 |
1267 |
30.58 |
|
|
|
| 17 |
A |
1306 |
1240 |
2.56 |
|
|
|
| 18 |
A |
1249 |
1185 |
17.27 |
|
|
|
| 19 |
A |
1223 |
1161 |
53.76 |
|
|
|
| 20 |
A |
1160 |
1101 |
28.27 |
|
|
|
| 21 |
A |
1119 |
1062 |
62.46 |
|
|
|
| 22 |
A |
1107 |
1050 |
67.66 |
|
|
|
| 23 |
A |
980 |
930 |
1.64 |
|
|
|
| 24 |
A |
935 |
887 |
1.65 |
|
|
|
| 25 |
A |
927 |
880 |
15.49 |
|
|
|
| 26 |
A |
837 |
794 |
8.38 |
|
|
|
| 27 |
A |
565 |
536 |
169.60 |
|
|
|
| 28 |
A |
516 |
489 |
2.70 |
|
|
|
| 29 |
A |
393 |
373 |
4.26 |
|
|
|
| 30 |
A |
340 |
322 |
20.86 |
|
|
|
| 31 |
A |
289 |
274 |
74.13 |
|
|
|
| 32 |
A |
221 |
210 |
2.56 |
|
|
|
| 33 |
A |
111 |
105 |
0.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25520.5 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24224.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.017 |
1.029 |
-0.361 |
| C2 |
1.284 |
0.467 |
0.183 |
| C3 |
-1.242 |
0.415 |
0.312 |
| H4 |
-0.070 |
0.806 |
-1.435 |
| H5 |
-0.014 |
2.123 |
-0.244 |
| O6 |
1.288 |
-0.938 |
-0.089 |
| O7 |
-1.504 |
-0.904 |
-0.140 |
| H8 |
2.089 |
-1.325 |
0.292 |
| H9 |
-0.665 |
-1.385 |
-0.062 |
| H10 |
2.150 |
0.966 |
-0.285 |
| H11 |
1.337 |
0.646 |
1.271 |
| H12 |
-1.102 |
0.431 |
1.408 |
| H13 |
-2.142 |
1.001 |
0.086 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
H4 |
H5 |
O6 |
O7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5177 | 1.5265 | 1.0981 | 1.1008 | 2.3759 | 2.4482 | 3.2251 | 2.5176 | 2.1690 | 2.1547 | 2.1600 | 2.1718 |
C2 | 1.5177 | | 2.5295 | 2.1365 | 2.1470 | 1.4309 | 3.1231 | 1.9675 | 2.6998 | 1.1034 | 1.1038 | 2.6822 | 3.4687 | C3 | 1.5265 | 2.5295 | | 2.1398 | 2.1759 | 2.8965 | 1.4181 | 3.7579 | 1.9269 | 3.4875 | 2.7609 | 1.1058 | 1.0978 | H4 | 1.0981 | 2.1365 | 2.1398 | | 1.7771 | 2.5879 | 2.5801 | 3.4903 | 2.6541 | 2.5050 | 3.0538 | 3.0483 | 2.5780 | H5 | 1.1008 | 2.1470 | 2.1759 | 1.7771 | | 3.3302 | 3.3752 | 4.0743 | 3.5732 | 2.4542 | 2.5103 | 2.6029 | 2.4282 | O6 | 2.3759 | 1.4309 | 2.8965 | 2.5879 | 3.3302 | | 2.7924 | 0.9675 | 2.0036 | 2.0992 | 2.0874 | 3.1346 | 3.9439 | O7 | 2.4482 | 3.1231 | 1.4181 | 2.5801 | 3.3752 | 2.7924 | | 3.6431 | 0.9706 | 4.1067 | 3.5298 | 2.0837 | 2.0213 | H8 | 3.2251 | 1.9675 | 3.7579 | 3.4903 | 4.0743 | 0.9675 | 3.6431 | | 2.7771 | 2.3632 | 2.3253 | 3.8094 | 4.8327 | H9 | 2.5176 | 2.6998 | 1.9269 | 2.6541 | 3.5732 | 2.0036 | 0.9706 | 2.7771 | | 3.6745 | 3.1474 | 2.3773 | 2.8104 | H10 | 2.1690 | 1.1034 | 3.4875 | 2.5050 | 2.4542 | 2.0992 | 4.1067 | 2.3632 | 3.6745 | | 1.7841 | 3.7046 | 4.3080 | H11 | 2.1547 | 1.1038 | 2.7609 | 3.0538 | 2.5103 | 2.0874 | 3.5298 | 2.3253 | 3.1474 | 1.7841 | | 2.4517 | 3.6924 | H12 | 2.1600 | 2.6822 | 1.1058 | 3.0483 | 2.6029 | 3.1346 | 2.0837 | 3.8094 | 2.3773 | 3.7046 | 2.4517 | | 1.7769 | H13 | 2.1718 | 3.4687 | 1.0978 | 2.5780 | 2.4282 | 3.9439 | 2.0213 | 4.8327 | 2.8104 | 4.3080 | 3.6924 | 1.7769 | |
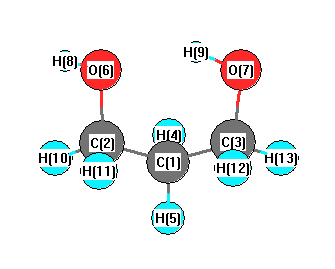 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
107.332 |
|
C1 |
C2 |
H10 |
110.691 |
| C1 |
C2 |
H11 |
109.544 |
|
C1 |
C3 |
O7 |
112.442 |
| C1 |
C3 |
H12 |
109.237 |
|
C1 |
C3 |
H13 |
110.636 |
| C2 |
C1 |
C3 |
112.385 |
|
C2 |
C1 |
H4 |
108.453 |
| C2 |
C1 |
H5 |
109.120 |
|
C2 |
O6 |
H8 |
108.700 |
| C3 |
C1 |
H4 |
108.122 |
|
C3 |
C1 |
H5 |
110.786 |
| C3 |
O7 |
H9 |
106.023 |
|
H4 |
C1 |
H5 |
107.840 |
| O6 |
C2 |
H10 |
111.196 |
|
O6 |
C2 |
H11 |
110.219 |
| O7 |
C3 |
H12 |
110.685 |
|
O7 |
C3 |
H13 |
106.217 |
| H10 |
C2 |
H11 |
107.863 |
|
H12 |
C3 |
H13 |
107.483 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.327 |
|
|
|
| 2 |
C |
-0.026 |
|
|
|
| 3 |
C |
0.001 |
|
|
|
| 4 |
H |
0.174 |
|
|
|
| 5 |
H |
0.146 |
|
|
|
| 6 |
O |
-0.686 |
|
|
|
| 7 |
O |
-0.697 |
|
|
|
| 8 |
H |
0.422 |
|
|
|
| 9 |
H |
0.443 |
|
|
|
| 10 |
H |
0.145 |
|
|
|
| 11 |
H |
0.137 |
|
|
|
| 12 |
H |
0.116 |
|
|
|
| 13 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.327 |
-0.205 |
0.136 |
| y |
-0.205 |
6.009 |
0.043 |
| z |
0.136 |
0.043 |
5.165 |
<r2> (average value of r
2) Å
2
| <r2> |
128.498 |
| (<r2>)1/2 |
11.336 |
