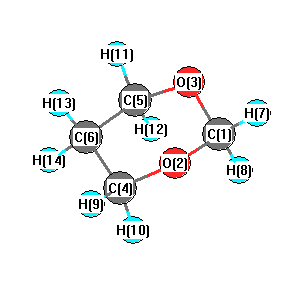
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3158 |
3022 |
35.96 |
|
|
|
| 2 |
A' |
3131 |
2996 |
31.33 |
|
|
|
| 3 |
A' |
3121 |
2986 |
23.97 |
|
|
|
| 4 |
A' |
3069 |
2937 |
19.48 |
|
|
|
| 5 |
A' |
2987 |
2858 |
110.64 |
|
|
|
| 6 |
A' |
2935 |
2808 |
81.68 |
|
|
|
| 7 |
A' |
1536 |
1469 |
5.69 |
|
|
|
| 8 |
A' |
1521 |
1455 |
0.16 |
|
|
|
| 9 |
A' |
1492 |
1427 |
5.34 |
|
|
|
| 10 |
A' |
1428 |
1367 |
23.81 |
|
|
|
| 11 |
A' |
1333 |
1276 |
5.70 |
|
|
|
| 12 |
A' |
1223 |
1170 |
113.18 |
|
|
|
| 13 |
A' |
1198 |
1146 |
79.09 |
|
|
|
| 14 |
A' |
1127 |
1078 |
13.00 |
|
|
|
| 15 |
A' |
1022 |
978 |
39.17 |
|
|
|
| 16 |
A' |
930 |
890 |
11.01 |
|
|
|
| 17 |
A' |
860 |
823 |
9.95 |
|
|
|
| 18 |
A' |
653 |
625 |
5.02 |
|
|
|
| 19 |
A' |
491 |
470 |
0.59 |
|
|
|
| 20 |
A' |
436 |
417 |
9.95 |
|
|
|
| 21 |
A' |
268 |
257 |
1.95 |
|
|
|
| 22 |
A" |
3127 |
2992 |
46.87 |
|
|
|
| 23 |
A" |
2979 |
2850 |
21.69 |
|
|
|
| 24 |
A" |
1519 |
1453 |
5.12 |
|
|
|
| 25 |
A" |
1462 |
1399 |
17.20 |
|
|
|
| 26 |
A" |
1402 |
1342 |
0.79 |
|
|
|
| 27 |
A" |
1381 |
1321 |
0.12 |
|
|
|
| 28 |
A" |
1348 |
1289 |
1.22 |
|
|
|
| 29 |
A" |
1270 |
1215 |
32.01 |
|
|
|
| 30 |
A" |
1242 |
1189 |
0.13 |
|
|
|
| 31 |
A" |
1098 |
1051 |
65.63 |
|
|
|
| 32 |
A" |
1047 |
1002 |
67.51 |
|
|
|
| 33 |
A" |
935 |
895 |
20.44 |
|
|
|
| 34 |
A" |
907 |
868 |
0.04 |
|
|
|
| 35 |
A" |
464 |
444 |
7.17 |
|
|
|
| 36 |
A" |
266 |
254 |
1.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27184.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 26007.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.130 |
|
|
|
| 2 |
O |
-0.456 |
|
|
|
| 3 |
O |
-0.456 |
|
|
|
| 4 |
C |
-0.092 |
|
|
|
| 5 |
C |
-0.092 |
|
|
|
| 6 |
C |
-0.344 |
|
|
|
| 7 |
H |
0.182 |
|
|
|
| 8 |
H |
0.128 |
|
|
|
| 9 |
H |
0.180 |
|
|
|
| 10 |
H |
0.144 |
|
|
|
| 11 |
H |
0.180 |
|
|
|
| 12 |
H |
0.144 |
|
|
|
| 13 |
H |
0.187 |
|
|
|
| 14 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.653 |
1.888 |
0.000 |
1.998 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.552 |
1.655 |
0.000 |
| y |
1.655 |
-34.568 |
0.000 |
| z |
0.000 |
0.000 |
-38.336 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.900 |
1.655 |
0.000 |
| y |
1.655 |
2.376 |
0.000 |
| z |
0.000 |
0.000 |
-3.277 |
|
| Polar |
|---|
| 3z2-r2 | -6.553 |
|---|
| x2-y2 | -0.984 |
|---|
| xy | 1.655 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.477 |
0.441 |
0.000 |
| y |
0.441 |
7.418 |
0.000 |
| z |
0.000 |
0.000 |
6.912 |
<r2> (average value of r
2) Å
2
| <r2> |
138.710 |
| (<r2>)1/2 |
11.778 |